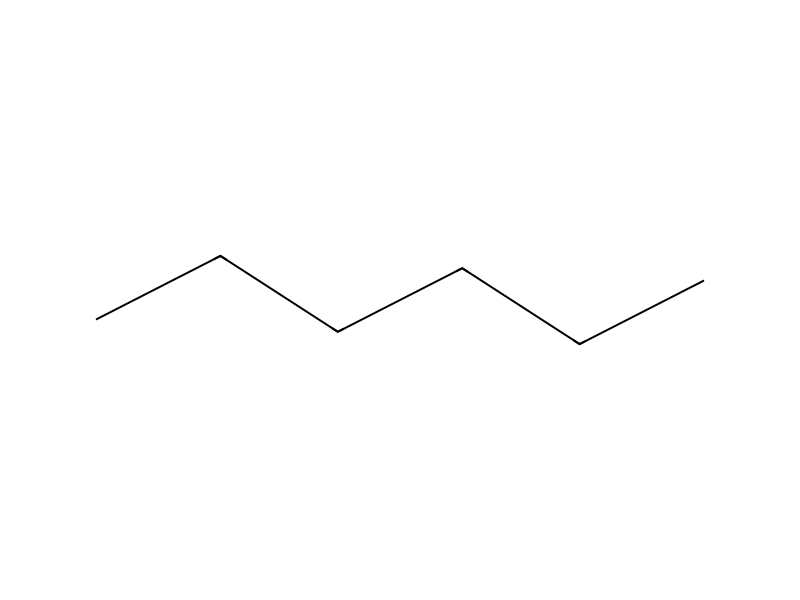
Hexane → 5-chloro-2'-pyridin-4-yl-6',7'-dihydro-3H,5'H-spiro[2-benzofuran-1,4'-[1]benzothiophen]-3-one
Reaction equation

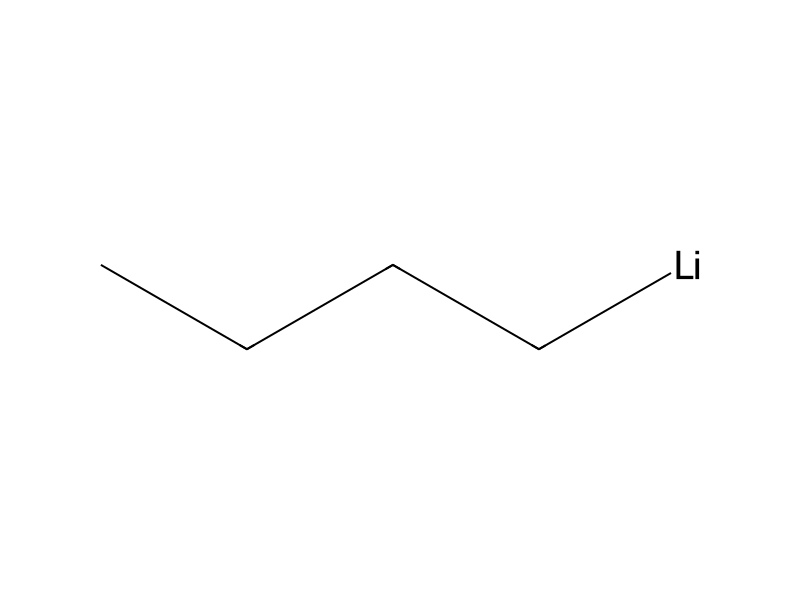
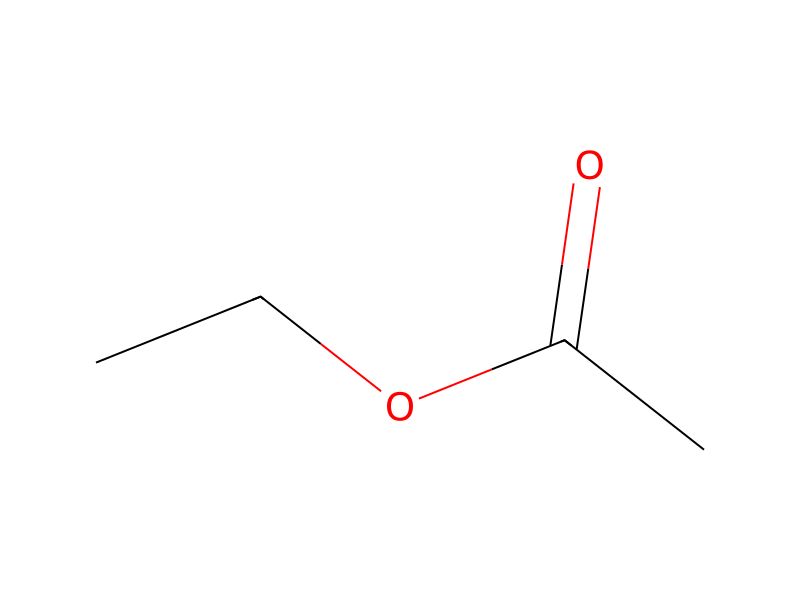
[Cl:1][C:2]1[CH:3]=[CH:4][C:5](I)=[C:6]([CH:14]=1)[C:7](N(CC)CC)=[O:8].C([Li])CCC.CCCCCC.[N:27]1[CH:32]=[CH:31][C:30]([C:33]2[S:34][C:35]3[CH2:41][CH2:40][CH2:39][C:38](=[O:42])[C:36]=3[CH:37]=2)=[CH:29][CH:28]=1>O1CCCC1.CCOC(C)=O>[Cl:1][C:2]1[CH:3]=[CH:4][C:5]2[C:38]3([O:42][C:7](=[O:8])[C:6]=2[CH:14]=1)[C:36]1[CH:37]=[C:33]([C:30]2[CH:31]=[CH:32][N:27]=[CH:28][CH:29]=2)[S:34][C:35]=1[CH2:41][CH2:40][CH2:39]3Participants
Reactant 4


Product 1

Reagents, catalysts & solvents 2

Conditions
- Temperature
- -78 °C
Procedure
In a 100 mL round bottomed flask was placed 5-chloro-N,N-diethyl-2-iodobenzamide (840 mg, 2.5 mmol) and tetrahydrofuran (10 mL). The mixture was stirred for 5 min at −78° C. then n-butyllithium in hexane (1.6M, 1.8 mL, 2.9 mmol) was added and the mixture was stirred for an additional 30 min at the same temperature. To the mixture was added a solution of 2-pyridin-4-yl-6,7-dihydro-1-benzothiophen-4(5H)-one (445 mg, 1.94 mmol) in tetrahydrofuran (15 mL). The mixture was stirred for an additional 16 hr during which time the reaction temperature was allowed to slowly rise from −78° C. to rt. The reaction mixture was then refluxed for 4 hr, cooled to rt and then diluted with EtOAc (50 mL). The resulting solution was washed with water (50 mL). The aqueous phase was separated and neutralized by the addition of 1N HCl (ca 1.21 mL, pH ca 7.0) and then extracted with EtOAc (50 mL). Finally, the combined organic phases were washed with brine (30 mL) then dried over anhydrous MgSO4. Insoluble materials were removed by filtration and the filtrate was concentrated under reduced pressure. The residue was purified by pre-packed silica gel (40 g) column chromatography, eluting with 30% to 100% EtOAc/hexane to yield the title compound (550 mg; 77%) as an off-white foam, which included a small amount of 2-pyridin-4-yl-1-benzothiophene-4-ol as an impurity. The crude material was used in the next step without further purification. LCMS: (FA) ES+ 368, 370; 1H NMR (400 MHz, d6-DMSO) δ ppm 8.46 (dd, 2H), 7.98 (dd, 1H), 7.82 (dd, 1H), 7.57 (d, 1H), 7.49 (dd, 2H), 7.11 (s, 1H), 3.03-3.09 (m, 1H), 2.89-2.97 (m, 1H), 2.29-2.37 (m, 1H), 2.00-2.16 (m, 3H).
Workup
- customIn a 100 mL round bottomed flask was placed
- stirringthe mixture was stirred for an additional 30 min at the same temperature
- stirringThe mixture was stirred for an additional 16 hr during which time the reaction temperature
- customto slowly rise from −78° C. to rt
- temperatureThe reaction mixture was then refluxed for 4 hr
- temperaturecooled to rt
- washThe resulting solution was washed with water (50 mL)
- customThe aqueous phase was separated
- additionneutralized by the addition of 1N HCl (ca 1.21 mL, pH ca 7.0)
- extractionextracted with EtOAc (50 mL)
- washFinally, the combined organic phases were washed with brine (30 mL)
- dry with materialthen dried over anhydrous MgSO4
- customInsoluble materials were removed by filtration
- concentrationthe filtrate was concentrated under reduced pressure
- customThe residue was purified by pre-packed silica gel (40 g) column chromatography
- washeluting with 30% to 100% EtOAc/hexane
Sources & evidence
- ORD record
- ord-058d628454b3456cac4aa745d99403d7
- Source dataset
- uspto-grants-2014_07
- Patent number
- US08765746B2
Reaction notes
Loading…