Acetonitrile → tert-butyl 2-(4-(hydroxymethyl)-1H-1,2,3-triazol-1-yl)benzylcarbamate
Reaction equation

[N:1]([C:4]1[CH:18]=[CH:17][CH:16]=[CH:15][C:5]=1[CH2:6][NH:7][C:8](=[O:14])[O:9][C:10]([CH3:13])([CH3:12])[CH3:11])=[N+:2]=[N-:3].[C:19](#N)[CH3:20].[C:22]([O:26]N=O)(C)(C)C.C[Si](N=[N+]=[N-])(C)C>C(O)C#C>[OH:26][CH2:22][C:19]1[N:3]=[N:2][N:1]([C:4]2[CH:18]=[CH:17][CH:16]=[CH:15][C:5]=2[CH2:6][NH:7][C:8](=[O:14])[O:9][C:10]([CH3:13])([CH3:12])[CH3:11])[CH:20]=1Participants
Reactant 4
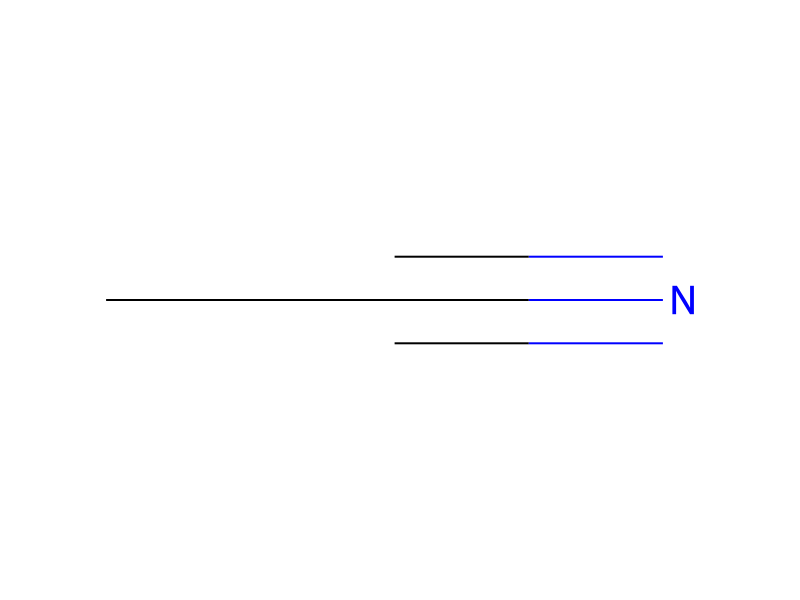

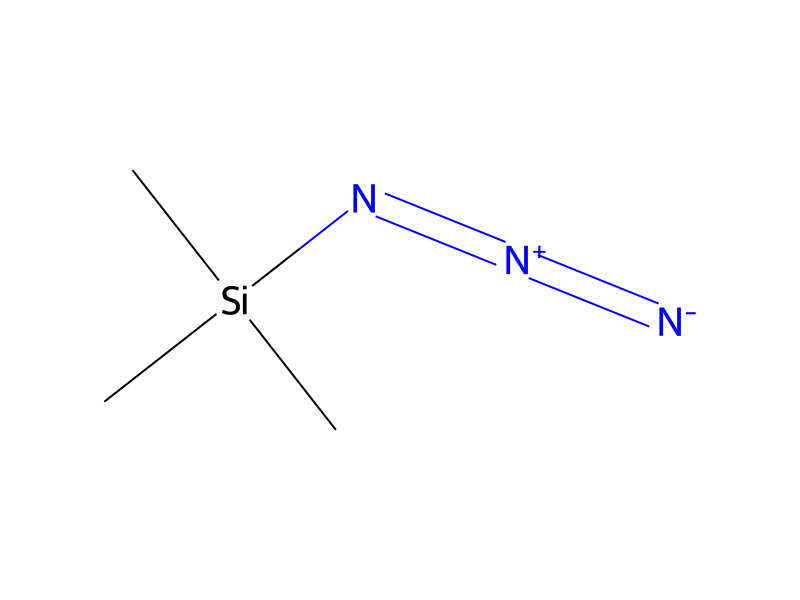

Product 1

Reagents, catalysts & solvents 1

Conditions
- Temperature
- 0 °C
Procedure
To a THF (30 mL) solution of 2-aminobenzyl amine (2.0 g, 16.4 mmol) was added triethylamine (4.60 mL, 32.7 mmol) and di-tert-butyldicarbonate (4.3 g, 19.6 mmol). The reaction mixture was stirred at room temperature for 16 h. The crude reaction mixture was then poured over saturated aqueous NaHCO3 and the product was then extracted with ethyl acetate. The organic phase was washed with brine and then dried over MgSO4, filtered and concentrated in vacuo. The crude product was purified by silica gel chromatography (0-5% methanol/dichloromethane) yielding 730 mg (20%) of 11: 1H NMR (400 MHz, CDCl3) δ 7.12-7.80, (m, 1H), 7.00 (d, J=7.6 Hz, 1H), 6.69-6.62 (m, 2H), 4.77, (bs, 1H), 4.29-4.11, (m, 4H), 1.43 (s, 9H); ESI+ MS: m/z (rel intensity) 223 (80, M+H). Step b: Preparation of tert-butyl 2-azidobenzylcarbamate (12). To a acetonitrile (40 mL) solution of 15 (5.00 g, 22.5 mmol) at 0° C. was added tert-butylnitrite (4.00 mL, 33.7 mmol) and trimethylsilylazide (33.55 mL, 26.9 mmol). The reaction stirred at 0° C. for 3 h. The reaction mixture was concentrated in vacuo, and the crude material was taken on to the next step without further purification. ESI+ MS: m/z (rel intensity) 271 (80, M+H). Step c: Preparation of tert-butyl 2-(4-(hydroxymethyl)-1H-1,2,3-triazol-1-yl)benzylcarbamate (13). tert-butyl 2-azidobenzylcarbamate, 12, (5.4 g, 21.7 mmol) was dissolved in neat propargyl alcohol (10 mL). The reaction mixture was stirred at 90° C. for 18 h. The reaction mixture was then concentrated in vacuo and the resulting residue dissolved in diethyl ether. Upon standing, triazole 13 crystallized. The crystals were collected by filtration yielding 2.5 g (38%) of 13: 1H NMR (400 MHz, CDCl3) δ 7.82, (s, 1H), 7.64-7.35 (m, 3H), 7.34-7.30 (m, 1H), 5.35, (bs, 1H), 4.87, (s, 2H), 4.15 (s, 2H), 1.42 (s, 9H); ESI+ MS: m/z (rel intensity) 249 (100, M-tBu). Step d: Preparation of tert-butyl 2-(4-formyl-1H-1,2,3-triazol-1-yl)benzylcarbamate (14). To a solution of tert-butyl 2-(4-(hydroxymethyl)-1H-1,2,3-triazol-1-yl)benzylcarbamate, 13, (2.5 g, 8.31 mmol) in CH2Cl2 (10 mL) was added MnO2 (5.0 g, 57.5 mmol). The reaction mixture was stirred at room temperature for 18 h. The crude reaction mixture was then filtered through a plug of Celite® and then concentrated in vacuo. The crude material was purified by silica gel chromatography (0-5% methanol/dichloromethane) yielding 610 mg (25%) of 14: ESI+ MS: m/z (rel intensity) 225 (100, M+Na). Step e: Preparation of tert-butyl(2-(4-((methyl(5,6,7,8-tetrahydroquinolin-8-yl)amino)methyl)-1H-1,2,3-triazol-1-yl)phenyl) (15). To a solution of tert-butyl 2-(4-formyl-1H-1,2,3-triazol-1-yl)benzylcarbamate, 14, (0.61 g, 2.03 mmol) in 1,2-dichloroethane (4 mL) was added N-methyl-5,6,7,8-tetrahydroquinolin-8-amine, 11, (0.30 g, 1.85 mmol), sodium triacetoxyborohydride (0.70 g, 3.33 mmol), and AcOH (10 drops). The reaction mixture was heated to 65° C. and stirred for 18 h. The reaction mixture was poured into saturated aqueous NaHCO3 (10 mL) and was extracted with EtOAc (2×10 mL). The organic phases were combined and washed with brine, dried over MgSO4, filtered, and concentrated in vacuo. The crude material was purified by silica gel chromatography (0-5% methanol/dichloromethane) yielding 360 mg (44%) of 15: 1H NMR (400 MHz, CDCl3) δ 8.48 (d, J=4.4 Hz, 1H), 8.04 (s, 1H), 7.79 (bs, 1H), 7.64 (d, J=7.6 Hz, 1H), 7.47-7.30 (m, 4H), 7.08 (dd, J=8.0, 4.8 Hz, 1H), 4.18-3.95 (m, 5H), 2.88-2.65, (m, 2H), 2.41, (s, 3H), 2.24-2.14, (m, 1H), 2.10-1.90, (m, 2H), 1.76-1.64 (m, 1H), 1.39 (s, 9H); ESI+ MS: m/z (rel intensity) 449 (100, M+H). Step f: Preparation of N-((1-(2-(aminomethyl)phenyl)-1H-1,2,3-triazol-4-yl)methyl)-N-methyl-5,6,7,8-tetrahydroquinolin-8-amine tetrahydrochloride (AD). Through a methanol (2 mL) solution of tert-butyl(2-(4-((methyl(5,6,7,8-tetrahydroquinolin-8-yl)amino)methyl)-1H-1,2,3-triazol-1-yl)phenyl), 15, (0.36 g, 0.80 mmol) was bubbled HCl(g) (generated by adding conc. H2SO4 to dry NaCl) until complete conversion was observed by LC-MS. The reaction mixture was then concentrated in vacuo yielding 250 mg (68%) of AD: 1H NMR (400 MHz, d6-DMSO) δ 8.81-8.68, (m, 4H), 8.52, (d, J=4.4 Hz, 1H), 7.87, (d, J=6.8 Hz, 1H), 7.75-7.50, (m, 4H), 7.38, (dd, J=7.6, 4.8 Hz, 1H), 4.81-4.73, (m, 1H), 4.61, (d, J=13.6 Hz, 1H), 4.47, (d, J=13.6 Hz, 1H), 3.91-3.83, (m, 2H), 2.90-2.70, (m, 5H), 2.51-2.49, (m, 1H), 2.15-2.03, (m, 2H), 1.72-1.60, (m, 1H); ESI+ MS: m/z (rel intensity) 349 (100, M+H).
Workup
- concentrationThe reaction mixture was concentrated in vacuo
- customthe crude material was taken on to the next step without further purification
- customStep c: Preparation of tert-butyl 2-(4-(hydroxymethyl)-1H-1,2,3-triazol-1-yl)benzylcarbamate (13)
- stirringThe reaction mixture was stirred at 90° C. for 18 h
- concentrationThe reaction mixture was then concentrated in vacuo
- dissolutionthe resulting residue dissolved in diethyl ether
- customtriazole 13 crystallized
- filtrationThe crystals were collected by filtration
Sources & evidence
- ORD record
- ord-c629cb0e3cb441b6925ee4c900d2649d
- Source dataset
- uspto-grants-2012_12
- Patent number
- US08338448B2
Reaction notes
Loading…