Azobisisobutyronitrile → 2-(4-Fluoro-2-methoxy-phenyl)-5-(5-methoxymethyl-3-phenyl-isoxazol-4-yl)-[1,3,4]oxadiazole
Reaction equation

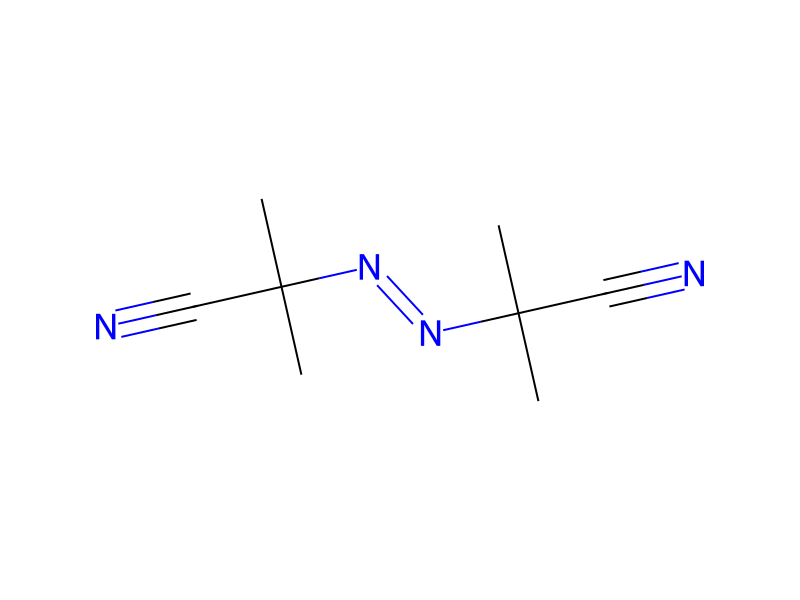
[F:1][C:2]1[CH:7]=[CH:6][C:5]([C:8]2[O:9][C:10]([C:13]3[C:14]([C:19]4[CH:24]=[CH:23][CH:22]=[CH:21][CH:20]=4)=[N:15][O:16][C:17]=3[CH3:18])=[N:11][N:12]=2)=[C:4]([O:25][CH3:26])[CH:3]=1.BrN1[C:32](=[O:33])CCC1=O.N(C(C)(C)C#N)=NC(C)(C)C#N.C[O-].[Na+]>C(Cl)(Cl)(Cl)Cl.C(OCC)(=O)C>[F:1][C:2]1[CH:7]=[CH:6][C:5]([C:8]2[O:9][C:10]([C:13]3[C:14]([C:19]4[CH:24]=[CH:23][CH:22]=[CH:21][CH:20]=4)=[N:15][O:16][C:17]=3[CH2:18][O:33][CH3:32])=[N:11][N:12]=2)=[C:4]([O:25][CH3:26])[CH:3]=1Participants
Reactant 4


Product 1

Reagents, catalysts & solvents 2
Conditions
- Temperature
- 70 °C
Procedure
To a solution of 2-(4-fluoro-2-methoxy-phenyl)-5-(5-methyl-3-phenyl-isoxazol-4-yl)-[1,3,4]oxadiazole (1.95 g, 5.56 mmol) in carbon tetrachloride (20 ml) was added N-bromo succinimide (990 mg, 5.56 mmol) and 2,2′-azobis(2-methylpropionitrile) (46 mg, 0.28 mmol) and the reaction mixture was stirred for 4 h at 70° C. Further 2,2′-azobis(2-methylpropionitrile) (46 mg, 0.28 mmol) was added and stirring was continued for another 14 h at 70° C. N-Bromo succinimide (378 mg, 2.12 mmol) was added and the mixture stirred for another 4 h at 90° C. After the reaction mixture was cooled to 0° C. the suspension was filtered off and was washed with dichloromethane (30 mL) and aqueous sodium hydrogencarbonate (1 N). It was extracted with dichloroemthane and dried over sodium sulfate resulting in crude material (1.66 g). A part of this light brown solid (200 mg) was dissolved in tetrahydropyran (2 mL), sodium methoxide (30% in methanol, 0.43 ml, 2.32 mmol) was added and the resulting mixture was stirred for 1 h at ambient temperature. It was diluted with ethyl acetate (20 mL) and washed aqueous ammonium chloride (saturated). The aqueous layers were extracted with ethyl acetate and the combined organic layers were dried over sodium sulfate. Purification by chromatography (SiO2, heptane:ethyl acetate:dichloromethane=70:10:20 to 40:40:20) afforded the title compound (25 mg, 14%) as a white solid. MS: m/e=382.3 [M+H]+.
Workup
- stirringstirring
- waitwas continued for another 14 h at 70° C
- stirringthe mixture stirred for another 4 h at 90° C
- temperatureAfter the reaction mixture was cooled to 0° C. the suspension
- filtrationwas filtered off
- washwas washed with dichloromethane (30 mL) and aqueous sodium hydrogencarbonate (1 N)
- extractionIt was extracted with dichloroemthane
- dry with materialdried over sodium sulfate resulting in crude material (1.66 g)
- dissolutionA part of this light brown solid (200 mg) was dissolved in tetrahydropyran (2 mL)
- stirringthe resulting mixture was stirred for 1 h at ambient temperature
- washwashed aqueous ammonium chloride (saturated)
- extractionThe aqueous layers were extracted with ethyl acetate
- dry with materialthe combined organic layers were dried over sodium sulfate
- customPurification by chromatography (SiO2, heptane:ethyl acetate:dichloromethane=70:10:20 to 40:40:20)
Sources & evidence
- ORD record
- ord-d4bd7784a1aa49eba9cb0e4046b89949
- Source dataset
- uspto-grants-2011_06
- Patent number
- US07956074B2
Reaction notes
Loading…