Iodoethane → 6-Amino-7-ethyl-2-[3-(2-fluorobenzyl)-1H-pyrazolo[4,3-b]pyridin-1-yl]-7,9-dihydro-8H-purin-8-one
Reaction equation

[F:1][C:2]1[CH:26]=[CH:25][CH:24]=[CH:23][C:3]=1[CH2:4][C:5]1[C:9]2=[N:10][CH:11]=[CH:12][CH:13]=[C:8]2[N:7]([C:14]2[N:19]=[C:18]([NH2:20])[C:17]([NH2:21])=[C:16]([NH2:22])[N:15]=2)[N:6]=1.I[CH2:28][CH3:29].[O:30]1CCC[CH2:31]1>C(OCC)(=O)C>[NH2:20][C:18]1[N:19]=[C:14]([N:7]2[C:8]3[C:9](=[N:10][CH:11]=[CH:12][CH:13]=3)[C:5]([CH2:4][C:3]3[CH:23]=[CH:24][CH:25]=[CH:26][C:2]=3[F:1])=[N:6]2)[N:15]=[C:16]2[C:17]=1[N:21]([CH2:28][CH3:29])[C:31](=[O:30])[NH:22]2Participants




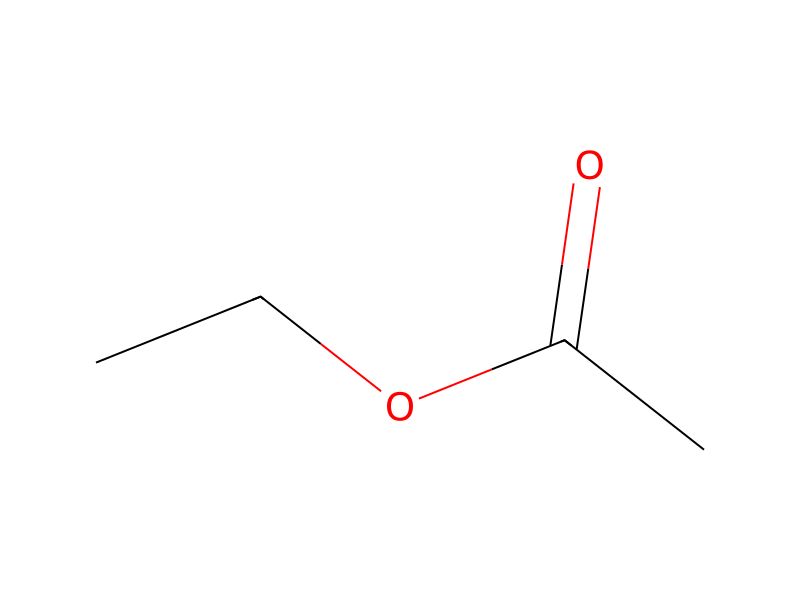
Conditions
- Temperature
- 0 °C
Procedure
200 mg (0.392 mmol) of 2-[3-(2-fluorobenzyl)-1H-pyrazolo[4,3-b]pyridin-1-yl]pyrimidine-4,5,6-triamine (example 2A) were initially charged in tetrahydrofuran (11.1 ml) and the suspension was cooled to 0° C. Subsequently, 0.43 ml (0.43 mmol) of bis(trimethylsilyl)sodium amide solution (1.0 M in tetrahydrofuran) was added and the mixture was stirred at 0° C. for a further 30 min Thereafter, 31.3 μl (0.392 mmol) of iodoethane were added dropwise and the reaction mixture was stirred at RT overnight. The reaction mixture was cooled again to 0° C. and a further 0.47 ml (0.47 mmol) of bis(trimethylsilyl)sodium amide solution was added. Thereafter, the reaction mixture was stirred at RT overnight and heated to reflux for a further 2 days. Subsequently, the reaction mixture was diluted with ethyl acetate and washed twice with saturated aqueous sodium hydrogencarbonate solution. The organic phase was dried over magnesium sulfate, filtered and concentrated by rotary evaporation. The residue was separated by means of preparative HPLC (eluent: acetonitrile-water with 0.1% formic acid gradient) and the product fractions were concentrated. The crude product was extracted by stirring in methyl tert-butyl ether, and the solids were filtered off and dried under high vacuum. This gave 4.6 mg (2.9% of theory) of the title compound in solid form.
Workup
- stirringthe reaction mixture was stirred at RT overnight
- temperatureThe reaction mixture was cooled again to 0° C.
- stirringThereafter, the reaction mixture was stirred at RT overnight
- temperatureheated
- temperatureto reflux for a further 2 days
- washwashed twice with saturated aqueous sodium hydrogencarbonate solution
- dry with materialThe organic phase was dried over magnesium sulfate
- filtrationfiltered
- concentrationconcentrated by rotary evaporation
- customThe residue was separated by means of preparative HPLC (eluent: acetonitrile-water with 0.1% formic acid gradient)
- concentrationthe product fractions were concentrated
- extractionThe crude product was extracted
- stirringby stirring in methyl tert-butyl ether
- filtrationthe solids were filtered off
- customdried under high vacuum
Sources & evidence
- ORD record
- ord-1a547e7b188f493ea8a6a50afdd5dcee
- Source dataset
- uspto-grants-2015_08
- Patent number
- US09096592B2
Reaction notes
Loading…