ホームMCPワークベンチSkills
ホームワークベンチ通知マイページ
llms.txt · MCPガイドプライバシーポリシー · 利用規約 · サポートv1.8.0
  1. ホーム
  2. /反応

Potassium Carbonate → 2-(2,4-Difluorophenyl)-N,N-dimethylpyridin-4-amine

HRID 1808271
ノート
印刷プレビュー

反応式

HRID 1808271 の反応式
I[C:2]1[CH:7]=[C:6]([N:8]([CH3:10])[CH3:9])[CH:5]=[CH:4][N:3]=1.[F:11][C:12]1[CH:17]=[C:16]([F:18])[CH:15]=[CH:14][C:13]=1B(O)O.C([O-])([O-])=O.[K+].[K+]>C1(C)C=CC=CC=1.O.C1C=CC([P]([Pd]([P](C2C=CC=CC=2)(C2C=CC=CC=2)C2C=CC=CC=2)([P](C2C=CC=CC=2)(C2C=CC=CC=2)C2C=CC=CC=2)[P](C2C=CC=CC=2)(C2C=CC=CC=2)C2C=CC=CC=2)(C2C=CC=CC=2)C2C=CC=CC=2)=CC=1>[F:11][C:12]1[CH:17]=[C:16]([F:18])[CH:15]=[CH:14][C:13]=1[C:2]1[CH:7]=[C:6]([N:8]([CH3:10])[CH3:9])[CH:5]=[CH:4][N:3]=1
参加物反応条件実験手順後処理出典と根拠反応ノート

参加物

7

反応物 3

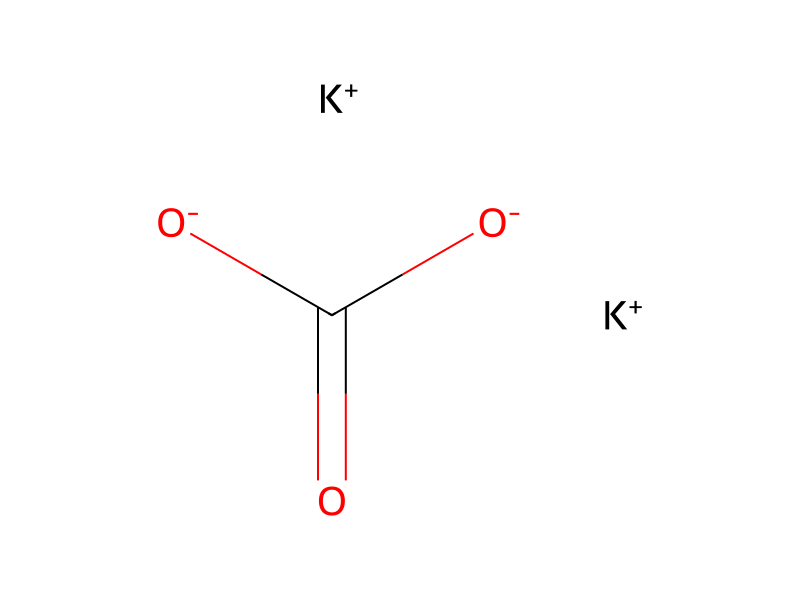
HCID 11430Potassium CarbonateHCID 27343342,4-Difluorophenylboronic acidHCID 110861002-Iodo-4-dimethylaminopyridine

生成物 1

HCID 208073522-(2,4-Difluorophenyl)-N,N-dimethylpyridin-4-amine収率 78.3%

試薬・触媒・溶媒 3

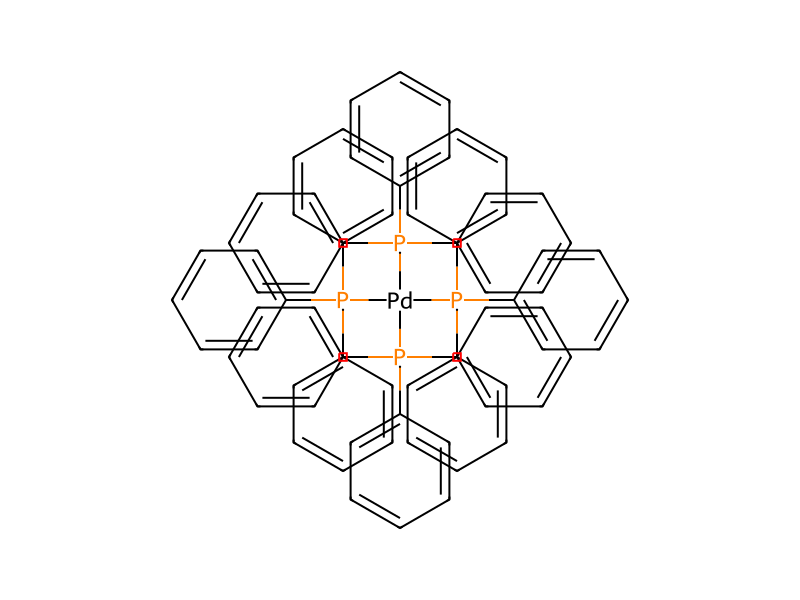
HCID 178418397HCID 178418397、化合物触媒
HCID 962
Water
溶媒
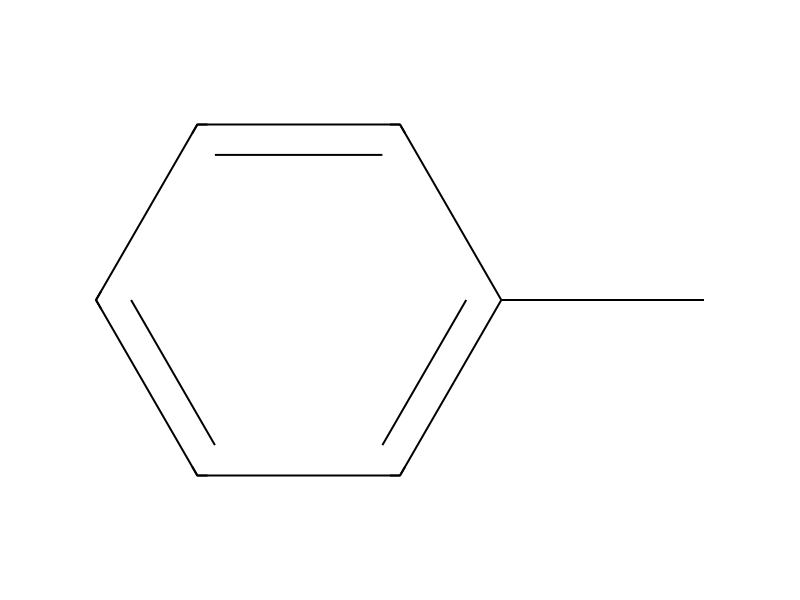
HCID 1140Toluene溶媒

反応条件

温度
90 °C

実験手順

A mixture of 2-iodo-4-dimethylaminopyridine (3 g, 12 mmol), 2,4-difluorophenylboronic acid (2.3 g, 14.5 mmol) and K2CO3 (6 g, 43.5 mmol) in toluene (60 ml) and water (10 ml) were degassed with nitrogen for 15 minutes. Pd(PPh3)4 (800 mg, 0.66 mmol) was added and the resulting mixture was heated to 90° C. for 48 hours under nitrogen. After being cooled to room temperature, the aqueous phase was separated and extracted with EtOAc (3×100 ml). The combined organic fractions were washed with brine, dried over MgSO4, filtered and evaporated. To further purify the compound, the so-obtained oil was dissolved in Et2O and extracted with 10% HCl solution (3×50 ml). The combined aqueous fractions were washed with Et2O (2×100 ml) and neutralized with concentrated NaOH aqueous solution. The resulting mixture was extracted with EtOAc (4×100 ml), the combined organic fractions were washed with brine (50 ml), dried over MgSO4, filtered and evaporated to dryness. The crude compound was purified by column chromatography (SiO2, CHCl3 then CHCl3/MeOH, 97/3) to afford 2.2 g (78%) of the titled compound as slightly yellow oil, which solidified upon standing.

後処理

  1. customwere degassed with nitrogen for 15 minutes
  2. temperatureAfter being cooled to room temperature
  3. customthe aqueous phase was separated
  4. extractionextracted with EtOAc (3×100 ml)
  5. washThe combined organic fractions were washed with brine
  6. dry with materialdried over MgSO4
  7. filtrationfiltered
  8. customevaporated
  9. customTo further purify the compound
  10. dissolutionthe so-obtained oil was dissolved in Et2O
  11. extractionextracted with 10% HCl solution (3×50 ml)
  12. washThe combined aqueous fractions were washed with Et2O (2×100 ml) and neutralized with concentrated NaOH aqueous solution
  13. extractionThe resulting mixture was extracted with EtOAc (4×100 ml)
  14. washthe combined organic fractions were washed with brine (50 ml)
  15. dry with materialdried over MgSO4
  16. filtrationfiltered
  17. customevaporated to dryness
  18. customThe crude compound was purified by column chromatography (SiO2, CHCl3

出典と根拠

ORDレコード
ord-9bf32757c9b8452db311499b706f3a6a
出典データセット
uspto-grants-2013_01
DOI
10.6084/m9.figshare.5104873.v1
特許番号
US08357799B2

反応ノート

読み込み中…

このページの内容参加物反応条件実験手順後処理出典と根拠反応ノート