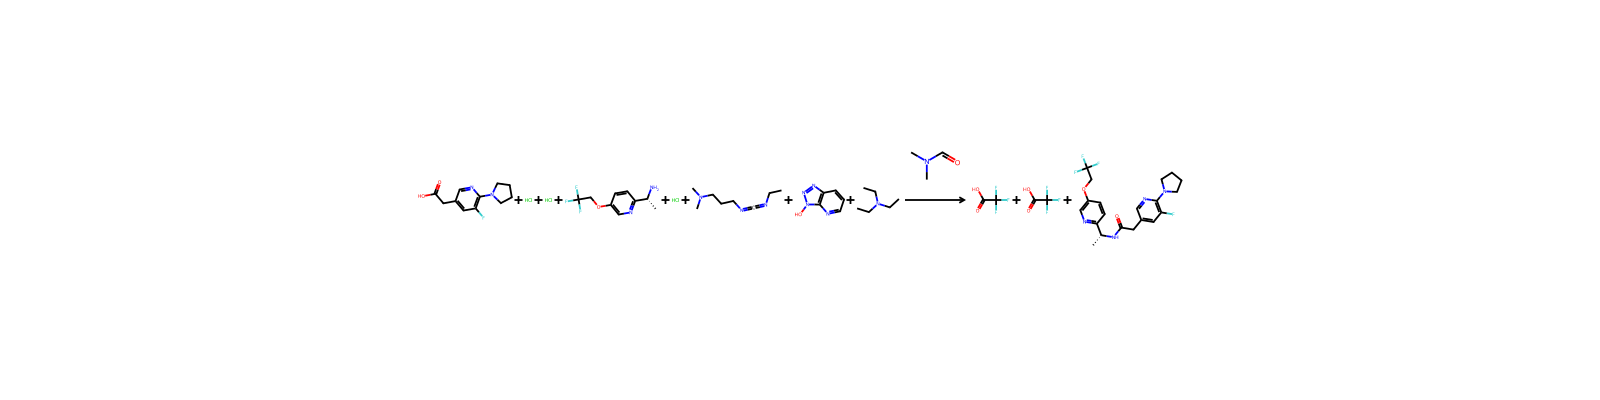
反応手順
A mixture of methyl 5-fluoro-6-methoxynicotinate (3.5 g, 18.9 mmol) and KOH (4.2 g, 61.5 mmol) in MeOH (70 ml) was stirred at room temperature for 5 h. After concentration, the residue was dissolved in water, and the obtained solution was washed with ether. The aqueous phase was acidified to pH=1 with dilute hydrochloric acid, and extracted with ether, and the combined organic layers were washed with brine, dried over Na2SO4 and concentrated under vacuum to afford 5-fluoro-6-methoxynicotinic acid (3.2 g, 98%) as a white solid. To a dry-ice cooled solution of 5-fluoro-6-methoxynicotinic acid (3.5 g, 20.5 mmol) and Et3N (2.3 g, 22.8 mmol) in dry THF (50 ml), was added isobutyl chloroformate (3.1 g, 22.7 mmol) slowly to keep the reaction temperature below −20° C. After the mixture was stirred for 1 h at the same temperature, to it ice-water was added, and the resulting mixture was extracted with ether, and the combined organic layers were washed with brine, dried over Na2SO4 and concentrated under vacuum to afford 5.5 g of (5-fluoro-6-methoxypyridin-3-yl)carbonyl isobutyl carbonate as a yellow oil (98%). 1H-NMR (400 MHz, CDCl3) δ 8.65 (d, J=2 Hz, 1H, Ar—H), 7.90 (dd, J1=10 Hz, J2=2 Hz, 1H, Ar—H), 4.11 (s, 3H, —OCH3), 3.73 (m, 2H, —OCH2), 2.09 (m, 1H, —CH), 0.98 (m, 6H, —CH3). A mixture of (5-fluoro-6-methoxypyridin-3-yl)carbonyl isobutyl carbonate (3.7 g, 13.6 mmol) and CH2N2-ether (30 mmol) was stirred for 5 h at −40° C. then overnight at room temperature. The reaction was quenched by the addition of water, and the resulting mixture was extracted with ether, and the combined organic layers were washed with brine, dried over Na2SO4 and concentrated under vacuum. The residue was purified by column to afford 3 g of 2-diazo-1-(5-fluoro-6-methoxypyridin-3-yl)ethanone (90%). 1H-NMR (400 MHz, CDCl3) δ 8.29 (d, J=2 Hz, 1H, Ar—H), 7.79 (dd, J1=10.4 Hz, J2=2 Hz, 1H, Ar—H), 5.85 (s, —CH), 4.09 (s, 3H, —OCH3). A mixture of 2-diazo-1-(5-fluoro-6-methoxypyridin-3-yl)ethanone (6 g, 30.8 mmol) in EtOH (100 ml) was heated to 40° C., then to it was added AgOAc (2 g, 12 mmol) and the resulting mixture was stirred at 70° C. for 2 h. After filtration, the filter cake was washed with THF. The filtrate was concentrated under vacuum, and the residue was purified by column to afford 2.5 g of ethyl (5-fluoro-6-methoxypyridin-3-yl)acetate as a solid (38.5%). 1H-NMR (400 MHz, CDCl3) δ 7.86 (d, J=1.6 Hz, 1H, Ar—H), 7.33 (dd, J=10 Hz, J2=2 Hz, 1H, Ar—H), 4.16 (q, J=7.8 Hz, 2H, —CH2CH3), 4.00 (s, 3H, —OCH3), 3.54 (s, 2H, Ar—CH2), 1.26 (t, J=7.8 Hz, 3H, —CH3). A mixture of ethyl (5-fluoro-6-methoxypyridin-3-yl)acetate (2.5 g, 11.7 mol), PCl5 (5 g, 24 mmol) and DMF (0.5 ml) in POCl3 (20 ml) was stirred overnight at 80° C. After concentration to remove the excess POCl3, the residue was poured into ice-water (100 ml), and the solution was neutralized to pH=7 with diluted aq. NaOH, and extracted with ether. The combined organic layers were washed with brine, dried over Na2SO4 and concentrated under vacuum, the residue was purified by column to afford 1.5 g of ethyl (6-chloro-5-fluoropyridin-3-yl)acetate as an oil (60%). 1H-NMR (400 MHz, CDCl3) δ 8.12 (d, J=1.6 Hz, 1H, Ar—H), 7.50 (dd, J1=8.4 Hz, J2=2 Hz, 1H, Ar—H), 4.19 (q, J=8.8 Hz, 2H, —CH2CH3), 3.63 (s, 3H, —OCH3), 1.27 (t, J=8.8 Hz, 3H, —CH3). A solution of ethyl (6-chloro-5-fluoropyridin-3-yl)acetate (1.5 g, 6.9 mmol) in conc. HCl (30 ml) was stirred at 120° C. for 5 h. After concentration, the residue was poured into water (50 ml), and the resulting mixture was extracted with ether. The combined organic layers was washed with brine, dried over Na2SO4 and concentrated under vacuum to afford 0.8 of (6-chloro-5-fluoropyridin-3-yl)acetic acid (61.2%). 1H-NMR (300 MHz, CDCl3) δ 8.15 (d, J=1.5 Hz, 1H, Ar—H), 7.51 (dd, J=8.4 Hz, J2=1.8 Hz, 1H, Ar—H), 3.70 (s, 3H, —OCH3). A mixture of (6-chloro-5-fluoropyridin-3-yl)acetic acid (2 g, 10.6 mmol) and pyrrolidine (2.2 g, 31.7 mmol) in dioxane (50 ml) was stirred at 110° C. overnight. After concentration, the residue was purified by prep. HPLC to afford 1.2 g (20%) of (5-fluoro-6-pyrrolidin-1-ylpyridin-3-yl)acetic acid TFA salt as a white solid. 1H-NMR (400 MHz CDCl3) δ 10.11 (br, 3H, COOH), 7.77 (s, 1H, Ar—H), 7.54 (d, J=13.2 Hz, 1H, Ar—H), 3.80 (m, 4H, CH2—CH2), 3.49 (m, Ar—CH2), 2.09 (m, 4H, CH2—CH2). To a 15 ml round bottom flask equipped with a magnetic stir bar were added (5-fluoro-6-pyrrolidin-1-ylpyridin-3-yl)acetic acid (45.9 mg, 0.21 mmol), (R)-1-[5-(2,2,2-trifluoroethoxy)pyridine-2-yl]ethylamine bis-hydrochloride salt (60.0 mg, 0.21 mmol), 1-(3-dimethylaminopropyl)-3-ethylcarbodiimide hydrochloride (47.1 mg, 0.25 mmol), 1-hydroxy-7-azabenzotriazole (33.4 mg, 0.25 mmol) and triethylamine (0.09 ml, 0.61 mmol) into 2.0 ml of dimethylformamide. The resulting mixture was stirred at room temperature for 16 hours. Reaction mixture was directly purified by preparative reversed-phase HPLC to give 2-(5-fluoro-6-pyrrolidin-1-ylpyridin-3-yl)-N-{(1R)-1-[5-(2,2,2-trifluoroethoxy)pyridin-2-yl]ethyl}acetamide bis-TFA salt as an oil (53.0 mg, 39.2%); 1H NMR (400 MHz, CD3OD) δ 8.42 (s, 1H), 7.79 (dd, J=2.8 Hz and 8.8 Hz, 1H), 7.73 (dd, J=1.6 Hz and 14 Hz, 1H), 7.62 (d, J=8.8 Hz, 1H), 7.29 (d, J=0.8 Hz, 1H), 5.04 (q, J=6.8 Hz, 1H), 4.71 (q, J=8.4 Hz, 2H), 3.74 (m, 4H), 3.56 (s, 2H), 2.11-2.05 (m, 4H), 1.50 (d, J=6.8 Hz, 3H); MS (Electrospray): m/z 427.1 (M+H).