ホームMCPワークベンチSkills
ホームワークベンチ通知マイページ
llms.txt · MCPガイドプライバシーポリシー · 利用規約 · サポートv1.8.0
  1. ホーム
  2. /反応

Hydrochloric Acid → 3-Fluoro-5-iodobenzenesulfonamide

HRID 720842
ノート
印刷プレビュー

反応式

HRID 720842 の反応式
[F:1][C:2]1[CH:3]=[C:4]([CH:6]=[C:7]([I:9])[CH:8]=1)N.Cl.[N:11]([O-])=O.[Na+].[S:15](=[O:17])=[O:16]>O.[Cu]Cl.C(O)(=O)C>[F:1][C:2]1[CH:3]=[C:4]([S:15]([NH2:11])(=[O:17])=[O:16])[CH:6]=[C:7]([I:9])[CH:8]=1
参加物反応条件実験手順後処理出典と根拠反応ノート

参加物

8

反応物 4

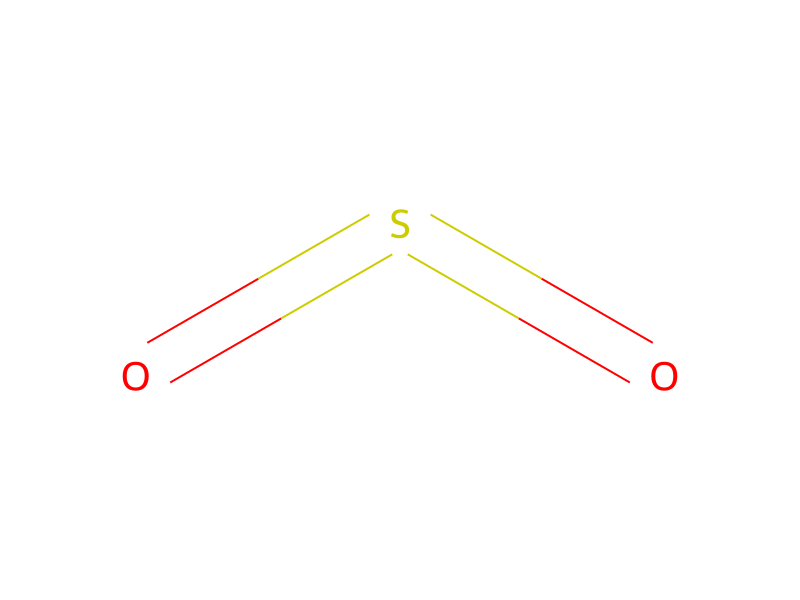
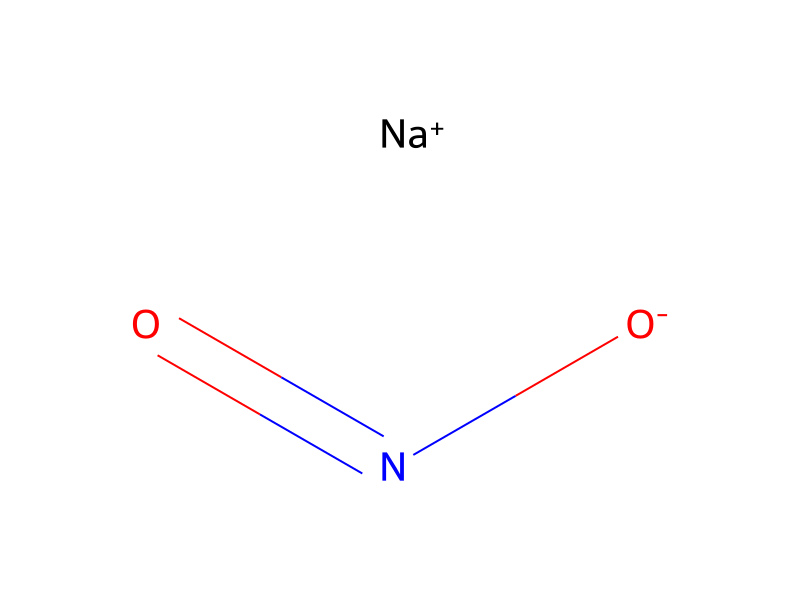
HCID 313Hydrochloric AcidHCID 1119Sulfur DioxideHCID 27589863-Fluoro-5-iodoanilineHCID 23668193Sodium Nitrite

生成物 1

HCID 218955263-Fluoro-5-iodobenzenesulfonamide

試薬・触媒・溶媒 3

HCID 62652Cuprous chloride触媒
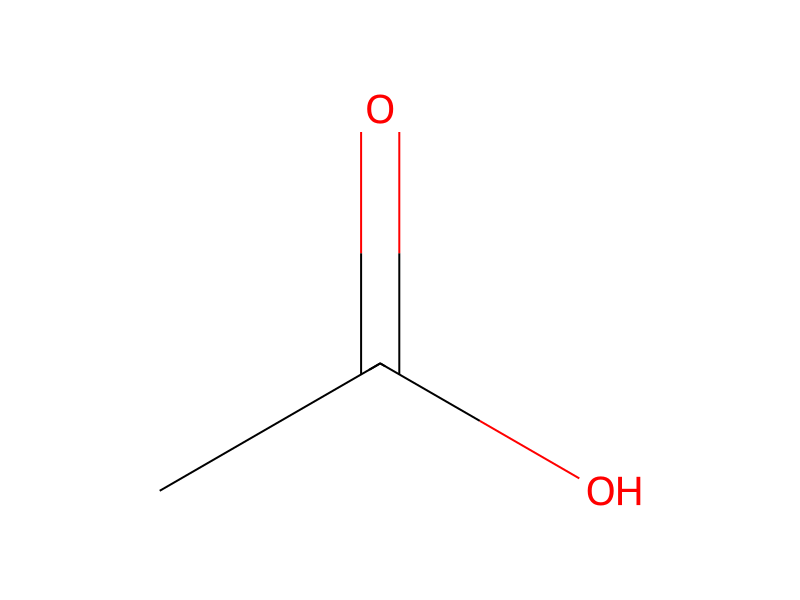
HCID 176
Acetic Acid
溶媒
2 回
HCID 962Water溶媒2 回

反応条件

温度
-5 °C

実験手順

3-Fluoro-5-iodoaniline (3.06 g) (WO 9623783) was added to a stirred mixture of concentrated hydrochloric acid (4 ml) and water (4 ml). Glacial acetic acid (8 ml) was added and the reaction mixture cooled to −5° C. A solution of sodium nitrite (0.99 g) in water (8 ml) was added dropwise maintaining the temperature between −5° C. and −2° C. After the addition was complete the reaction was stirred for 10 min. In the meantime glacial acetic acid (20 ml) was saturated with sulfur dioxide gas for 0.25 h, then copper(I) chloride (0.353 g) was added. Additional sulfur dioxide was bubbled through the solution until a fine suspension was obtained. The mixture was cooled to 5° C. and then treated portionwise with the diazonium salt prepared above. After stirring at room temperature for 1 h ice (50 g) was added. The mixture was extracted with ether (100 ml) and the organic phases washed with NaHCO3 solution (2×100 ml) then water (100 ml), dried (MgSO4) and concentrated. The residue was dissolved in THF (30 ml) at 0° C. and aqueous ammonia (0.880; 5 ml) was added. After stirring at room temperature the mixture was partitioned between EtOAc (100 ml) and water (100 ml). The organic phase was washed with brine (50 ml), dried (MgSO4) and concentrated. The residue was purified by chromatography using cyclohexane-EtOAc (5:1 then 3:1). Evaporation of the fractions and trituration of the residue with cyclohexane afforded the title compound (0.886 g). ES−ve 299 (M−H)−

後処理

  1. temperaturemaintaining the temperature between −5° C. and −2° C
  2. additionAfter the addition
  3. customAdditional sulfur dioxide was bubbled through the solution until a fine suspension
  4. customwas obtained
  5. temperatureThe mixture was cooled to 5° C.
  6. customprepared above
  7. additionwas added
  8. extractionThe mixture was extracted with ether (100 ml)
  9. washthe organic phases washed with NaHCO3 solution (2×100 ml)
  10. dry with materialwater (100 ml), dried (MgSO4)
  11. concentrationconcentrated
  12. dissolutionThe residue was dissolved in THF (30 ml) at 0° C.
  13. additionaqueous ammonia (0.880; 5 ml) was added
  14. stirringAfter stirring at room temperature the mixture
  15. customwas partitioned between EtOAc (100 ml) and water (100 ml)
  16. washThe organic phase was washed with brine (50 ml)
  17. dry with materialdried (MgSO4)
  18. concentrationconcentrated
  19. customThe residue was purified by chromatography
  20. customEvaporation of the fractions and trituration of the residue with cyclohexane

出典と根拠

ORDレコード
ord-a4b44d43f82f45dbabdf04819ff32c1d
出典データセット
uspto-grants-2006_11
DOI
10.6084/m9.figshare.5104873.v1
特許番号
US07135600B2

反応ノート

読み込み中…

このページの内容参加物反応条件実験手順後処理出典と根拠反応ノート