Acetonitrile, 2,2-diethoxy- → Methyl 2,2-diethoxyethanecarboximidate
Reaction equation

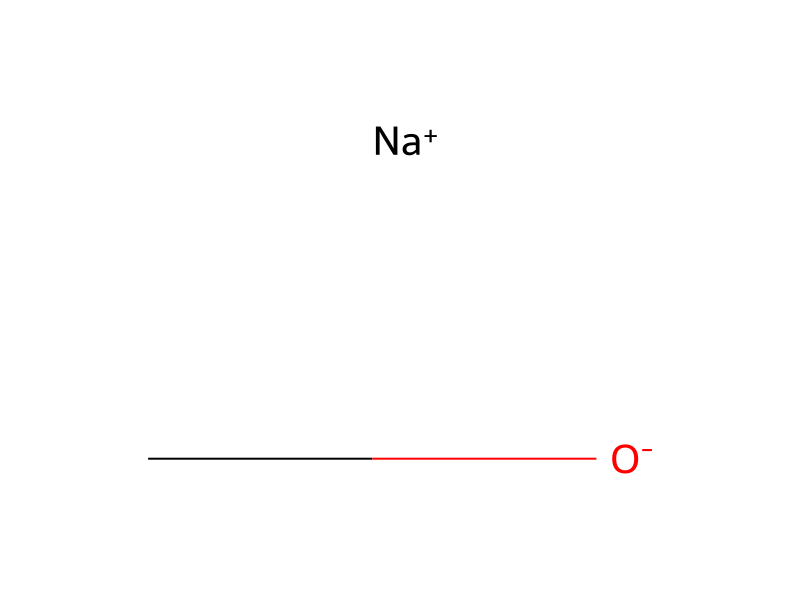
[CH2:1]([O:3][CH:4]([O:7][CH2:8][CH3:9])[C:5]#[N:6])[CH3:2].[CH3:10][O-:11].[Na+]>CO>[CH2:1]([O:3][CH:4]([O:7][CH2:8][CH3:9])[C:5](=[NH:6])[O:11][CH3:10])[CH3:2]Participants
Reactant 2

Product 1

Reagents, catalysts & solvents 1
Procedure
Cyclopropanecarbonitrile (100 g, 1.49 mol), methanol (59.7 g, 76 mL, 1.86 mol), and diethyl ether (184 mL) were charged to a 2 L three-necked flask. The mixture was cooled to −10° C. while stirring and HCl (g) (232 g, 6.36 mol) was then bubbled through the mixture solution such that the internal temperature was maintained below −5° C. Upon completion of gas addition, the mixture was stirred for 1.5 h at −10° C. Ether (685 mL) was added slowly over a 1-h period while continuing to maintain a the temperature below 0° C. The resulting solids were filtered and rinsed with diethyl ether (3×300 mL). The solids were further dried under vacuum, affording 181.5 g (89.8% yield) of methyl cyclopropanecarboximidoate hydrochloride. mp: 112-114° C. In a 2 L three-neck flask, a solution of diethoxyacetonitrile (125 g, 968 mmol) in methanol (300 mL, anhydrous) was treated with a 25 wt % solution of sodium methoxide (21.0 g, 389 mmol) in methanol, producing a slightly exothermic reaction. After cooling, the mixture was stirred at room temperature for 15 h. Upon completion of the reaction, the solvent was slowly evaporated under vacuum. The remaining brown-colored residue was solvated with diethyl ether (1 L), then washed with water (3×500 mL), followed by brine (1×200 mL). The organic layer was dried using magnesium sulfate, the salts filtered, and the solvent evaporated carefully under vacuum to afford 132 g (85% yield) of methyl 2,2-diethoxyethanimidoate as a colorless liquid. Methyl 2,2-diethoxyethanimidoate (39.0 g, 242 mmol) was dissolved in tetrahydrofuran (345 mL), mixed with methylhydrazine (11.1 g, 242 mmol), then the mixture stirred at room temperature for 5 h, at which point LC/MS analysis confirmed the formation of the amidrazone intermediate. Methyl cyclopropanecarboximidoate hydrochloride (32.8 g, 242 mmol) was added to the reaction mixture, resulting in a suspension. Acetic acid (21.8 g, 363 mmol) was added slowly to the suspension, and the resulting exothermic reaction controlled with a cooling bath. After addition was completed, the reaction mixture was stirred at ambient temperature overnight. After LC/MS analysis indicated that the starting reactants had been consumed, the mixture was diluted with dichloromethane (500 mL) and stirred for 1 h. The solids were filtered and rinsed with dichloromethane (3×200 mL), and the filtrate concentrated in vacuo. The remaining oil was purified by column chromatography on silica gel (mobile phase: ethyl acetate, then with 1, 3, 5, 10% methanol, consecutively). The product-containing fractions were pooled and the solvent evaporated in vacuo. After further drying under high vacuum, 27.4 g of the title acetal was obtained as an oil containing 12.4 weight % of ethyl acetate as an impurity.
Workup
- customproducing
- customa slightly exothermic reaction
- temperatureAfter cooling
- customUpon completion of the reaction
- customthe solvent was slowly evaporated under vacuum
- washwashed with water (3×500 mL)
- customThe organic layer was dried
- filtrationthe salts filtered
- customthe solvent evaporated carefully under vacuum
Sources & evidence
- ORD record
- ord-f1b0553c51464f25a18a1f76d000f99a
- Source dataset
- uspto-grants-2009_11
- Patent number
- US07622605B2
Reaction notes
Loading…