Triethylamine → (2S)-N-((1S,2S)-3-{(2S,4S)-2-[N-(tert-butyl)carbamoyl]-4-chloropyrrolidinyl}-2-hydroxy-3-oxo-1-benzylpropyl)-3-carbamoyl-2-[(phenylmethoxy)carbonylamino]propanamide
Reaction equation

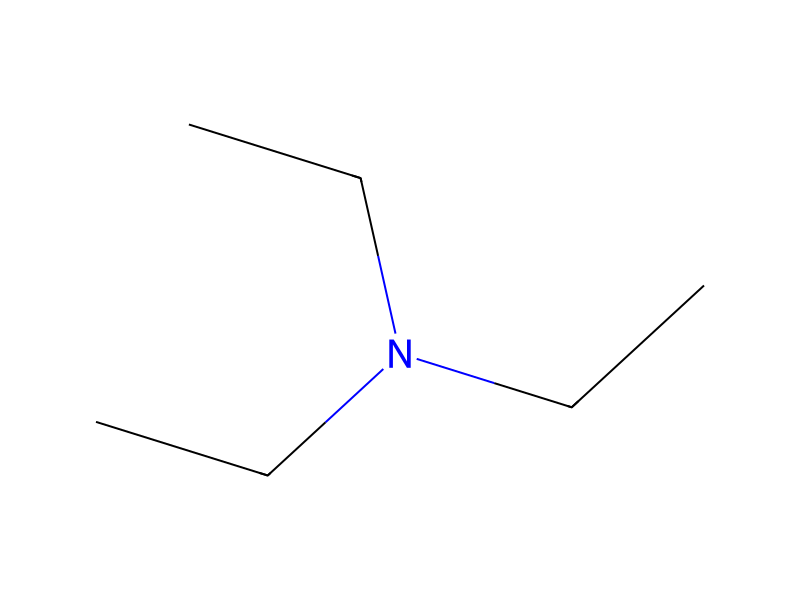
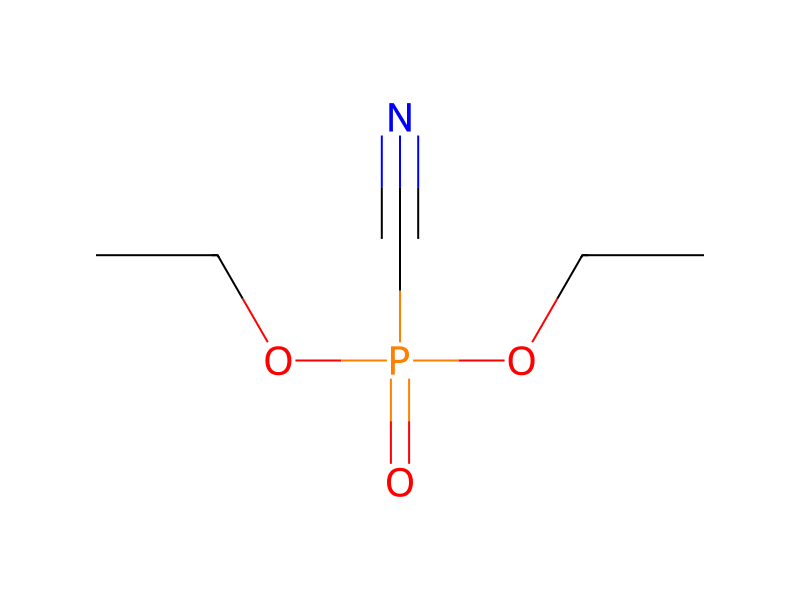
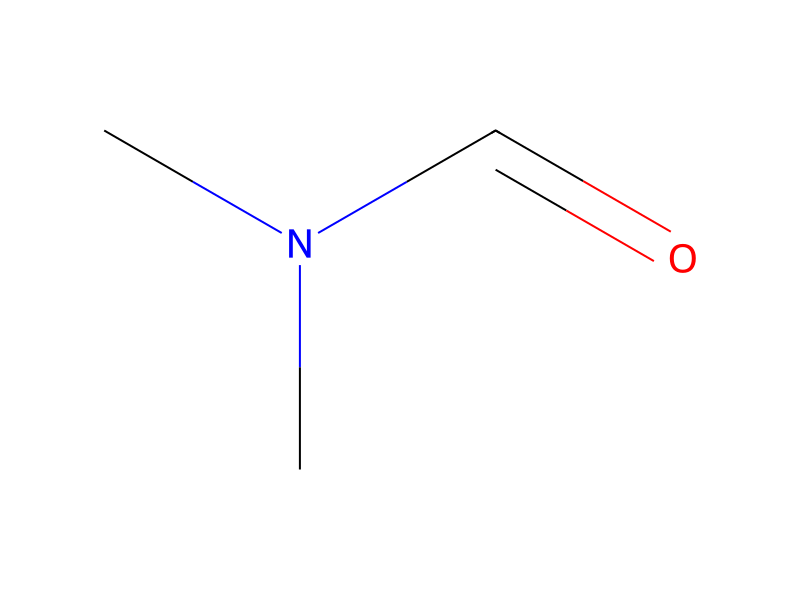
Cl.[C:2]([NH:6][C:7](=[O:14])[C@@H:8]1[CH2:12][C@H:11]([Cl:13])[CH2:10][NH:9]1)([CH3:5])([CH3:4])[CH3:3].[CH2:15]([O:22][C:23]([NH:25][C@H:26]([C:31]([NH:33][C@@H:34]([CH2:40][C:41]1[CH:46]=[CH:45][CH:44]=[CH:43][CH:42]=1)[C@H:35]([OH:39])[C:36](O)=[O:37])=[O:32])[CH2:27][C:28](=[O:30])[NH2:29])=[O:24])[C:16]1[CH:21]=[CH:20][CH:19]=[CH:18][CH:17]=1.P(C#N)(OCC)(OCC)=O.C(N(CC)CC)C>CN(C)C=O>[CH2:15]([O:22][C:23]([NH:25][C@H:26]([C:31]([NH:33][C@@H:34]([CH2:40][C:41]1[CH:42]=[CH:43][CH:44]=[CH:45][CH:46]=1)[C@H:35]([OH:39])[C:36]([N:9]1[CH2:10][C@@H:11]([Cl:13])[CH2:12][C@H:8]1[C:7]([NH:6][C:2]([CH3:5])([CH3:3])[CH3:4])=[O:14])=[O:37])=[O:32])[CH2:27][C:28](=[O:30])[NH2:29])=[O:24])[C:16]1[CH:21]=[CH:20][CH:19]=[CH:18][CH:17]=1Participants
Reactant 4


Product 1

Reagents, catalysts & solvents 1
Procedure
680 mg (2.23 mmol) of (4S)-1-t-butoxycarbonyl-N-t-butyl-4-chloro-L-prolinamide (prepared as described in Preparation 2) were treated with 10 ml of a 4N solution of hydrogen chloride in dioxane, in order to eliminate the t-butoxycarbonyl group and give (4S)-N-t-butyl-4-chloro-L-prolinamide hydrochloride. The whole of the hydrochloride thus obtained and 880 mg (1.99 mmol) of (2S,3S)-3-(N2 -benzyloxycarbonyl-L-asparaginyl)amino-2-hydroxy-4-phenylbutyric acid (prepared as described in Preparation 1) were dissolved in 5 ml of dimethylformamide, and the solution was cooled with ice. 420 mg (2.58 mmol) of diethyl cyanophosphate, followed by 640 mg (6.40 mmol) of triethylamine, were then added to the ice-cooled mixture, and the mixture was stirred for 3 hours. At the end of this time, the reaction mixture was concentrated by evaporation under reduced pressure, and the residue was mixed with a 10% w/v aqueous solution of citric acid and then extracted with ethyl acetate. The organic extract was washed with a 10% w/v aqueous solution of sodium hydrogencarbonate and with a saturated aqueous solution of sodium chloride, in that order, after which it was dried over anhydrous sodium sulfate. The solvent was then removed by distillation under reduced pressure, and the residue was purified by preparative thin layer chromatography using a 10:1 by volume mixture of methylene chloride and methanol as the developing solvent, to give 850 mg of the title compound as a colorless powder, melting at 107°-109° C.
Workup
- temperaturethe solution was cooled with ice
- temperaturecooled mixture
- concentrationAt the end of this time, the reaction mixture was concentrated by evaporation under reduced pressure
- additionthe residue was mixed with a 10% w/v aqueous solution of citric acid
- extractionextracted with ethyl acetate
- extractionThe organic extract
- washwas washed with a 10% w/v aqueous solution of sodium hydrogencarbonate and with a saturated aqueous solution of sodium chloride, in that order
- dry with materialafter which it was dried over anhydrous sodium sulfate
- customThe solvent was then removed by distillation under reduced pressure
- customthe residue was purified by preparative thin layer chromatography
- additionby volume mixture of methylene chloride and methanol as the developing solvent
Sources & evidence
- ORD record
- ord-f0b75d9522a04845a90692a068cb670e
- Source dataset
- uspto-grants-1997_05
- Patent number
- US05629406
Reaction notes
Loading…