HCID 86759966, compound → HCID 22220914, compound
Reaction equation

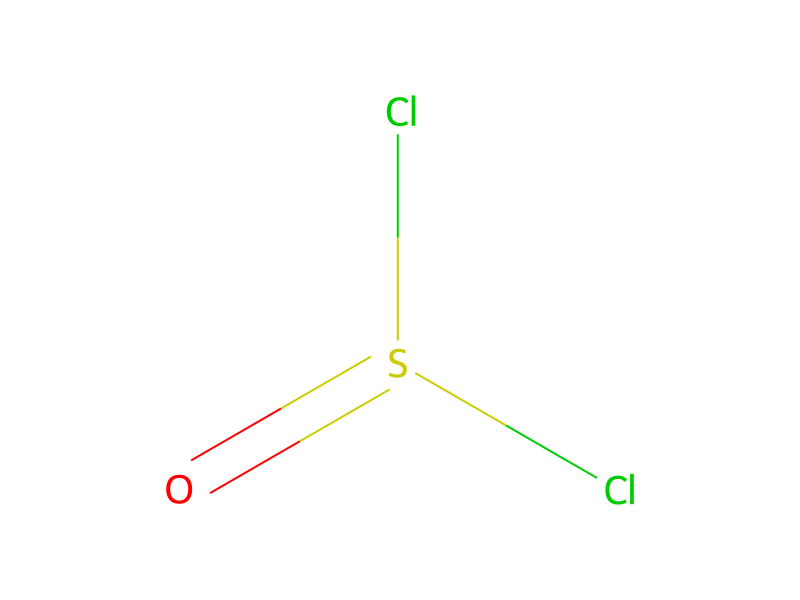
[C:1]1([C:7]2[C:11]([C:12]([NH:14][NH:15][S:16]([C:19]3[CH:24]=[CH:23][C:22]([CH3:25])=[CH:21][CH:20]=3)(=[O:18])=[O:17])=[O:13])=[C:10]([CH3:26])[O:9][N:8]=2)[CH:6]=[CH:5][CH:4]=[CH:3][CH:2]=1.CC1C=CC(S(C2C(C3C=CC=CC3=NN)N(CCl)OC=2C)(=O)=O)=CC=1>S(Cl)(Cl)=O>[C:1]1([C:7]2[C:11]([C:12]([NH:14][NH:15][S:16]([C:19]3[CH:20]=[CH:21][C:22]([CH3:25])=[CH:23][CH:24]=3)(=[O:18])=[O:17])=[O:13])=[C:10]([CH3:26])[O:9][N:8]=2)[CH:6]=[CH:5][CH:4]=[CH:3][CH:2]=1Participants
Reactant 1

Product 1

Reagents, catalysts & solvents 1
Procedure
3-Phenyl-5-methyl-4-isoxazoyl hydrazide (40 g, 0.184 mol) (prepared as in Example 1) was dissolved in anhydrous pyridine (300 ml) and cooled to 0° C. p-Toluenesulfonyl chloride (35.15 g, 0.184 mol) was dissolved in anhydrous pyridine (200 ml) and added dropwise to the cooled hydrazide solution. The reaction was stirred for an additional 15 minutes at 0° C., warmed to 25° C., and the pyridine removed in vacuo. The residue was poured into 30% HCl and the product extracted into EtOAc. The organic layer was dried over MgSO4 and evaporated. The residue was recrystallised from 70% EtOH to obtain the title compound as a beige solid (65 g, 95%). 1H NMR (360 MHz, CDCl3) δ 7.75 (2H, d, J=2.2 Hz), 7.57–7.48 (5H, m), 7.29 (2H, d, J=2.2 Hz), 2.49 (3H, s), 2.43 (3H, s). b) 4-(4-Methylphenylsulfonyl)hydrazono(chloro)methyl-5-methyl-3-phenylisoxazole 1-(3-phenyl-5-methyl-4-isoxazoloyl)-2-p-toluenesulfonylhydrazine (65 g, 0.175 mol) was placed in a flask containing thionyl chloride (300 ml) and the reaction heated to reflux under nitrogen for 2 h. The reaction mixture was cooled, the solvent removed in vacuo, and ethanol added to the residue. The resulting solid was filtered, washed with ethanol and dried in vacuo at 25° C. to obtain the title compound as a white solid (60 g, 88%). 1H NMR (400 MHz, CDCl3) δ 8.25 (1H, s), 7.76–7.73 (2H, m), 7.46–7.29 (7H, m), 2.45 (3H, s), 2.42 (3H, s). c) 6-Nitro-2,3-dihydro-isoindol-1-one To a solution of 2,3-dihydroisoindole-1-one (5 g, 38 mmol) in concentrated sulfuric acid (100 ml) at 0° C., KNO3 was added portionwise whilst maintaining the temperature at 0° C. The reaction was gradually allowed to warm to room temperature and was stirred overnight. The reaction mixture was poured into ice and the resulting precipitate was filtered off and dried to give title compound as a yellow solid (4.7 g, 70%). 1H NMR (360 MHz, d6 DMSO) δ 8.96 (1H, s), 8.45 (1H, dd, J=8.3 Hz, 2.2 Hz), 8.34 (1H, d, J=2 Hz), 7.87 (1H, d, J=8.3 Hz), 4.54 (2H, s). m/z (ES+) 179 (M+). d) 6-Amino-2,3-dihydro-isoindol-1-one 6-Nitro-2,3-dihydroisoindol-1-one (4.7 g, 26 mmol) was dissolved in acetic acid (130 ml) and hydrogenated at 50 psi (3 bar) in the presence of palladium on charcoal (0.47 g) overnight. The catalyst was filtered off through a pad of Hyflo™ and the acetic acid was removed in vacuo. The residue was triturated with dichloromethane/hexane and the resulting solid was filtered off to afford the title compound (3.56 g, 91%). 1H NMR (360 MHz, d6 DMSO) δ 8.28 (1H, s), 7.17 (1H, d, J=8 Hz), 6.82 (1H, s), 6.78 (1H, dd, J=8.1, 1.8 Hz), 5.25 (2H, s), 4.16 (2H, s). m/z (ES+) 149 (M+H)+. e) (3-Oxo-2,3-dihydro-1H-isoindol-5-yl)carbamic acid tert-butyl ester To a solution of 6-amino-2,3-dihydroisoindol-1-one (1.78 g, 12 mmol) in DMF/H2O (2:1, 75 ml) was added K2CO3 (3.32 g, 24 mmol, 2 eq.) and di-tert-butyl dicarbonate (5.67 g, 26 mmol, 2.2 eq.). The reaction was stirred at room temperature overnight. The reaction was partitioned between ethyl acetate and water, and the organic layer was washed with saturated NaCl (aq), dried (MgSO4), filtered and concentrated in vacuo. The residue was triturated with ethyl acetate and the resulting solid was filtered off to afford title compound (2.04 g, 68%). 1H NMR (360 MHz, d6 DMSO) δ 8.48 (1H, s), 7.83 (1H, d, J=1.2 Hz), 7.6 (1H, dd, J=8.2, 1.9 Hz), 7.43 (1H, d, J=8.2 Hz), 4.28 (2H, s), 1.49 (9H, s). f) 5-tert-Butoxycarbonylamino-3-methoxy-1H-isoindolium trifluoromethane sulfonate Under nitrogen atmosphere, (3-oxo-2,3-dihydro-1H-isoindol-5-yl)-carbamic acid tert-butyl ester (2.14 g, 8.23 mmol) was dissolved in dichloromethane (190 ml) and methyl triflate (1.40 ml, 12 mmol, 1.5 eq.) was added. The reaction was heated to reflux for 3 hours and then stirred at room temperature overnight. The reaction mixture was evaporated to one fifth of the original volume followed by the addition of diethyl ether. The resulting white precipitate was filtered off to afford the title compound title compound (2.71 g, 80%). 1H NMR (360 MHz, d6 DMSO) δ 9.87 (1H, s), 8.11 (1H, s), 7.75 (1H, d,J=1.9 Hz), 7.70 (1H, d, J=8.4 Hz), 4.93 (2H, s), 4.35 (3H, s), 1.50 (9H, s). g) 3-(5-Methyl-3-phenylisoxazol-4-yl)-5H-[1,2,4]triazolo[3,4-a]isoindol-8-ylamine To a solution of 5-tert-butoxycarbonylamino-3-methoxy-1H-isoindolium trifluoromethane sulfonate (0.53 g, 1.29 mmol) in 1,4-dioxane was added N-p-toluenesulfonyl)-5-methyl-3-phenylisoxazol-4-hydrazidoyl chloride 4-(4-methylphenylsulfonyl)hydrazono(chloro)methyl-5-methyl-3-phenylisoxazole (0.5 g, 1.29 mmol) and triethylamine (0.18 ml, 1.29 mmol). The reaction was heated to reflux for one hour. The solvent was concentrated in vacuo and the residue was purified by silica chromatography, cluting with 2% methanol/dichloromethane. The resulting solid was dissolved in trifluoroacetic acid (5 ml) and was stirred at room temperature for 10 minutes. The reaction was concentrated in vacuo and the residue was partitioned between ethyl acetate and saturated K2CO3 (aq). The organic layer was washed with saturated NaCl (aq), dried (MgSO4), filtered and concentrated in vacuo. The residue was purified by silica chromatography, eluting with ethyl acetate→3% methanol/ethyl acetate, to afford the title compound title compound (97 mg, 23%). 1H NMR (360 MHz, d6 DMSO) δ 7.53–7.44 (5H, m), 7.17 (1H, d, J=8.4 Hz), 7.14 (1H, s), 6.74 (1H, d, J=7.9 Hz), 4.33 (2H, s), 2.59 (3H, s). m/z (ES+) 330 (M+H)+.
Workup
- temperaturethe reaction heated
- temperatureto reflux under nitrogen for 2 h
- temperatureThe reaction mixture was cooled
- customthe solvent removed in vacuo, and ethanol
- additionadded to the residue
- filtrationThe resulting solid was filtered
- washwashed with ethanol
- customdried in vacuo at 25° C.
Sources & evidence
- ORD record
- ord-f8451293a9c1492ba8927366b5af2d1a
- Source dataset
- uspto-grants-2006_04
- Patent number
- US07022724B2
Reaction notes
Loading…