Dichloromethane → (2R)-2-({4-[(3,4-Dihydro-2H-[1,4]oxathiepino[2,3-c]pyridin-8-ylmethyl)amino]-1-piperidinyl}methyl)-1,2-dihydro-3H,8H-2a,5,8a-triazaacenaphthylene-3,8-dione hydrochloride
Reaction equation

[NH2:1][CH:2]1[CH2:7][CH2:6][N:5]([CH2:8][C@H:9]2[N:19]3[C:20]4[N:11]([C:12](=[O:22])[CH:13]=[CH:14][C:15]=4[N:16]=[CH:17][C:18]3=[O:21])[CH2:10]2)[CH2:4][CH2:3]1.[S:23]1[C:33]2[CH:32]=[C:31]([CH:34]=O)[N:30]=[CH:29][C:28]=2[O:27][CH2:26][CH2:25][CH2:24]1.S1C2C=C(C=O)N=CC=2OCC1.BrC(Br)(C)C.C(O[BH-](OC(=O)C)OC(=O)C)(=O)C.[Na+].C(=O)([O-])O.[Na+].[Cl:72]CCl>CO.C(Cl)(Cl)Cl>[ClH:72].[S:23]1[C:33]2[CH:32]=[C:31]([CH2:34][NH:1][CH:2]3[CH2:7][CH2:6][N:5]([CH2:8][C@H:9]4[N:19]5[C:20]6[N:11]([C:12](=[O:22])[CH:13]=[CH:14][C:15]=6[N:16]=[CH:17][C:18]5=[O:21])[CH2:10]4)[CH2:4][CH2:3]3)[N:30]=[CH:29][C:28]=2[O:27][CH2:26][CH2:25][CH2:24]1Participants
Reactant 7
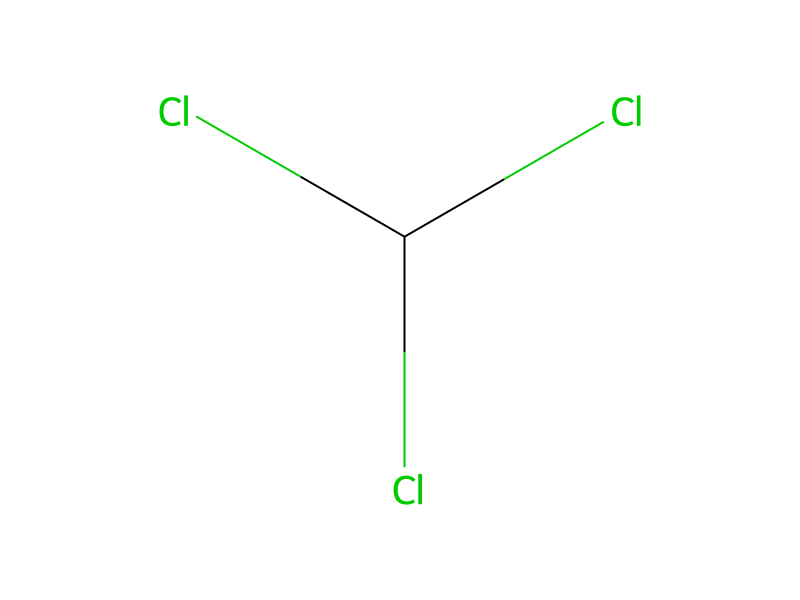
 HCID 6344Dichloromethane
 HCID 116582,2-Dibromopropane
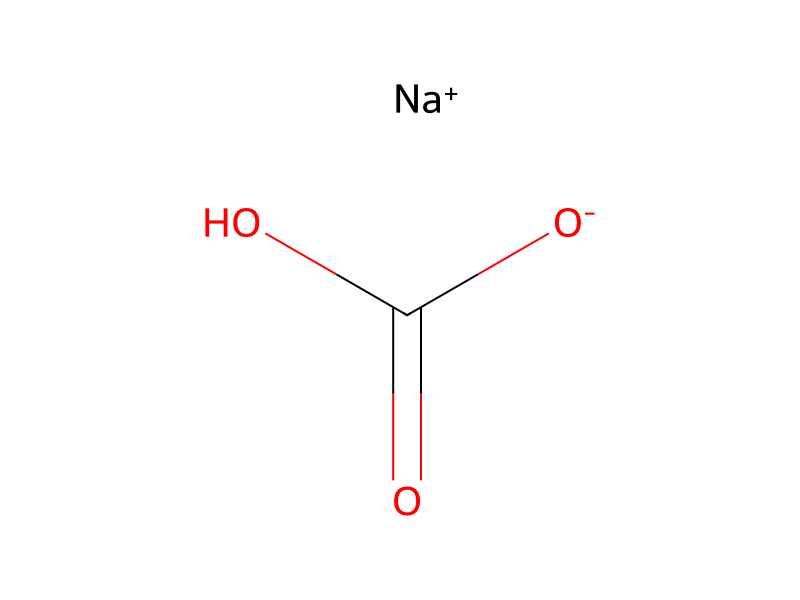 HCID 516892Sodium Bicarbonate
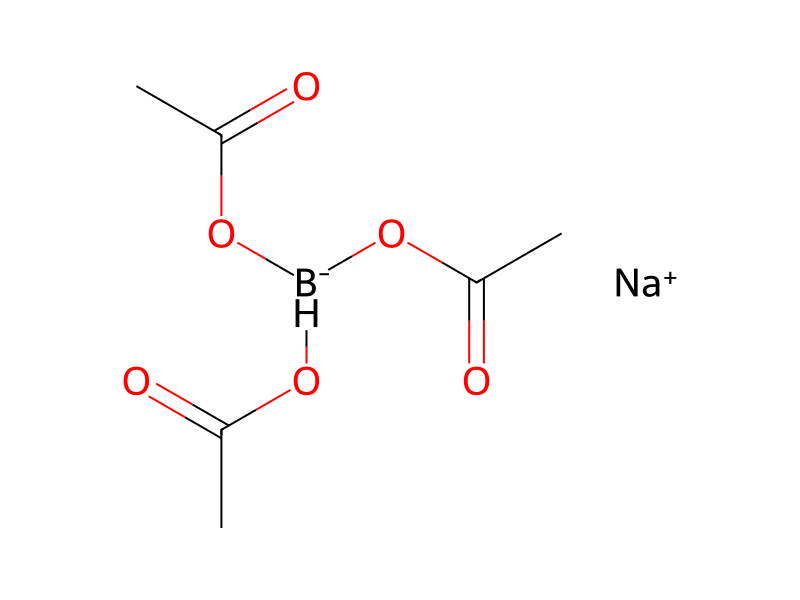 HCID 5049666Sodium triacetoxyborohydride
 HCID 574231152,3-Dihydro[1,4]oxathiino[2,3-c]pyridine-7-carbaldehyde
 HCID 58200877(2R)-2-[(4-Amino-1-piperidinyl)methyl]-1,2-dihydro-3H,8H-2a,5,8a-triazaacenaphthylene-3,8-dione
 HCID 866421063,4-dihydro-2H-[1,4]oxathiepino[2,3-c]pyridine-8-carbaldehyde
Product 1

Reagents, catalysts & solvents 2
Procedure
(2R)-2-[(4-Amino-1-piperidinyl)methyl]-1,2-dihydro-3H,8H-2a,5,8a-triazaacenaphthylene-3,8-dione (50 mg, 0.166 mmol) (for a preparation see Example 16(j)) and 3,4-dihydro-2H-[1,4]oxathiepino[2,3-c]pyridine-8-carbaldehyde (29 mg, 0.895 eq.) (may be prepared analogously to the synthesis of 2,3-dihydro[1,4]oxathiino[2,3-c]pyridine-7-carbaldehyde (WO2004058144, Example 60) but replacing dibromoethane with dibromopropane) were stirred in 9:1 v:v chloroform:methanol (1 ml) for 2.5 hours. Sodium triacetoxyborohydride (105 mg, 3.000 eq.) was then added in one portion and the mixture was stirred vigorously at room temperature for 30 minutes. Saturated aqueous sodium hydrogen carbonate (0.5 ml) was then added, followed by dichloromethane (10 ml) and the mixture was stirred vigorously at room temperature for 10 min and the phases were separated (hydrophobic frit). The organic phase was evaporated under reduced pressure and the crude product was purified by column chromatography on silica (eluted with 0-12% (2M NH3 in MeOH) in DCM). Appropriate fractions were combined and evaporated under reduced pressure to give the free base of the title compound as a pale yellow solid (65 mg).
Workup
- stirringthe mixture was stirred vigorously at room temperature for 30 minutes
- stirringthe mixture was stirred vigorously at room temperature for 10 min
- customthe phases were separated (hydrophobic frit)
- customThe organic phase was evaporated under reduced pressure
- customthe crude product was purified by column chromatography on silica (eluted with 0-12% (2M NH3 in MeOH) in DCM)
- customevaporated under reduced pressure
Sources & evidence
- ORD record
- ord-007ec7ceda7a4de2a6e15d444a20e033
- Source dataset
- uspto-grants-2013_03
- Patent number
- US08389524B2
Reaction notes
Loading…