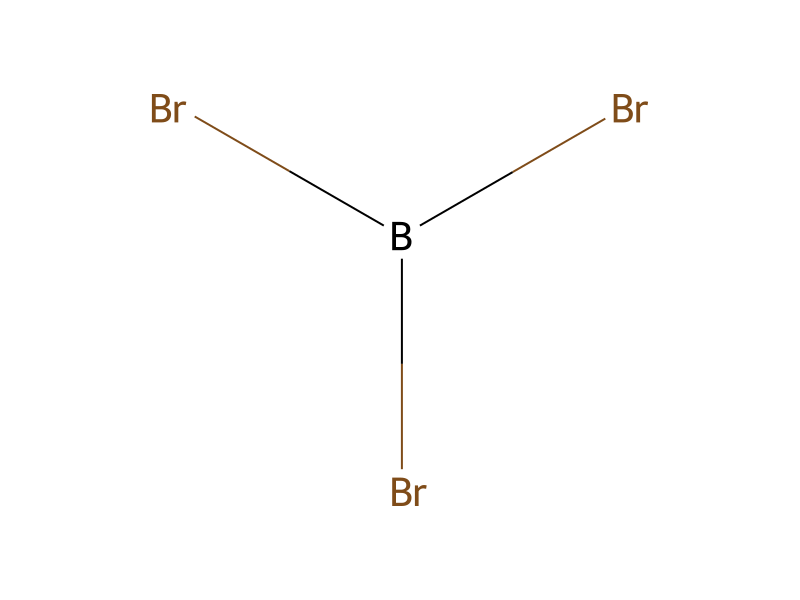
Boron tribromide → 4-((5-Bromo-2-chlorophenyl)methyl)phenol
Reaction equation

[Br:1][C:2]1[CH:7]=[CH:6][C:5]([Cl:8])=[C:4]([CH2:9][C:10]2[CH:15]=[CH:14][C:13]([O:16]CC)=[CH:12][CH:11]=2)[CH:3]=1.B(Br)(Br)Br>ClCCl>[Br:1][C:2]1[CH:7]=[CH:6][C:5]([Cl:8])=[C:4]([CH:3]=1)[CH2:9][C:10]1[CH:15]=[CH:14][C:13]([OH:16])=[CH:12][CH:11]=1Participants
Reactant 2

Product 1

Reagents, catalysts & solvents 1

Procedure
To a solution of 4-bromo-1-chloro-2-(4-ethoxy-benzyl)-benzene (5.0 g, 15.35 mmol) in dichloromethane (20.0 mL) cooled to 0 degrees Celsius under nitrogen was added drop-wise over 30 minutes a 1M solution of boron tribromide in dichloromethane (17.0 mL, 17.0 mmol). After the addition was complete, the reaction was allowed to warm to room temperature overnight (˜16 hours). The reaction was cooled to 0 degrees Celsius and quenched with an aqueous solution of 1N hydrochloric acid. The resulting mixture was stirred for 30 minutes and extracted using dichloromethane. The organic layer was separated and the aqueous layer was extracted two times with dichloromethane. The combined organic layers were dried over magnesium sulfate, filtered and concentrated under reduced pressure. The reaction was purified by flash chromatography over silica gel using the ISCO automated chromatography unit (120 g silica gel column) and eluting with a gradient of 0 to 100% ethyl acetate in heptane. 3.53 g of desired product obtained (77% yield). 1H NMR (400 MHz, METHANOL-d4) delta ppm 3.94 (s, 2H), 6.70 (d, J=8.6 Hz, 2H), 6.98 (d, J=8.4 Hz, 2H), 7.23-7.34 (m, 3H). Alternative Procedure: To a solution of 4-bromo-1-chloro-2-(4-ethoxy-benzyl)-benzene (10.0 g, 30.71 mmol) in dichloromethane (40.0 mL) cooled to 0 degrees Celsius under nitrogen was added drop-wise over 30 minutes a 1M solution of boron trichloride in dichloromethane (34 mL, 34.0 mmol). After the addition was complete, the reaction was allowed to warm to room temperature overnight (˜16 hours). The reaction mixture was cooled to 0 degrees Celsius and an aqueous solution of 1N hydrochloric acid was added. The resulting mixture was stirred for 30 minutes and then extracted using dichloromethane. The organic layer was separated and the aqueous layer was extracted two times with dichloromethane. The combined organic layers were dried over magnesium sulfate, filtered and concentrated under reduced pressure. The TLC showed only 50% conversion. The crude material was redissolved in dichloromethane (40 mL), cooled to 0 degrees Celsius, and a 1M solution of boron tribromide in dichloromethane (31 mL, 31 mmol) was added dropwise. The reaction mixture was allowed to warm to room temperature over the weekend (˜55 hours). The reaction mixture was cooled to 0 degrees Celsius and an aqueous solution of 1N hydrochloric acid was added dropwise. The resulting mixture was stirred for 30 minutes and then extracted using dichloromethane. The organic layer was separated and the aqueous layer was extracted two times with dichloromethane. The combined organic layers were dried over magnesium sulfate, filtered and concentrated under reduced pressure. The reaction was purified by flash chromatography over silica gel using the ISCO automated chromatography unit (120 g silica gel column) and eluting with a gradient of 0 to 100% ethyl acetate in heptane. 9 g (98% yield) of the desired product obtained as a white solid.
Workup
- additionAfter the addition
- temperatureThe reaction was cooled to 0 degrees Celsius
- customquenched with an aqueous solution of 1N hydrochloric acid
- extractionextracted
- customThe organic layer was separated
- extractionthe aqueous layer was extracted two times with dichloromethane
- dry with materialThe combined organic layers were dried over magnesium sulfate
- filtrationfiltered
- concentrationconcentrated under reduced pressure
- customThe reaction was purified by flash chromatography over silica gel using the ISCO automated chromatography unit (120 g silica gel column)
- washeluting with a gradient of 0 to 100% ethyl acetate in heptane
Sources & evidence
- ORD record
- ord-bc86afa720fc4ffbafeeefd4f308ce5b
- Source dataset
- uspto-grants-2016_04
- Patent number
- US09308204B2
Reaction notes
Loading…