Reaktionsgleichung

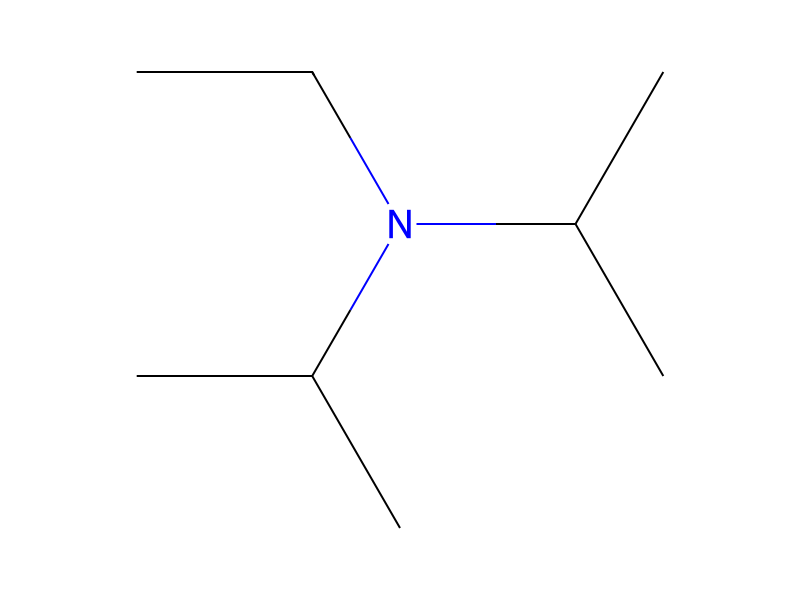
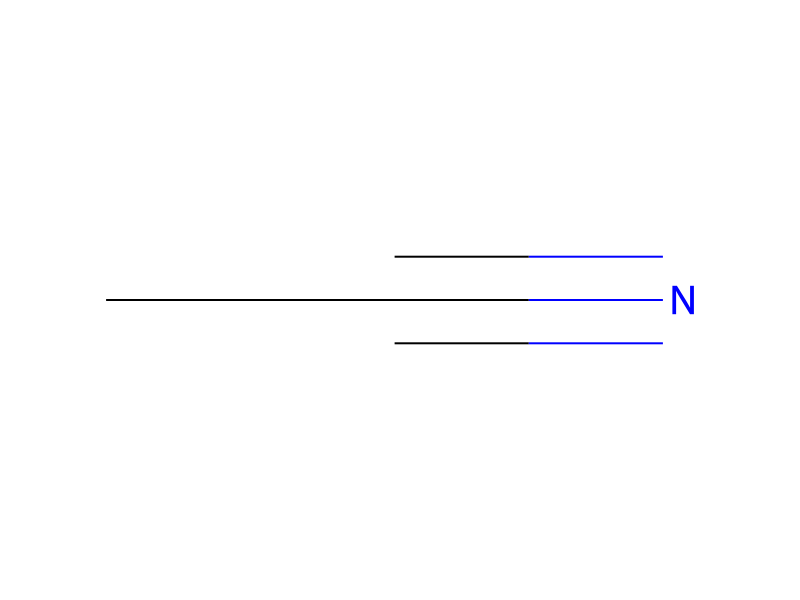
Cl.Cl.[N:3]1[CH:8]=[CH:7][CH:6]=[CH:5][C:4]=1[N:9]1[CH2:14][CH2:13][CH:12]([NH2:15])[CH2:11][CH2:10]1.[C:16](N1C=CN=C1)(N1C=CN=C1)=[O:17].C(N(CC)C(C)C)(C)C.[NH2:37][CH2:38][CH2:39][NH:40][C:41](=[O:47])[O:42][C:43]([CH3:46])([CH3:45])[CH3:44]>C(#N)C>[N:3]1[CH:8]=[CH:7][CH:6]=[CH:5][C:4]=1[N:9]1[CH2:10][CH2:11][CH:12]([NH:15][C:16]([NH:37][CH2:38][CH2:39][NH:40][C:41](=[O:47])[O:42][C:43]([CH3:44])([CH3:46])[CH3:45])=[O:17])[CH2:13][CH2:14]1Komponenten
Edukt 4



Produkt 1

Reagenzien, Katalysatoren, Lösungsmittel 1
Versuchsvorschrift
an ice-cooled solution of 1-(2-pyridinyl)-4-piperidinylamine dihydrochloride EP-A-0021973) (20.82 g, 0.0832 moles) and 1,1′-carbonyldiimidazole (14.85 g, 0.915 moles) in acetonitrile (140 ml) under an atmosphere of nitrogen was added N,N-di-iso-propylethylamine (22.0 g, 29.7 ml, 0.170 moles) over a period of 20 minutes. The resultant light brown solution was stirred at ambient temperature for 20 minutes and a solution of tert-butyl N-(2-aminoethyl)carbamate (14.0 g, 0.0874 moles) in acetonitrile (10 ml plus 5 ml rinse) was added over a period of 5 minutes. The resultant mixture was then heated under reflux for 2.5 hours. After cooling to ambient temperature, the reaction mixture was concentrated in vacuo and the residue was dissolved in ethyl acetate (150 ml). This solution was washed successively with saturated aqueous sodium bicarbonate solution (70 ml) and deionised water (20 ml). The aqueous phases were extracted with ethyl acetate (2×100 ml) and the combined organic phases were washed with water. The organic phase was dried over anhydrous magnesium sulphate and the solution was concentrated to a volume of approximately 200 ml in vacuo. The resultant solution was then distilled at atmospheric pressure until the volume was approximately 75 ml. The solution was allowed to cool to ambient temperature during which time crystallisation occurred. The resultant thick slurry was diluted with ethyl acetate (60 ml) and was cooled in ice. The solid was collected by filtration and the filter cake was washed with cold ethyl acetate (2×30 ml). The resultant solid was dried in vacuo at 50° C. for 20 hours to give the crude product (24.0 g) that was recrystallised from ethyl acetate (270 ml) to give the title compound (20.7 g) as a colourless solid.
Aufarbeitung
- temperatureunder reflux for 2.5 hours
- temperatureAfter cooling to ambient temperature
- concentrationthe reaction mixture was concentrated in vacuo
- dissolutionthe residue was dissolved in ethyl acetate (150 ml)
- washThis solution was washed successively with saturated aqueous sodium bicarbonate solution (70 ml) and deionised water (20 ml)
- extractionThe aqueous phases were extracted with ethyl acetate (2×100 ml)
- washthe combined organic phases were washed with water
- dry with materialThe organic phase was dried over anhydrous magnesium sulphate
- concentrationthe solution was concentrated to a volume of approximately 200 ml in vacuo
- distillationThe resultant solution was then distilled at atmospheric pressure until the volume
- temperatureto cool to ambient temperature during which time crystallisation
- additionThe resultant thick slurry was diluted with ethyl acetate (60 ml)
- temperaturewas cooled in ice
- filtrationThe solid was collected by filtration
- washthe filter cake was washed with cold ethyl acetate (2×30 ml)
- customThe resultant solid was dried in vacuo at 50° C. for 20 hours
- customto give the crude product (24.0 g) that
- customwas recrystallised from ethyl acetate (270 ml)
Quellen und Nachweise
- ORD-Eintrag
- ord-9de8626dfbfe4853a1c297e1bbb92197
- Quelldatensatz
- uspto-grants-2006_08
- Patentnummer
- US07094769B2
Verknüpfte Notizen
Wird geladen…