Reaction equation

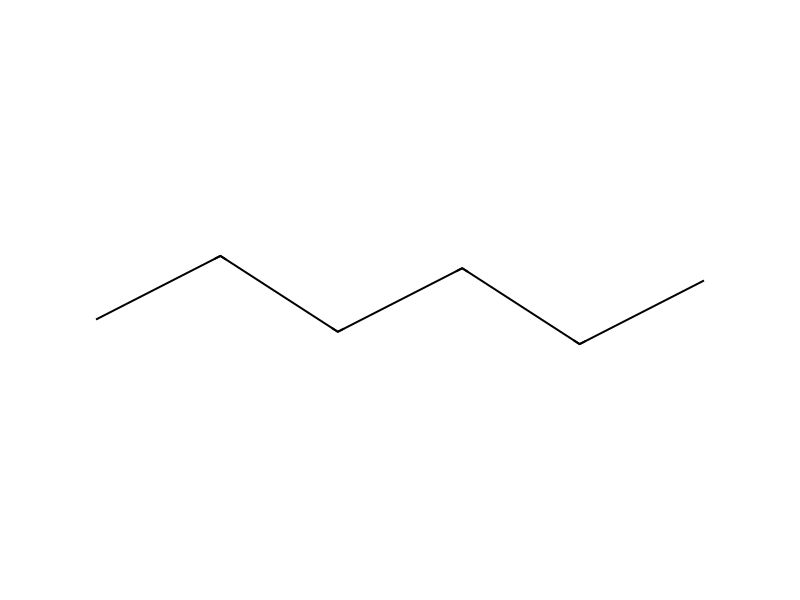
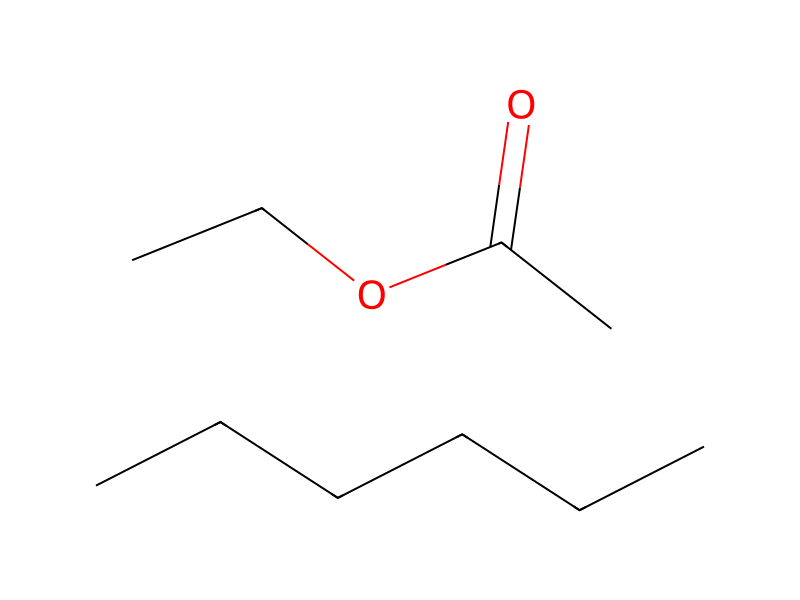
[O:1]=[C:2]1[C:8]2[CH:9]=[CH:10][CH:11]=[CH:12][C:7]=2[CH2:6][O:5][C:4]2[CH:13]=[CH:14][C:15]([CH2:17][C:18](O)=[O:19])=[CH:16][C:3]1=2.O1CCCC1.B.CO>O.CCCCCC.CCCCCC.C(OCC)(=O)C>[O:1]=[C:2]1[C:8]2[CH:9]=[CH:10][CH:11]=[CH:12][C:7]=2[CH2:6][O:5][C:4]2[CH:13]=[CH:14][C:15]([CH2:17][CH2:18][OH:19])=[CH:16][C:3]1=2Participants


Conditions
- Temperature
- 5 °C
Procedure
A stirred solution (-17° C.) of 6.71 g (0.025 mol) of 6,11-dihydro-11-oxodibenz[b,e]oxepin-2-acetic acid and 18 ml of anhydrous tetrahydrofuran was treated dropwise over 15 minutes with 24 ml of 1.06M borane in tetrahydrofuran solution (approximately 0.025 mol BH3). After total addition, the cooling bath was allowed to equilibrate overnight (about 16 hours) to ambient temperature. The resultant stirred, chilled (5° C.) solution was treated with 5 ml of methanol, diluted with 50 ml of water and concentrated on a rotary evaporator at 40° C. to remove the organic solvents. The residue was diluted with water (70 ml) and CH2Cl2 (70 ml) and was washed with 5% NaHCO3 solution until the aqueous phase had a pH of 8. The organic phase was dried (Na2SO4) and then filtered. The resultant organic phase was concentrated to an oil (5.51 g) which was dried by azeotropic distillation of toluene (70 ml). A portion of the oil was dissolved in 8 ml of anhydrous ethyl acetate, diluted to the cloud point with hexane (7 ml) and was then treated dropwise with ethyl acetate (0.6 ml) until a clear solution was obtained. The solution was subjected to high pressure liquid chromatography with hexane/ethyl acetate (60:40). The product-containing fractions were combined and concentrated to an oil (2.08 g). On standing at 5° C., some crystals formed and were stirred into the oil. On trituration of the mixture with hexane the oil solidified. The material was isolated by vacuum filtration, washed with hexane and dried in vacuo at ambient temperature to afford 1.42 g (22.3%) of a solid of 2-(6,11-dihydro-11-oxodibenz[b,e]oxepin-2-yl)ethanol m.p. 78°-81.5° C.
Workup
- additionAfter total addition
- custom(about 16 hours)
- customto ambient temperature
- concentrationconcentrated on a rotary evaporator at 40° C.
- customto remove the organic solvents
- additionThe residue was diluted with water (70 ml) and CH2Cl2 (70 ml)
- washwas washed with 5% NaHCO3 solution until the aqueous phase
- dry with materialThe organic phase was dried (Na2SO4)
- filtrationfiltered
- concentrationThe resultant organic phase was concentrated to an oil (5.51 g) which
- dry with materialwas dried by azeotropic distillation of toluene (70 ml)
- dissolutionA portion of the oil was dissolved in 8 ml of anhydrous ethyl acetate
- additiondiluted to the cloud point with hexane (7 ml)
- additionwas then treated dropwise with ethyl acetate (0.6 ml) until a clear solution
- customwas obtained
- concentrationconcentrated to an oil (2.08 g)
- customOn standing at 5° C.
- customsome crystals formed
- stirringwere stirred into the oil
- customThe material was isolated by vacuum filtration
- washwashed with hexane
- customdried in vacuo at ambient temperature
Sources & evidence
- ORD record
- ord-2a9cd24603aa4e868f50bd97afeb9135
- Source dataset
- uspto-grants-1985_05
- Patent number
- US04515946
Linked notes
Loading…