ホームMCPワークベンチSkills
ホームワークベンチ通知マイページ
llms.txt · MCPガイドプライバシーポリシー · 利用規約 · サポートv1.8.3
  1. ホーム
  2. /反応

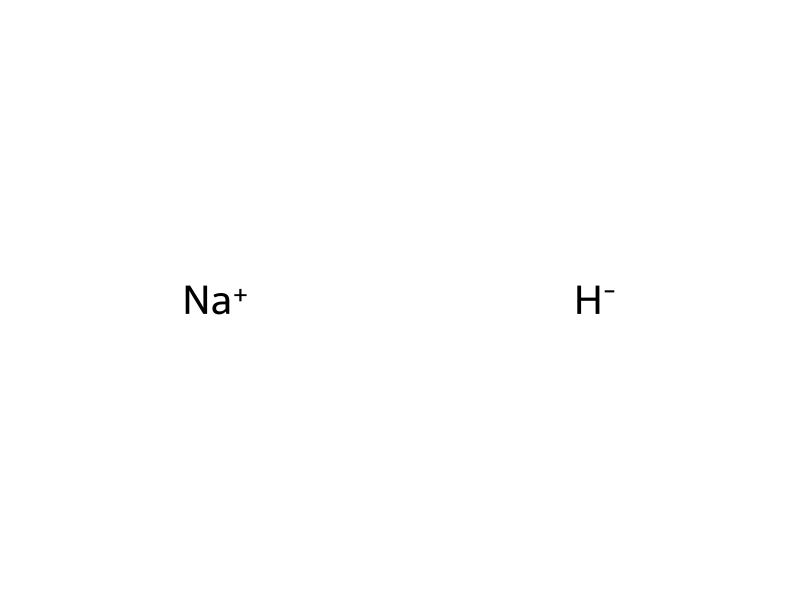
Sodium hydride → 5,11-Dihydro-11-[6-(1-piperidinyl)hexyl]-6H-pyrido[2,3-b][1,4]benzodiazepin-6-one

HRID 1084713
印刷

反応式

HRID 1084713 の反応式
C1(C[N:8]2[C:14](=[O:15])[C:13]3[CH:16]=[CH:17][CH:18]=[CH:19][C:12]=3[NH:11][C:10]3[N:20]=[CH:21][CH:22]=[CH:23][C:9]2=3)C=CC=CC=1.[H-].[Na+].Br[CH2:27][CH2:28][CH2:29][CH2:30][CH2:31][CH2:32][N:33]1[CH2:38][CH2:37][CH2:36][CH2:35][CH2:34]1>CN(C)C=O>[N:33]1([CH2:32][CH2:31][CH2:30][CH2:29][CH2:28][CH2:27][N:11]2[C:12]3[CH:19]=[CH:18][CH:17]=[CH:16][C:13]=3[C:14](=[O:15])[NH:8][C:9]3[CH:23]=[CH:22][CH:21]=[N:20][C:10]2=3)[CH2:38][CH2:37][CH2:36][CH2:35][CH2:34]1
参加物反応条件実験手順後処理出典と根拠反応ノート

参加物

5

反応物 3

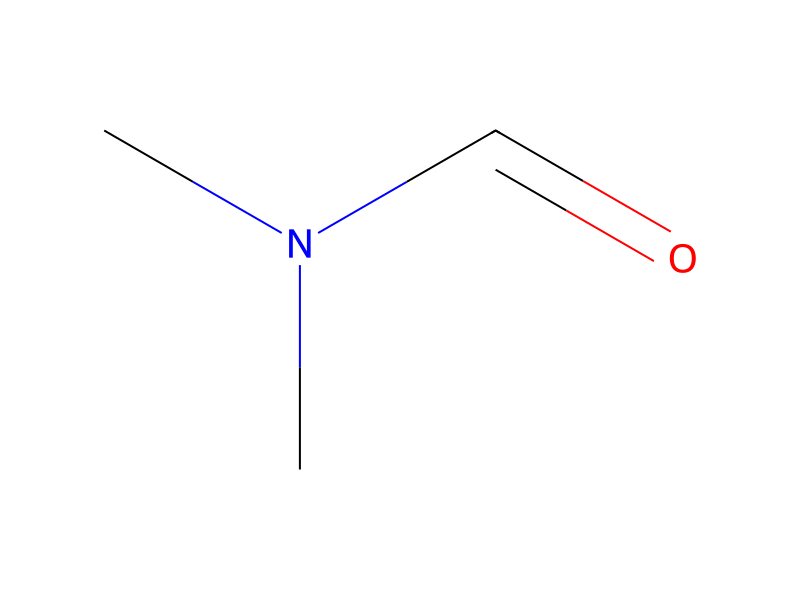
HCID 24758Sodium hydrideHCID 200624805,11-dihydro-5-(phenylmethyl)-6H-pyrido[2,3-b][1,4]benzodiazepin-6-oneHCID 200624841-Bromo6-(1-piperidinyl)hexane

生成物 1

HCID 145520925,11-Dihydro-11-[6-(1-piperidinyl)hexyl]-6H-pyrido[2,3-b][1,4]benzodiazepin-6-one

試薬・触媒・溶媒 1

HCID 6228Dimethylformamide溶媒

反応条件

温度
60 °C

実験手順

24.1 g (0.08 mol) of 5,11-dihydro-5-(phenylmethyl)-6H-pyrido[2,3-b][1,4]benzodiazepin-6-one (m.p. 152°-154° C.) were dissolved in 180 ml of anhydrous dimethylformamide, then 2.88 g (0.096 mol) of an 80% sodium hydride dispersion in mineral oil were added at ambient temperature and the resulting mixture was stirred for 45 minutes at 60° C. Then 23.8 g (0.096 mol) of 1-bromo-6-(1-piperidinyl)hexane were added dropwise and the mixture was heated to 120° C. for 30 minutes. After evaporation in vacuo, the residue was partitioned between dichloromethane and 5% aqueous hydrochloric acid, the aqueous phase was separated off and made alkaline with sodium hydroxide. The oil separated was taken up in dichloromethane, the solvent was distilled off in a water jet vacuum, the oily, highly viscous residue was carefully stirred with 250 g of 100% orthophosphoric acid and 7.5 g of phenol and the resulting mixture was heated to 120° C. for 3 hours. The mixture cooled to 70° C. was stirred into 2 kg of crushed ice, then made significantly alkaline by the addition of caustic soda and extracted exhaustively with dichloromethane. The dichloromethane extracts were dried over sodium sulphate and evaporated down, the residue remaining was digested with 10 ml of boiling ether and filtered while hot. The ether was evaporated off, the residue was purified by chromatography on silica gel (Macherey-Nagel, 0.2-0.5 mm) using dichloromethane/methanol/cyclohexane/ethyl acetate/conc. ammonia 59/7.5/7.5/25/1 (v/v/v/v/v) as eluant. By evaporating the appropriate fractions and subsequent recrystallisation from acetonitrile, colourless crystals with a melting point of 131°-132° C. were obtained in a yield of 21.8 g (72% of theory) which were found, according to their mixed melting point, thin layer chromatogram, IR, UV., 1H-NMR spectra, to be totally identical to a preparation prepared according to Example 6.

後処理

  1. temperaturethe mixture was heated to 120° C. for 30 minutes
  2. customAfter evaporation in vacuo
  3. customthe residue was partitioned between dichloromethane and 5% aqueous hydrochloric acid
  4. customthe aqueous phase was separated off
  5. customThe oil separated
  6. distillationthe solvent was distilled off in a water jet vacuum
  7. stirringthe oily, highly viscous residue was carefully stirred with 250 g of 100% orthophosphoric acid and 7.5 g of phenol
  8. temperaturethe resulting mixture was heated to 120° C. for 3 hours
  9. temperatureThe mixture cooled to 70° C.
  10. stirringwas stirred into 2 kg of crushed ice
  11. additionmade significantly alkaline by the addition of caustic soda
  12. extractionextracted exhaustively with dichloromethane
  13. dry with materialThe dichloromethane extracts were dried over sodium sulphate
  14. customevaporated down
  15. filtrationfiltered while hot
  16. customThe ether was evaporated off
  17. customthe residue was purified by chromatography on silica gel (Macherey-Nagel, 0.2-0.5 mm)
  18. customBy evaporating
  19. customthe appropriate fractions and subsequent recrystallisation from acetonitrile, colourless crystals with a melting point of 131°-132° C.
  20. customwere obtained in a yield of 21.8 g (72% of theory) which
  21. additionmixed melting point, thin layer chromatogram, IR, UV
  22. customprepared

出典と根拠

ORDレコード
ord-795c94f6f52045abbbec281450d82ab0
出典データセット
uspto-grants-1991_04
DOI
10.6084/m9.figshare.5104873.v1
特許番号
US05006522

反応ノート

このページの内容参加物反応条件実験手順後処理出典と根拠反応ノート