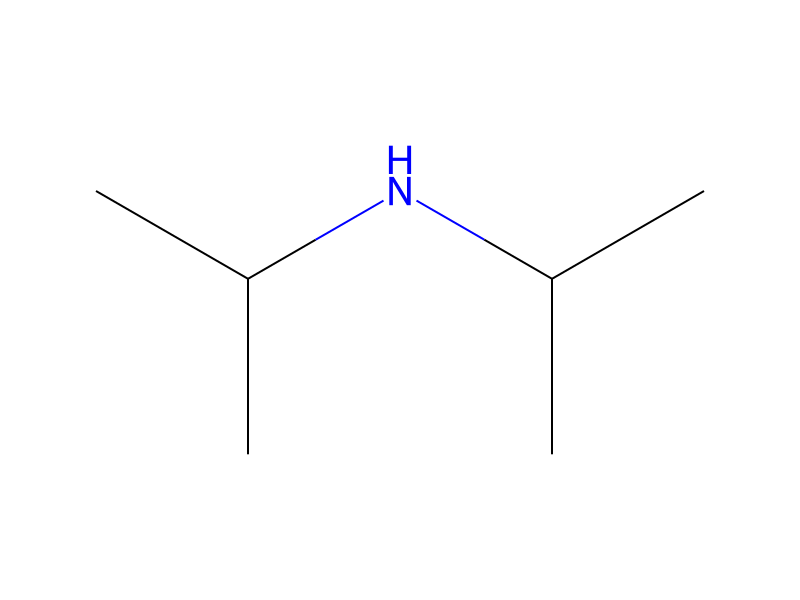
Diisopropylamine → 4-(Tert-butyldimethylsilyl)-3-chloro-2,5-difluoropyridine
反応式

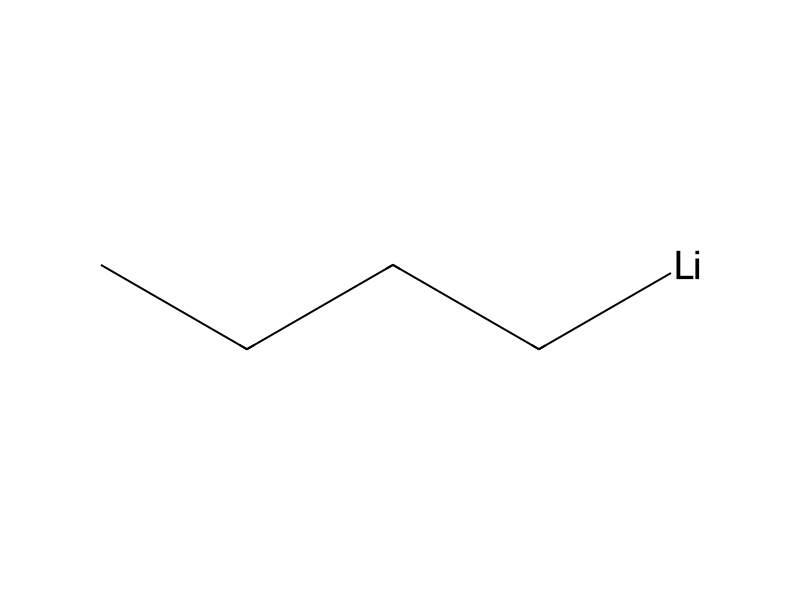
C(NC(C)C)(C)C.[Li]CCCC.[Cl:13][C:14]1[C:15]([F:21])=[N:16][CH:17]=[C:18]([F:20])[CH:19]=1.[C:22]([Si:26](Cl)([CH3:28])[CH3:27])([CH3:25])([CH3:24])[CH3:23].[Cl-].[NH4+]>C1COCC1.O.C(=O)(O)[O-].[Na+]>[Si:26]([C:19]1[C:18]([F:20])=[CH:17][N:16]=[C:15]([F:21])[C:14]=1[Cl:13])([C:22]([CH3:25])([CH3:24])[CH3:23])([CH3:28])[CH3:27]参加物
反応物 5


生成物 1

試薬・触媒・溶媒 3
反応条件
- 温度
- 0 °C
実験手順
A solution of diisopropylamine (78.52 g, 108.8 mL, 776.0 mmol) in THF (1.205 L) was cooled to −20° C. n-BuLi (2.5 M in hexanes) (298.4 mL of 2.5 M, 746.1 mmol) was added dropwise via cannula over 30 min at such a rate that the temperature was kept below −15° C. Once the addition was complete the cooling bath was removed and the reaction mixture was allowed to warm to 0° C. The reaction mixture was stirred for 15 min at 0° C., then re-cooled to −78° C. 3-chloro-2,5-difluoro-pyridine (95.963 g, 596.9 mmol) was added dropwise via cannula over 20 min at such a rate that the temperature was kept below −70° C. The reaction mixture was stirred for 45 min at −78° C. during which time the solution turned orange red. A solution of tert-butyl-chloro-dimethyl-silane (117.0 g, 776.0 mmol) in THF (133.9 mL) was added via cannula at such a rate to maintain the reaction temperature below −70° C. The reaction mixture was stirred at −78° C. for 90 minutes, during which time the solution turned to a deep red colour. Lc/Ms after this time indicated that the reaction was complete. A saturated ammonium chloride solution (300 ml) was then added and mixture was allowed to warm to ambient. The reaction mixture was diluted with water (200 ml) and saturated aqueous sodium bicarbonate (500 ml) and extracted with ethyl acetate (1.5 L then 500 ml). The combined organics were washed with brine (400 ml), dried over magnesium sulfate, filtered and concentrated in vacuo to an oil (187 g). The crude was purified by flash chromatography (CombiFlash Companion XL, 1.5 kg column, 0.5-10% ethyl acetate in petroleum ether). This afforded the title compound as a pale yellow oil (113.8 g, 72%); 1H NMR (CDCl3) 0.46 (6H, d), 0.93 (9H, s), 7.84 (1H, d); 19F NMR (decoupled) −112.8, −74.6; MS ES (+) 264.95 (M+1).
後処理
- additionwas added dropwise via cannula over 30 min at such a rate that the temperature
- customwas kept below −15° C
- additionOnce the addition
- customwas removed
- temperaturere-cooled to −78° C
- customwas kept below −70° C
- stirringThe reaction mixture was stirred for 45 min at −78° C. during which time the solution
- temperatureto maintain the reaction temperature below −70° C
- stirringThe reaction mixture was stirred at −78° C. for 90 minutes, during which time the solution
- temperatureto warm to ambient
- extractionextracted with ethyl acetate (1.5 L
- washThe combined organics were washed with brine (400 ml)
- dry with materialdried over magnesium sulfate
- filtrationfiltered
- concentrationconcentrated in vacuo to an oil (187 g)
- customThe crude was purified by flash chromatography (CombiFlash Companion XL, 1.5 kg column, 0.5-10% ethyl acetate in petroleum ether)
出典と根拠
- ORDレコード
- ord-e19eb72bc47e42a9a2f3430efc8e71fd
- 出典データセット
- uspto-grants-2015_06
- 特許番号
- US09067932B2
反応ノート
読み込み中…