ホームMCPワークベンチSkills
ホームワークベンチ通知マイページ
llms.txt · MCPガイドプライバシーポリシー · 利用規約 · サポートv1.8.3
  1. ホーム
  2. /反応

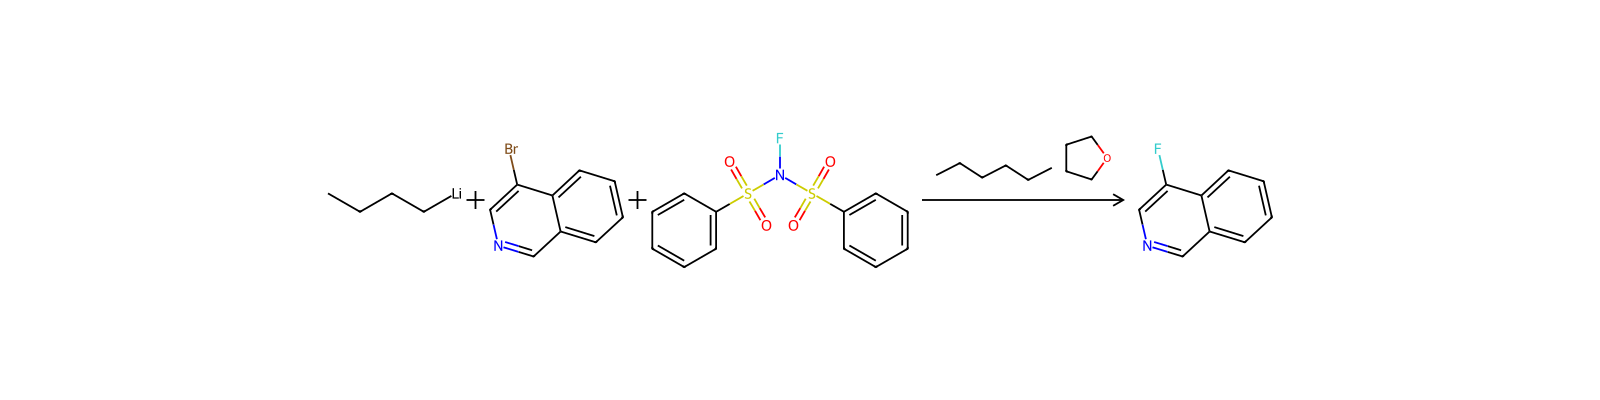
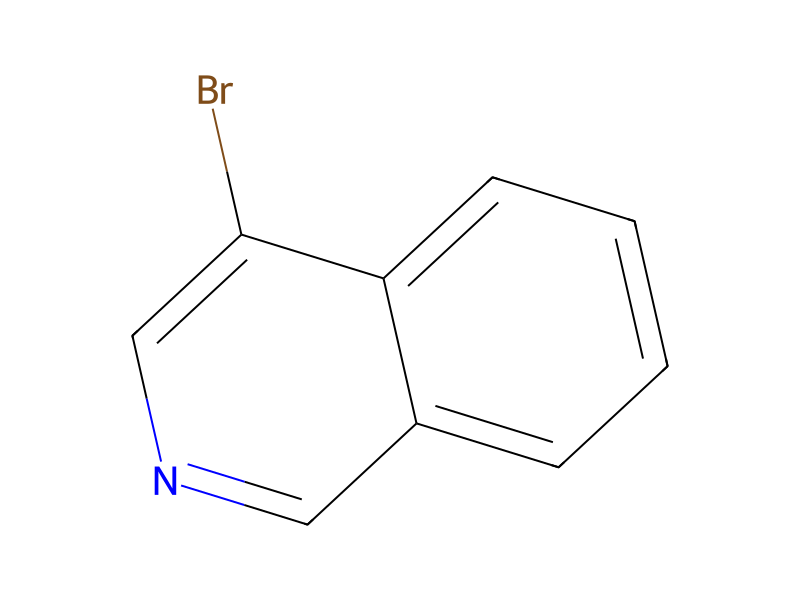
4-Bromoisoquinoline → 4-Fluoroisoquinoline

HRID 2425374
印刷

反応式

HRID 2425374 の反応式
C([Li])CCC.Br[C:7]1[C:16]2[C:11](=[CH:12][CH:13]=[CH:14][CH:15]=2)[CH:10]=[N:9][CH:8]=1.C1C=CC(S(N(S(C2C=CC=CC=2)(=O)=O)[F:27])(=O)=O)=CC=1>CCCCCC.O1CCCC1>[F:27][C:7]1[C:16]2[C:11](=[CH:12][CH:13]=[CH:14][CH:15]=2)[CH:10]=[N:9][CH:8]=1
参加物実験手順後処理出典と根拠反応ノート

参加物

6

反応物 3

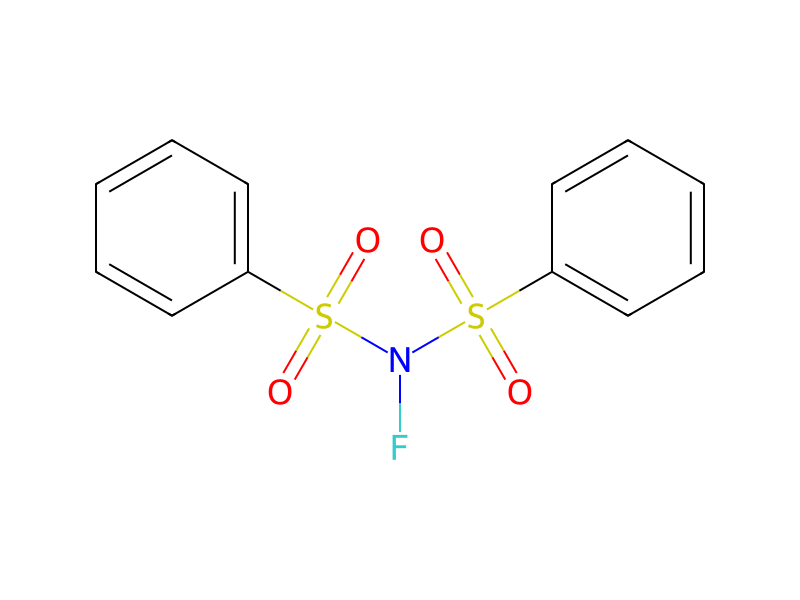
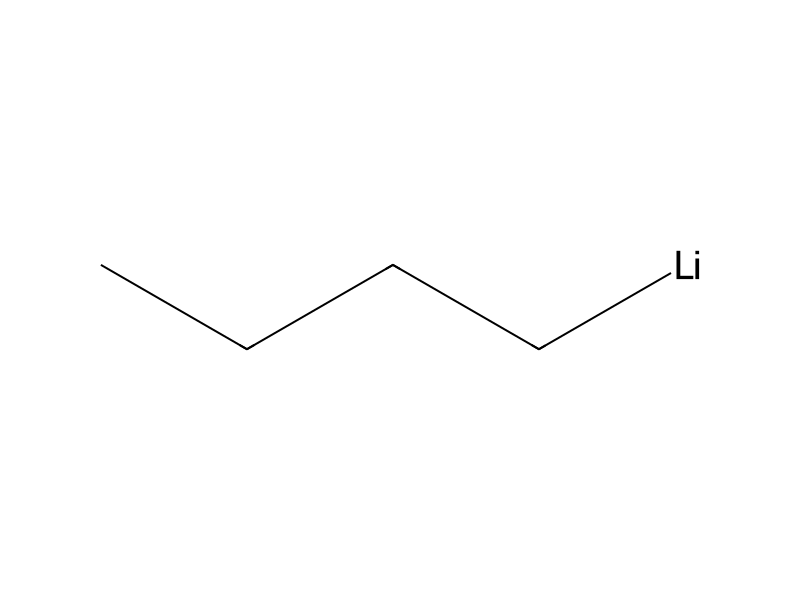
HCID 737434-BromoisoquinolineHCID 588007N-FluorobenzenesulfonimideHCID 178418339Butyllithium

生成物 1

HCID 6409734-Fluoroisoquinoline収率 56.6%

試薬・触媒・溶媒 2

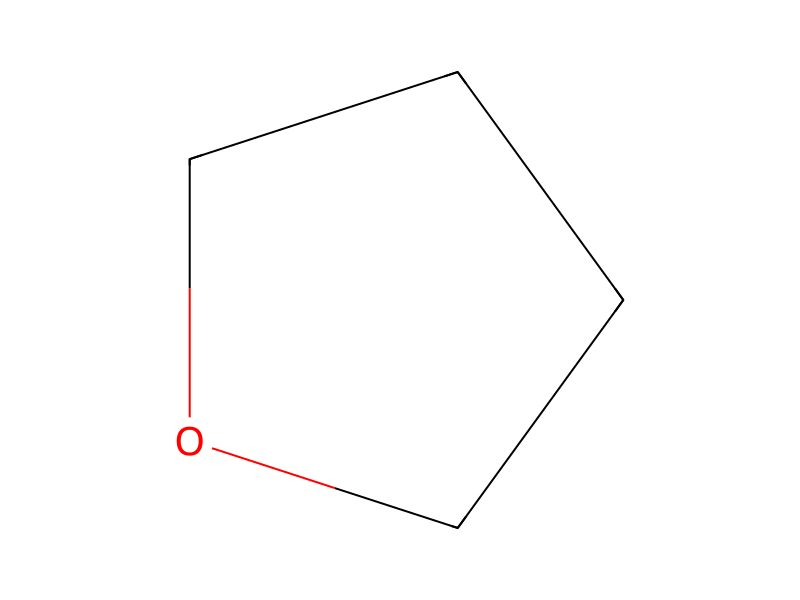
HCID 8028Tetrahydrofuran溶媒3 回HCID 8058Hexane溶媒

実験手順

A solution of n-butyllithium in n-hexane (1.58 M, 60.1 ml, Kanto Chemicals) was added with tetrahydrofuran (345 ml), and the mixture was sufficiently cooled on a dry ice-acetone bath. The mixture was added dropwise with a solution of 4-bromoisoquinoline (9.0 g, Tokyo Kasei Kogyo) in tetrahydrofuran (65 ml) over 1 hour so that the temperature of the reaction mixture should not exceed −65° C. The mixture was stirred at the same temperature for 30 minutes, and then added dropwise with a solution of N-fluorobenzenesulfonimide (30 g, Tokyo Kasei Kogyo) in tetrahydrofuran (100 ml) over 1 hour so that the temperature of the reaction mixture should not exceed −65° C. Subsequently, the mixture was stirred at the same temperature for 1 hour, then the cooling bath was removed, and the mixture was gradually warmed to room temperature. The reaction mixture was added with saturated aqueous sodium hydrogencarbonate (300 ml) and ethyl acetate (300 ml), and stirred at room temperature for 12 hours. The organic layer was separated, and the aqueous layer was extracted with ethyl acetate 3 times (200 ml for each time). The combined organic layer was washed with saturated brine (500 ml), and dried over anhydrous magnesium sulfate. The solvent was evaporated under reduced pressure, the residue was added with chloroform (250 ml), and the insoluble solids were removed by filtration. The solvent was evaporated under reduced pressure, and the residue was purified by silica gel chromatography (n-hexane:ethyl acetate=5:1) to obtain the title compound (3.6 g).
後処理

  1. temperaturethe mixture was sufficiently cooled on a dry ice-acetone bath
  2. customexceed −65° C
  3. customexceed −65° C
  4. stirringSubsequently, the mixture was stirred at the same temperature for 1 hour
  5. customthe cooling bath was removed
  6. additionThe reaction mixture was added with saturated aqueous sodium hydrogencarbonate (300 ml) and ethyl acetate (300 ml)
  7. stirringstirred at room temperature for 12 hours
  8. customThe organic layer was separated
  9. extractionthe aqueous layer was extracted with ethyl acetate 3 times (200 ml for each time)
  10. washThe combined organic layer was washed with saturated brine (500 ml)
  11. dry with materialdried over anhydrous magnesium sulfate
  12. customThe solvent was evaporated under reduced pressure
  13. additionthe residue was added with chloroform (250 ml)
  14. customthe insoluble solids were removed by filtration
  15. customThe solvent was evaporated under reduced pressure
  16. customthe residue was purified by silica gel chromatography (n-hexane:ethyl acetate=5:1)

出典と根拠

ORDレコード
ord-8f6fd553fbe44b20b5a6b565cea8f75a
出典データセット
uspto-grants-2011_06
DOI
10.6084/m9.figshare.5104873.v1
特許番号
US07964613B2

反応ノート

このページの内容参加物実験手順後処理出典と根拠反応ノート