反応式

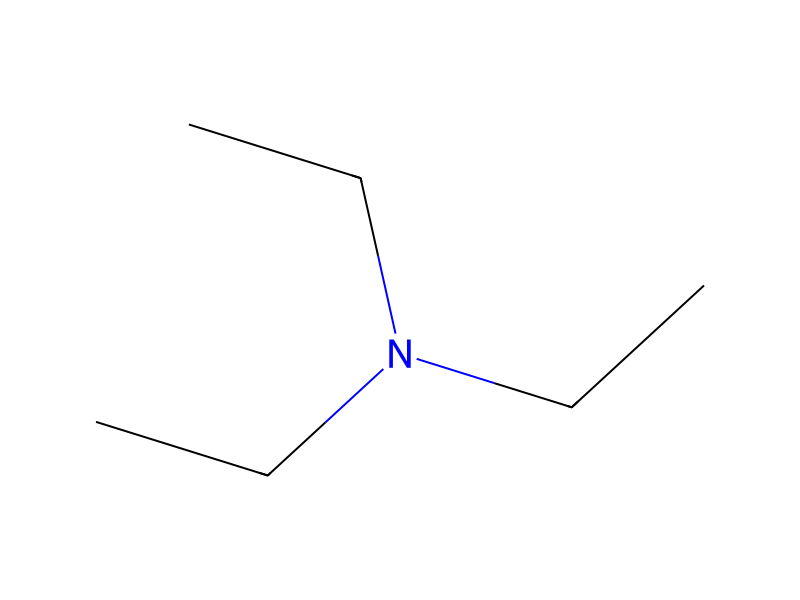
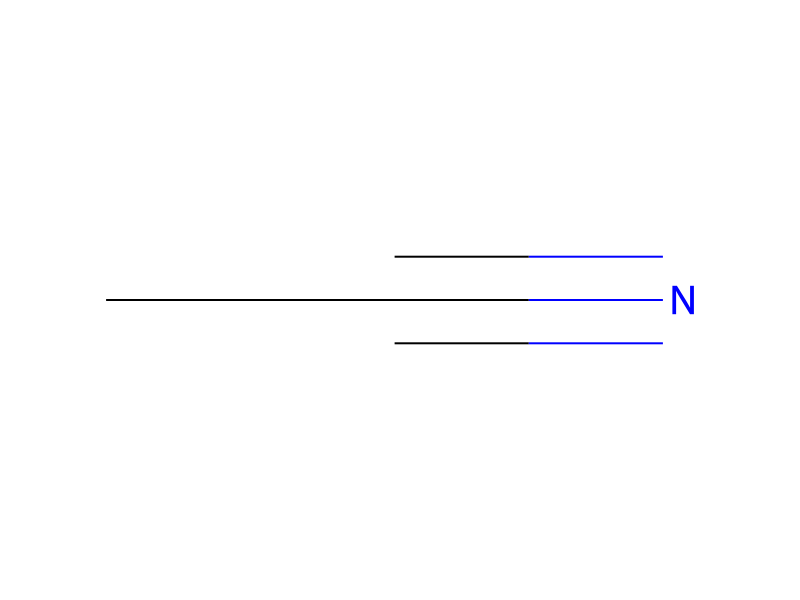
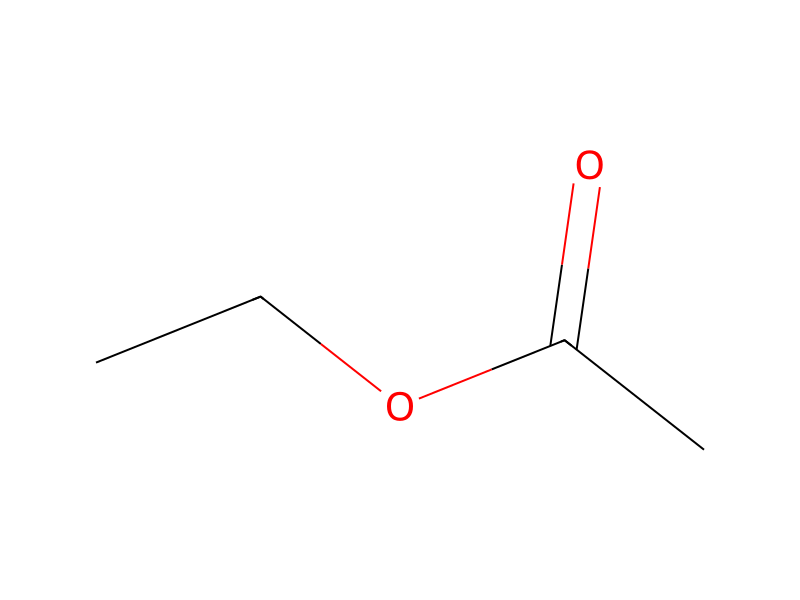
[CH2:1]=[C:2]1[CH:8]2[CH2:9][CH:5]([CH2:6][CH2:7]2)[C:4](=[O:10])[O:3]1.C(N(CC)CC)C.[C-]#N.[K+].[CH3:21][S:22]([C:25]1[CH:33]=[CH:32][C:28]([C:29](Cl)=[O:30])=[C:27]([N+:34]([O-:36])=[O:35])[CH:26]=1)(=[O:24])=[O:23].O=C1C2CC(CC2)C(OC(=O)C2C=CC(S(C)(=O)=O)=CC=2[N+]([O-])=O)=C1>C(#N)C.C(OCC)(=O)C>[OH:3][C:2]1[CH:8]2[CH2:9][CH:5]([CH2:6][CH2:7]2)[C:4](=[O:10])[C:1]=1[C:29](=[O:30])[C:28]1[CH:32]=[CH:33][C:25]([S:22]([CH3:21])(=[O:24])=[O:23])=[CH:26][C:27]=1[N+:34]([O-:36])=[O:35]参加物
反応物 5


生成物 1

試薬・触媒・溶媒 2
実験手順
2.76 g (0.02 mol) of 93% 4-methylene-3-oxabicyclo[3.2.1]octan-2-one and 4.45 g (0.044 mmol) of triethylamine are stirred for 7 hours at room temperature in 50 ml of acetonitrile in the presence of 130 mg (2 mmol) of potassium cyanide. 4.22 g (0.16 mol) of freshly prepared 4-methanesulfonyl-2-nitrobenzoyl chloride are then introduced and stirring is carried out for 12 hours at a temperature of 20° C. If, according to monitoring by thin-layer chromatography, relatively large amounts of the intermediate 4-methanesulfonyl-2-nitro-benzoic acid 4-oxobicyclo[3.2.1]oct-2-en-2-yl ester (known from U.S. Pat. No. 5,801,120) are still detectable, then a further 130 mg (2 mmol) of potassium cyanide are added and stirring is continued for a further 2 hours until conversion is complete. The reaction mixture is then taken up in ethyl acetate and extracted at a pH value of 13 against water. The aqueous phase is then acidified using hydrochloric acid and extracted at a pH value of approximately 4 using fresh ethyl acetate, dried over Na2SO4 and concentrated to a slight extent. The product, dissolved in warm ethyl acetate, is treated with activated carbon and filtered off with suction (suction filter) through a small amount of silica gel, then concentrated further and caused to crystallise by adding a small amount of hexane. 1.69 g of pure 4-hydroxy-3-(4-methanesulfonyl-2-nitrobenzoyl)-bicyclo[3.2.1]oct-3-en-2-one having a melting point of 170-170.5° C. are obtained. Further product (1.65 g) can be obtained from the mother liquor by recrystallisation. Total yield: 3.34 g, or 49.1% based on the starting material 4-methylene-3-oxabicyclo[3.2.1]octan-2-one used or 57.1% based on the starting material 4-methanesulfonyl-2-nitrobenzoic acid used. 1H-NMR (CDCl3): 15.94 ppm, s, 1H; 8.74, s, 1H; 8.24 ppm, d, 1H; 7.48 ppm, d, 1H; 3.20 ppm, m, 1H; 3.19 ppm, s, 3H; 2.84 ppm, m, 1H; 2.0-2.3 ppm, 4H; 1.6-1.8 ppm, 2H.
後処理
- stirringstirring
- waitis continued for a further 2 hours until conversion
- extractionextracted at a pH value of 13 against water
- extractionextracted at a pH value of approximately 4
- dry with materialdried over Na2SO4
- concentrationconcentrated to a slight extent
- dissolutionThe product, dissolved in warm ethyl acetate
- additionis treated with activated carbon
- filtrationfiltered off with suction (suction filter) through a small amount of silica gel
- concentrationconcentrated further
- customto crystallise
- additionby adding a small amount of hexane
出典と根拠
- ORDレコード
- ord-6f030612e75446b78372815cd92574d2
- 出典データセット
- uspto-grants-2005_02
- 特許番号
- US06855831B2
関連ノート
読み込み中…