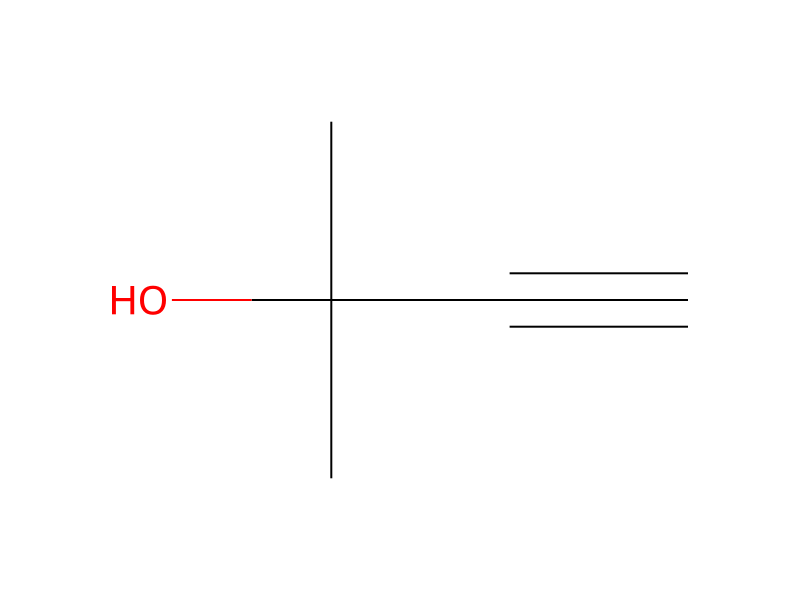
2-Methyl-3-butyn-2-ol → 8-fluoro-9-(3-hydroxy-3-methylbut-1-yn-1-yl)-5,6-dihydro-4H-4,6-methanobenzo[3,4]cyclohepta[1,2-b]thiophene-2-carboxamide
反応式

Br[C:2]1[C:3]([F:20])=[CH:4][C:5]2[CH:17]3[CH2:18][CH:15]([CH2:16]3)[C:8]3[S:9][C:10]([C:12]([NH2:14])=[O:13])=[CH:11][C:7]=3[C:6]=2[CH:19]=1.[CH3:21][C:22]([OH:26])([C:24]#[CH:25])[CH3:23]>[Cu]I.Cl[Pd](Cl)([P](C1C=CC=CC=1)(C1C=CC=CC=1)C1C=CC=CC=1)[P](C1C=CC=CC=1)(C1C=CC=CC=1)C1C=CC=CC=1.CN(C)C=O.C(N(CC)CC)C>[F:20][C:3]1[C:2]([C:25]#[C:24][C:22]([OH:26])([CH3:23])[CH3:21])=[CH:19][C:6]2[C:7]3[CH:11]=[C:10]([C:12]([NH2:14])=[O:13])[S:9][C:8]=3[CH:15]3[CH2:16][CH:17]([C:5]=2[CH:4]=1)[CH2:18]3参加物
反応物 2

生成物 1

試薬・触媒・溶媒 3

反応条件
- 温度
- 80 °C
実験手順
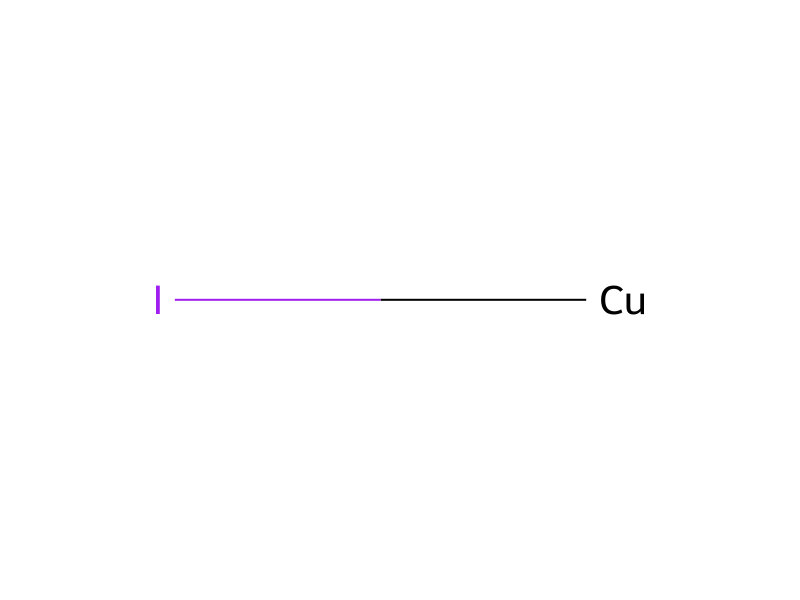
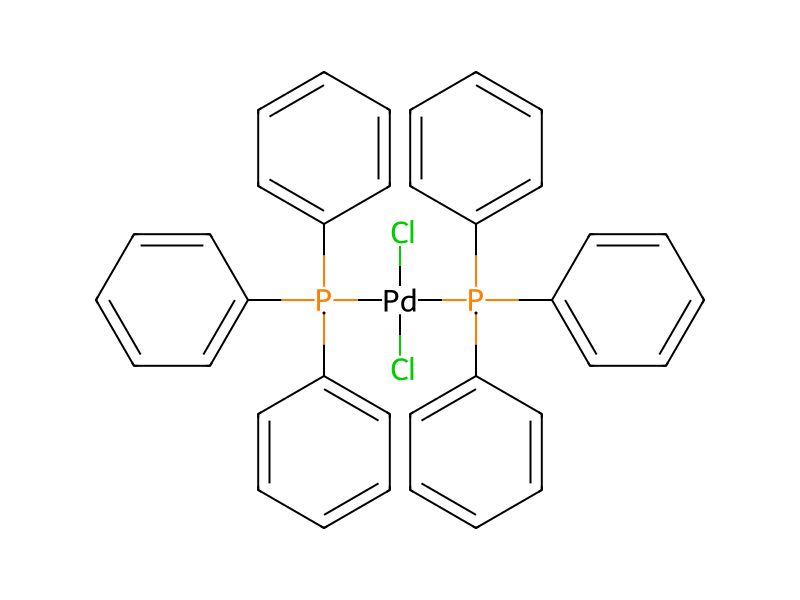
Into a pressure tube containing a solution of 9-bromo-8-fluoro-5,6-dihydro-4H-4,6-methanobenzo[3,4]cyclohepta[1,2-b]thiophene-2-carboxamide (31 mg, 0.08 mmol) in 1:1 N,N-dimethylformamide/triethylamine (0.8 mL) was introduced 2-methyl-3-butyn-2-ol (181 mg, 2.16 mmol), copper(I) iodide (2 mg, 0.01) and nitrogen gas bubbled through the solution for 3 minutes. Bis(triphenylphosphine)palladium(II) chloride (8 mg, 0.01 mmol) was added and nitrogen bubbling continued a further two minutes before the pressure tube was closed under nitrogen. The reaction mixture was warmed to 80° C. for 20 hr, re-treated with 2-methyl-3-butyn-2-ol (181 mg, 2.16 mmol), copper(I) iodide (2 mg, 0.01) and bis(triphenylphosphine)palladium(II) chloride (8 mg, 0.01 mmol) and warmed to 100° C. for an additional 7 hr. After cooling to room temperature, the reaction mixture was evaporated in vacuo and the residue re-dissolved in ethyl acetate (20 mL). This solution was washed with saturated aqueous sodium bicarbonate (10 mL), water (10 mL) and brine (5 mL). The organic extract was dried over sodium sulfate, filtered and the filtrate absorbed onto silica gel. Purification by silica gel flash column chromatography (eluent:heptane/ethyl acetate) furnished partially purified product as a brown solid. After several acetonitile solubilisation-evaporation (in vacuo) cycles, the resulting solid residue was slurried in acetonitrile (1.0 mL) and collected by filtration to furnish the title compound (17 mg, 0.05 mmol, 62%) as a pale-brown solid: 1H NMR (500 MHz, CDCl3) δ1.68 (s, 6H), 1.71-1.73 (m, 2H), 2.87-2.96 (m, 2H), 3.53-3.62 (m, 2H), 6.87 (d, J=10.09 Hz, 1H), 7.92 (d, J=6.94 Hz, 1H), 8.06 (s, 1H); M+H=356.
後処理
- custombubbled through the solution for 3 minutes
- customwas closed under nitrogen
- temperaturewarmed to 100° C. for an additional 7 hr
- temperatureAfter cooling to room temperature
- customthe reaction mixture was evaporated in vacuo
- dissolutionthe residue re-dissolved in ethyl acetate (20 mL)
- washThis solution was washed with saturated aqueous sodium bicarbonate (10 mL), water (10 mL) and brine (5 mL)
- extractionThe organic extract
- dry with materialwas dried over sodium sulfate
- filtrationfiltered
- customthe filtrate absorbed onto silica gel
- customPurification by silica gel flash column chromatography (eluent:heptane/ethyl acetate)
- customfurnished partially purified product as a brown solid
- customAfter several acetonitile solubilisation-evaporation (in vacuo) cycles
- filtrationcollected by filtration
出典と根拠
- ORDレコード
- ord-b6e43cd1f6374a08acb1fd05ced1aa46
- 出典データセット
- uspto-grants-2015_05
- 特許番号
- US09034866B2
反応ノート
読み込み中…