ホームMCPワークベンチSkills
ホームワークベンチ通知マイページ
llms.txt · MCPガイドプライバシーポリシー · 利用規約 · サポートv1.8.0
  1. ホーム
  2. /反応

Sodium Carbonate → HCID 21713907、化合物

HRID 649808
ノート
印刷プレビュー

反応式

HRID 649808 の反応式
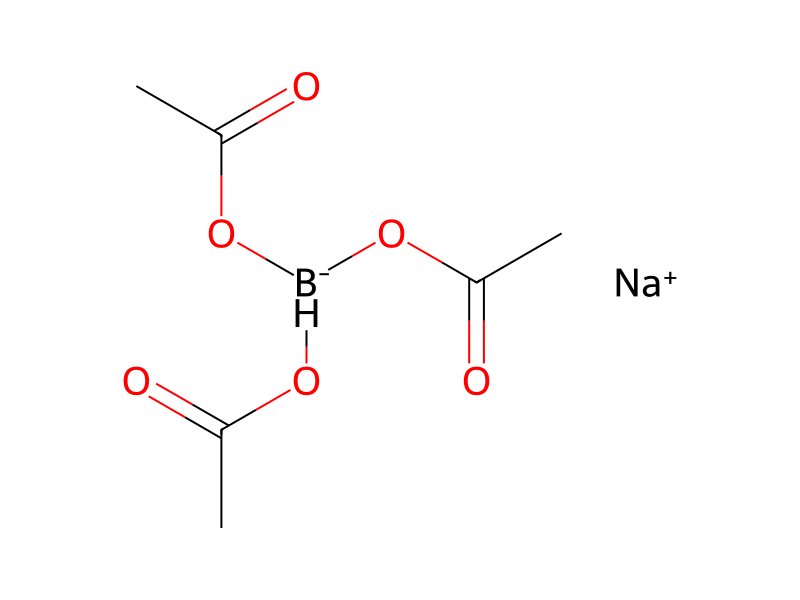
[CH:1]([CH2:3][N:4]([C:13]1[CH:18]=[CH:17][CH:16]=[CH:15][N:14]=1)[C:5]([CH:7]1[CH2:12][CH2:11][CH2:10][CH2:9][CH2:8]1)=[O:6])=O.[F:19][C:20]([F:35])([F:34])[O:21][C:22]1[CH:27]=[CH:26][CH:25]=[CH:24][C:23]=1[N:28]1[CH2:33][CH2:32][NH:31][CH2:30][CH2:29]1.C(O[BH-](OC(=O)C)OC(=O)C)(=O)C.[Na+].C([O-])([O-])=O.[Na+].[Na+]>O.ClCCCl.C(O)(=O)C>[CH:7]1([C:5]([N:4]([C:13]2[CH:18]=[CH:17][CH:16]=[CH:15][N:14]=2)[CH2:3][CH2:1][N:31]2[CH2:30][CH2:29][N:28]([C:23]3[CH:24]=[CH:25][CH:26]=[CH:27][C:22]=3[O:21][C:20]([F:34])([F:19])[F:35])[CH2:33][CH2:32]2)=[O:6])[CH2:12][CH2:11][CH2:10][CH2:9][CH2:8]1
参加物実験手順後処理出典と根拠反応ノート

参加物

8

反応物 4

HCID 10340Sodium CarbonateHCID 5049666Sodium triacetoxyborohydrideHCID 107529251-(2-Trifluoromethoxyphenyl)piperazineHCID 21713921N-Formylmethyl-N-(2-pyridyl)cyclohexanecarboxamide

生成物 1

HCID 21713907HCID 21713907、化合物収率 51.1%

試薬・触媒・溶媒 3

HCID 11
1,2-Dichloroethane
溶媒
HCID 176Acetic Acid溶媒
HCID 962Water溶媒

実験手順

A mixture comprising 0.459 g of Compound 1B, 0.445 g of 1-(2-trifluoromethoxyphenyl)piperaine (prepared as described by D. Clarke et al, in European Patent EP 711 757, 1996), 0.634 g sodium triacetoxyborohydride, 0.23 mL of glacial acetic acid and 15 mL of 1,2-dichloroethane was stirred for 4 h, under a nitrogen atmosphere. The resultant solution was then kept overnight at room temperature, followed by the addition of 10 mL of water, and alkalinization with a 20% aqueous Na2CO3 solution. This was followed by separation of phases, with the isolated aqueous phase being extracted with 2×20 mL dichloromethane, and with the collected organic phases being dried over anhydrous sodium sulphate. This was followed by the removal of solvents, via vacuum evaporation to dryness. The obtained product residue was purified by flash chromatography (ethyl acetate—petroleum ether gradient 90:10 to 10:0). This was followed by evaporation of the solvents, affording 0.44 g of the title compound (yield 51%).

後処理

  1. customThis was followed by separation of phases, with the isolated aqueous phase
  2. extractionbeing extracted with 2×20 mL dichloromethane
  3. dry with materialwith the collected organic phases being dried over anhydrous sodium sulphate
  4. customThis was followed by the removal of solvents, via vacuum evaporation to dryness
  5. customThe obtained product residue was purified by flash chromatography (ethyl acetate—petroleum ether gradient 90:10 to 10:0)
  6. customThis was followed by evaporation of the solvents

出典と根拠

ORDレコード
ord-0a90b7411b3c4eba8be42f1ec1927717
出典データセット
uspto-grants-2001_08
DOI
10.6084/m9.figshare.5104873.v1
特許番号
US06271234B1

反応ノート

読み込み中…

このページの内容参加物実験手順後処理出典と根拠反応ノート