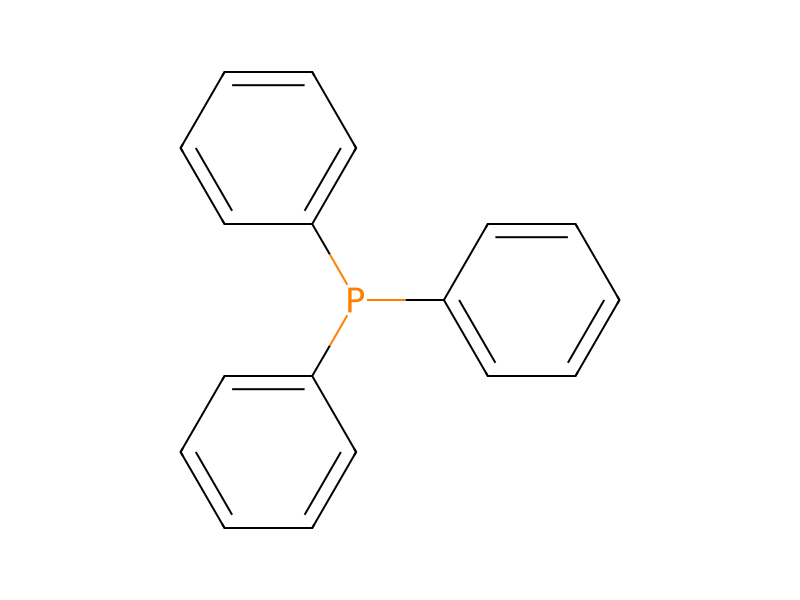
Triphenylphosphine → HCID 178476811, compound
Reaction equation

O[CH:2]1[CH2:6][CH2:5][C:4]([CH2:7][CH2:8][CH2:9][CH2:10][PH:11](=[O:15])[O:12][CH2:13][CH3:14])=[CH:3]1.CCOC(/[N:21]=N/C(OCC)=O)=O.N=[N+]=[N-].C1(P(C2C=CC=CC=2)C2C=CC=CC=2)C=CC=CC=1>C1C=CC=CC=1.C1COCC1.O>[NH2:21][CH:2]1[CH2:6][CH2:5][C:4]([CH2:7][CH2:8][CH2:9][CH2:10][PH:11](=[O:15])[O:12][CH2:13][CH3:14])=[CH:3]1Participants




Procedure
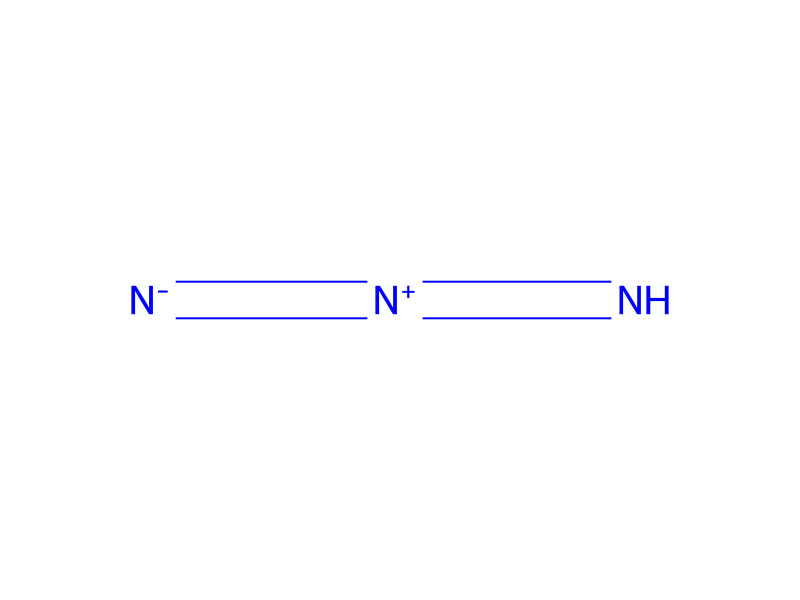
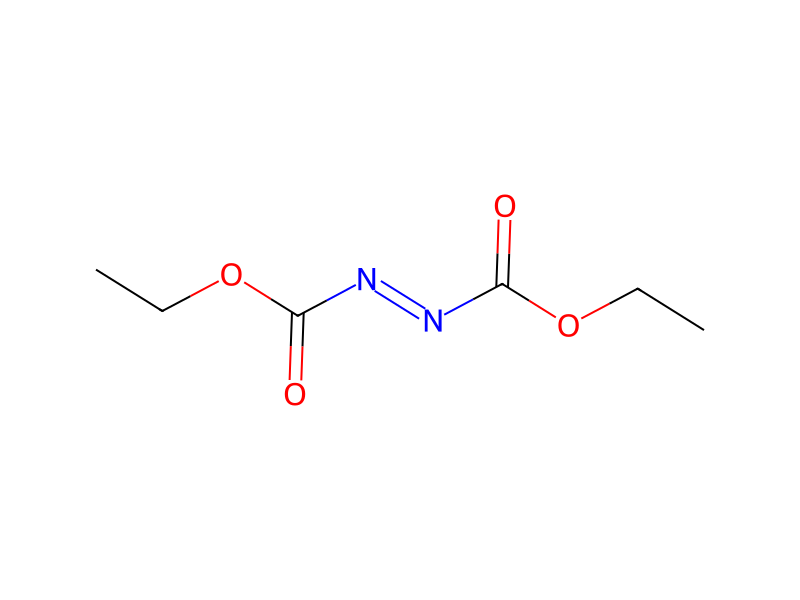
To a solution of (±)-ethyl (3-hydroxycyclopentenyl)butylphosphinate (7) (2.98 g, 12.8 mmol), DEAD (4.2 cm3) and HN3 (13.5 cm3 of a 1.9 M solution in benzene) in anhydrous THF (150 cm3) at 0° C. was added triphenylphosphine (13.43 g, 51.4 mmol) in small portions over a period of 1 hour. The reaction mixture was allowed to warm to room temperature and stirring continued for 12 hours. The reaction mixture was then heated to 50° C. for 3 hours after which time then water (2 cm3) was added and heating continued for a further 2 hours. The reaction mixture was cooled to room temperature and the solvent removed in vacuo. The residue was partitioned between aqueous HCl (1 M, 40 cm3) and DCM (40 cm3). The organic layer was separated and further extracted with water (3×30 cm3). The combined aqueous fractions were washed with DCM (2×30 cm3) and concentrated in vacuo. The crude product was purified by ion exchange chromatography (Dowex 50, H+), eluting first with water until the eluate was colourless and the pH 7. On further elution with aqueous ammonium hydroxide (1 M), ninhydrin positive fractions were collected and combined and the solvent was removed in vacuo. Recrystallisation from ethanol/acetone followed by drying gave the desired title compound (12) as a pale yellow oil (2.82 g, 95%): 1H-NMR (300 MHz, CDCl3) δH 0.88 (3H, 2×d, J=6.9 and 7.5 Hz, PCH2CH2CH2CH3),0.91 (3H, 2×d, J=7.2 and 7.2 Hz, PCH2CH2CH2CH3), 1.21-1.98 (9H, m, POCH2CH3, PCH2CH2CH2CH3, PCH2CH2CH2CH3,PCH2CH2CH2CH3), 2.38-2.71 (4H, m, C(4)—H, C(4)—H′, C(5)—H and C(5)—H′), 3.87-4.31 (5H, m, POCH2CH3, C(3)—NH2, C(3)—H), 6.51 (1H, d, J=9.6 Hz, C(2)—H); 13C-NMR (75 MHz, CDCl3) δC 13.43 (s, PCH2CH2CH2CH3),16.39 and 16.47 (d, 3JPOCC=6.00 Hz, POCH2CH3), 23.41 and 23.46 (2×d, 2JPC=3.38 and 3.75 Hz, PCH2CH2CH2CH3), 23.63 and 23.84 (d, 3JPC=15.90 Hz, PCH2CH2CH2CH3),27.15 and 28.47 (2×d, JPC=99.0 and 99.0 Hz, PCH2CH2CH2CH3), 31.93 and 32.09 (2×d, 3JPC=11.63 and 11.70 Hz, C(4)), 34.93 and 35.04 (2×d, 2JPC=8.25 and 8.55 Hz, C(5)), 58.86 and 59.09 (2×d, 3JPC=17.03 and 17.33 Hz, C(3)), 60.11 and 60.19 (d, 2JPOC=6.23 Hz, POCH2CH3), 135.63 and 137.72 (d, JPC=156.75 Hz, C(1)), 150.47 and 150.66 (2×d, 2JPC=13.88 and 13.95 Hz, C(2)); 31P-NMR (121 MHz, CDCl3) δP 43.18 and 43.28; MS (CI, CH4) m/z 232.0 (16%) (MH+), 215.1 (100), 446.0 (18), 260.2 (14). Synthesis of (±)-ethyl [3-(t-butyloxycarbonyl)aminocyclopentenyl]butylphosphinate (13)
Workup
- temperatureThe reaction mixture was then heated to 50° C. for 3 hours after which time
- temperatureheating
- waitcontinued for a further 2 hours
- temperatureThe reaction mixture was cooled to room temperature
- customthe solvent removed in vacuo
- customThe residue was partitioned between aqueous HCl (1 M, 40 cm3) and DCM (40 cm3)
- customThe organic layer was separated
- extractionfurther extracted with water (3×30 cm3)
- washThe combined aqueous fractions were washed with DCM (2×30 cm3)
- concentrationconcentrated in vacuo
- customThe crude product was purified by ion exchange chromatography (Dowex 50, H+)
- washeluting first with water until the eluate
- washOn further elution with aqueous ammonium hydroxide (1 M), ninhydrin positive fractions
- customwere collected
- customthe solvent was removed in vacuo
- customRecrystallisation from ethanol/acetone
- customby drying
Sources & evidence
- ORD record
- ord-7d69be26156d44be97ee41f13c144095
- Source dataset
- uspto-grants-2005_11
- Patent number
- US06962907B2
Reaction notes
Loading…