1,2-dimethyl-1H-imidazole-5-carbaldehyde → HCID 118005806, compound
Reaction equation

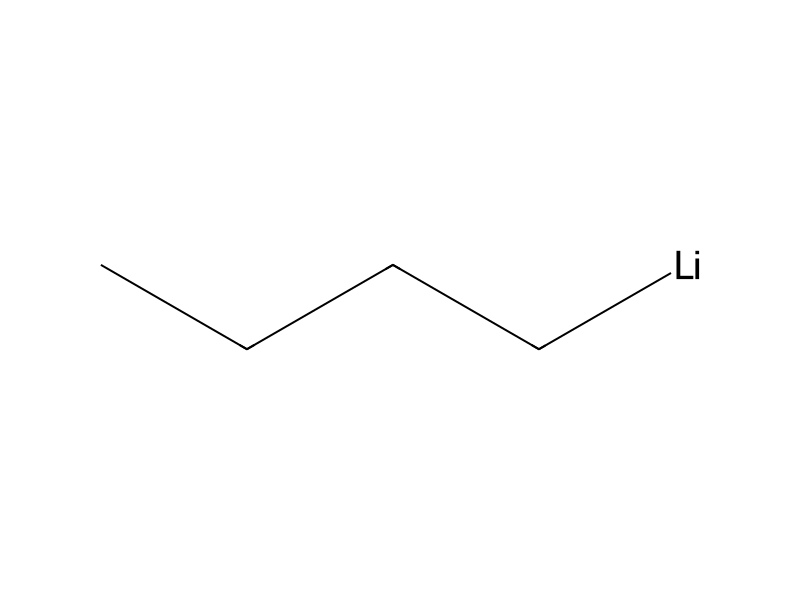
[N:1]1([C:6]2[CH:28]=[CH:27][C:9]([CH2:10][C:11]3[C:12]([N:23]4[CH2:26][CH2:25][CH2:24]4)=[N:13][C:14]4[C:19]([C:20]=3[Cl:21])=[CH:18][C:17](Br)=[CH:16][CH:15]=4)=[CH:8][CH:7]=2)[CH:5]=[CH:4][CH:3]=[N:2]1.[Li]CCCC.[CH3:34][N:35]1[C:39]([CH:40]=[O:41])=[CH:38][N:37]=[C:36]1[CH3:42]>C1COCC1>[N:23]1([C:12]2[C:11]([CH2:10][C:9]3[CH:27]=[CH:28][C:6]([N:1]4[CH:5]=[CH:4][CH:3]=[N:2]4)=[CH:7][CH:8]=3)=[C:20]([Cl:21])[C:19]3[C:14](=[CH:15][CH:16]=[C:17]([CH:40]([C:39]4[N:35]([CH3:34])[C:36]([CH3:42])=[N:37][CH:38]=4)[OH:41])[CH:18]=3)[N:13]=2)[CH2:26][CH2:25][CH2:24]1Participants
Reactant 3


Product 1

Reagents, catalysts & solvents 1

Conditions
- Temperature
- -78 °C
Procedure
3-(4-(1H-Pyrazol-1-yl)benzyl)-2-(azetidin-1-yl)-6-bromo-4-chloroquinoline (1.03 g, 2.27 mmol, Intermediate 17) and THF (15 mL) were combined in the reaction vessel under an N2 atmosphere and cooled to −78° C. in a dry ice acetone bath. n-BuLi (1.6 M in hexanes, 1.70 mL, 2.72 mmol) was then added dropwise via syringe over approximately one minute and contents were allowed to stir at −78° C. for an additional 5 minutes. 1,2-Dimethyl-1H-imidazole-5-carbaldehyde (0.31 g, 2.5 mmol) in THF (5 mL) was then cannulated into the reaction vessel and the reaction was stirred at −78° C. for 10 minutes. The dry ice bath was removed and replaced by an ice water bath and the reaction continued for approximately one hour at 0° C. The reaction was then quenched with a saturated, aqueous NH4Cl solution, then transferred to a separatory funnel with EtOAc. The organic phase was separated and the aqueous layer was back extracted with EtOAc, then the combined organic layers were dried over MgSO4, filtered and concentrated under reduced pressure. The crude product was purified by flash column chromatography (silica gel, 0-10% DCM/(10% of a 2 M NH3 MeOH in DCM)) to provide the title compound. MS (ESI): mass calcd. for C28H27ClN6O, 498.2; m/z found, 499.5 [M+H]+. 1H NMR (400 MHz, CDCl3) δ ppm 8.16-8.12 (m, 1H), 7.88 (dd, J=2.5, 0.6 Hz, 1H), 7.75 (d, J=8.6 Hz, 1H), 7.70 (dd, J=1.8, 0.6 Hz, 1H), 7.62-7.54 (m, 3H), 7.19 (d, J=8.5 Hz, 2H), 6.53 (s, 1H), 6.44 (dd, J=2.5, 1.8 Hz, 1H), 5.97 (s, 1H), 4.33 (s, 2H), 4.19-4.12 (m, 4H), 3.49 (s, 3H), 2.34 (s, 3H), 2.27-2.18 (m, 2H).
Workup
- stirringthe reaction was stirred at −78° C. for 10 minutes
- customThe dry ice bath was removed
- customreplaced by an ice water bath
- waitthe reaction continued for approximately one hour at 0° C
- customThe reaction was then quenched with a saturated, aqueous NH4Cl solution
- customtransferred to a separatory funnel with EtOAc
- customThe organic phase was separated
- extractionthe aqueous layer was back extracted with EtOAc
- dry with materialthe combined organic layers were dried over MgSO4
- filtrationfiltered
- concentrationconcentrated under reduced pressure
- customThe crude product was purified by flash column chromatography (silica gel, 0-10% DCM/(10% of a 2 M NH3 MeOH in DCM))
Sources & evidence
- ORD record
- ord-43a6bac3f83944b1b82f8951805a315f
- Source dataset
- uspto-grants-2015_12
- Patent number
- US09221804B2
Reaction notes
Loading…