Acetaldehyde → HCID 13833724, compound
Reaction equation

[CH3:1][CH2:2][O:3]CC.Br[C:7]1[CH:8]=[CH:9][C:10]2[O:14][CH2:13][C:12]([CH3:16])([CH3:15])[C:11]=2[CH:17]=1.C(=O)C.[NH4+].[Cl-]>C1COCC1>[CH3:15][C:12]1([CH3:16])[C:11]2[CH:17]=[C:7]([CH:2]([OH:3])[CH3:1])[CH:8]=[CH:9][C:10]=2[O:14][CH2:13]1Participants
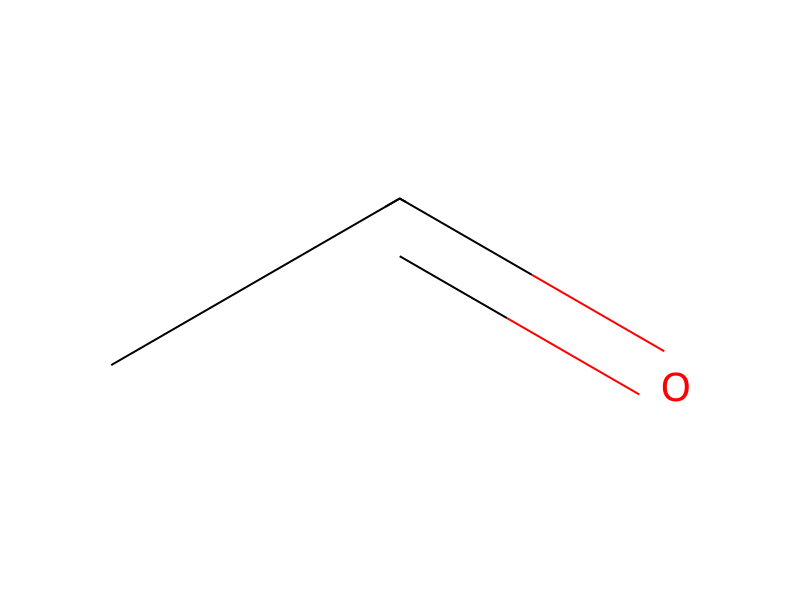
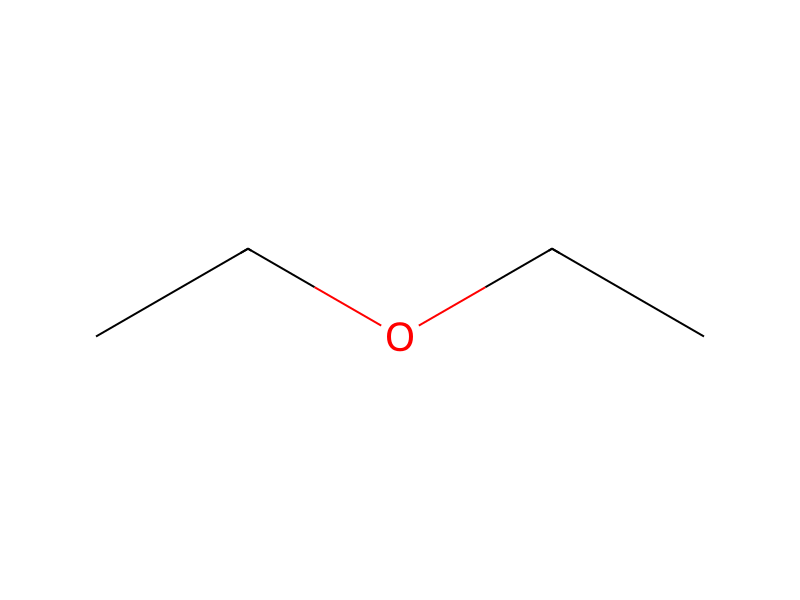



Procedure
A mixture of the precursor ether (0.21 g 0.9 mmol), Mg turnings (1.0 g 0.041 g at) and dry THF (2 mL) was heated (heat gun) under N2 until the mixture turned cloudy (ca 15 min). Dry THF (15 mL) was added to the mixture which was then heated to reflux. A solution of ether (42) (2.92 g, 12.9 mmol) in dry THF (25 mL) was added dropwise to the vigorously stirred mixture over a period of 0.75 hours. After vigorous stirring at reflux for 2.75 hours, another 0.25 g (0.010 g at) of Mg turnings were added. The new mixture was stirred at reflux for 0.75 hour and with no external heat for 0.5 hours. Upon cooling the mixture to -5° to -10° C. (ice-salt bath), a solution of freshly distilled acetaldehyde (2.0 g, 0.045 mol) in dry THF (20 mL) was added dropwise to the vigorously stirred mixture over a period of 0.7 hours. This reaction mixture was stirred in an ice-salt bath (-5° to -10° C.), for 1.5 hours, after which time a solution of acetaldehyde (0.9 g, 0.020 mol) in dry THF (5 mL) was added dropwise (over a period of about 0.2 h) and the new mixture was stirred 0.3 hours. With continued cooling (-5° to -10° C.), saturated aqueous NH4Cl (3 mL) was added and the excess Mg turnings were removed by filtration. Saturated aqueous NH4Cl (10 mL) and ether rinses (of glassware and Mg turnings, 60 mL) were added to the filtrate. After separating the organic layer, the aqueous phase (pH≥8) was acidified (pH 6.5 to 7) with saturated aqueous NH4Cl (15 mL) and 4% H2SO4 (14 mL). The aqueous solution was extracted with ether (5 ×40 mL) and ether (90 mL) was added to the combined organics. The organic solution was washed with saturated aqueous NaHCO3 (75 mL) and brine (50 mL). After drying (MgSO4) the solution, the solvent was removed, and the residual oil was chromatographed through a circular silica gel plate (4 mm) spun by a Chromatotron (Model 7924T, Harrison Research, 840 Moana Court, Palo Alto, Calif.). Half of the product was eluted with ether/petroleum ether (bp 50°-110° C.) 20:1, 8:1, and 4:1 and then the same solvent system was used to elute the other half. In both separations, the 4:1 ratio was required to elute the title compound. Concentration of the eluent in the desired fractions gave 1.18 g (44%) of the alcohol as a light yellow, viscous oil. TLC analysis (1:4 ether:petroleum ether, bp 50°-110° C.) indicated the compound was essentially pure.
Workup
- temperaturewas heated
- temperature(heat gun) under N2 until the mixture
- custom(ca 15 min)
- temperaturewas then heated to reflux
- temperatureat reflux for 2.75 hours
- stirringThe new mixture was stirred
- temperatureat reflux for 0.75 hour
- temperaturewith no external heat for 0.5 hours
- stirringThis reaction mixture was stirred in an ice-salt bath (-5° to -10° C.)
- stirringwas stirred 0.3 hours
- additionwas added
- customthe excess Mg turnings were removed by filtration
- additionSaturated aqueous NH4Cl (10 mL) and ether rinses (of glassware and Mg turnings, 60 mL) were added to the filtrate
- customAfter separating the organic layer
- extractionThe aqueous solution was extracted with ether (5 ×40 mL) and ether (90 mL)
- additionwas added to the combined organics
- washThe organic solution was washed with saturated aqueous NaHCO3 (75 mL) and brine (50 mL)
- dry with materialAfter drying (MgSO4) the solution
- customthe solvent was removed
- customthe residual oil was chromatographed through a circular silica gel plate (4 mm)
- washHalf of the product was eluted with ether/petroleum ether (bp 50°-110° C.) 20:1, 8:1, and 4:1
- washto elute the other half
- washto elute the title compound
Sources & evidence
- ORD record
- ord-072e23647ac34293aa251295c1291be1
- Source dataset
- uspto-grants-1989_05
- Patent number
- US04833254
Reaction notes
Loading…