Water → 2',6'-Dimethyl-4'-(3-(methylsulfonyl)propoxy)biphenyl-3-carbaldehyde
Reaction equation

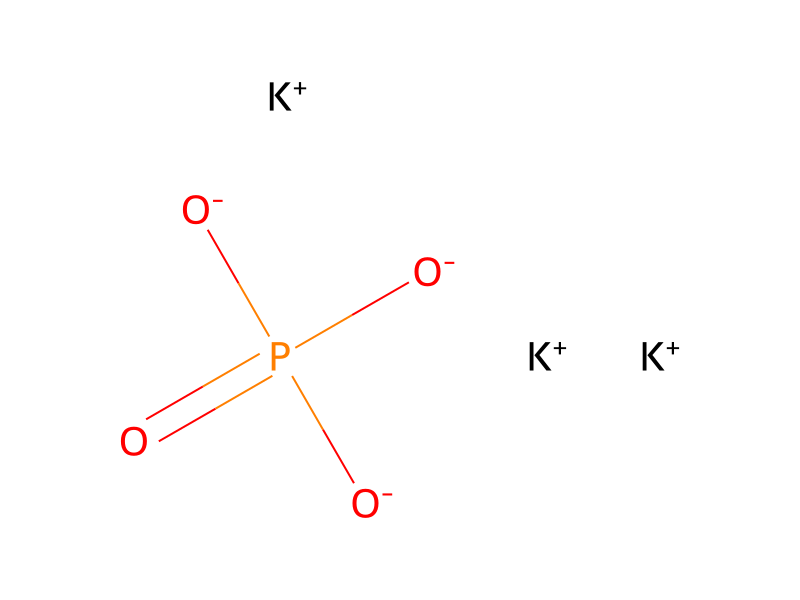
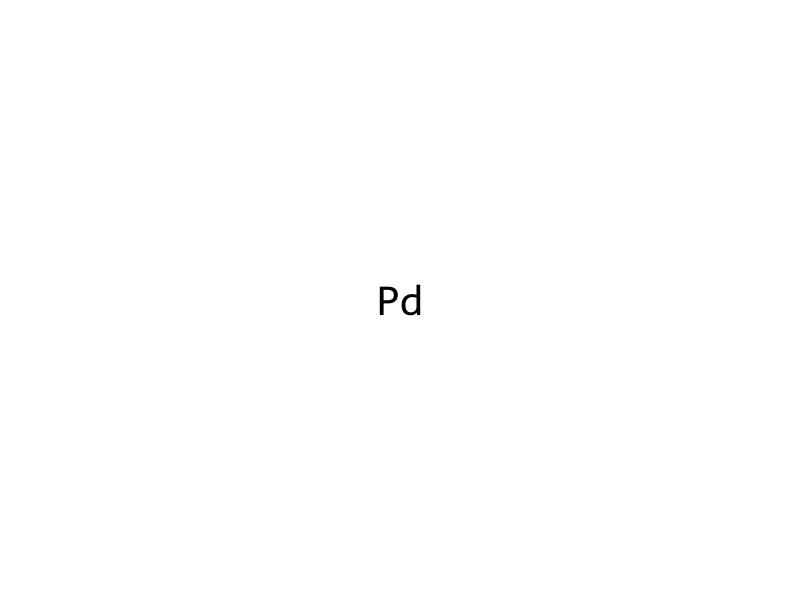
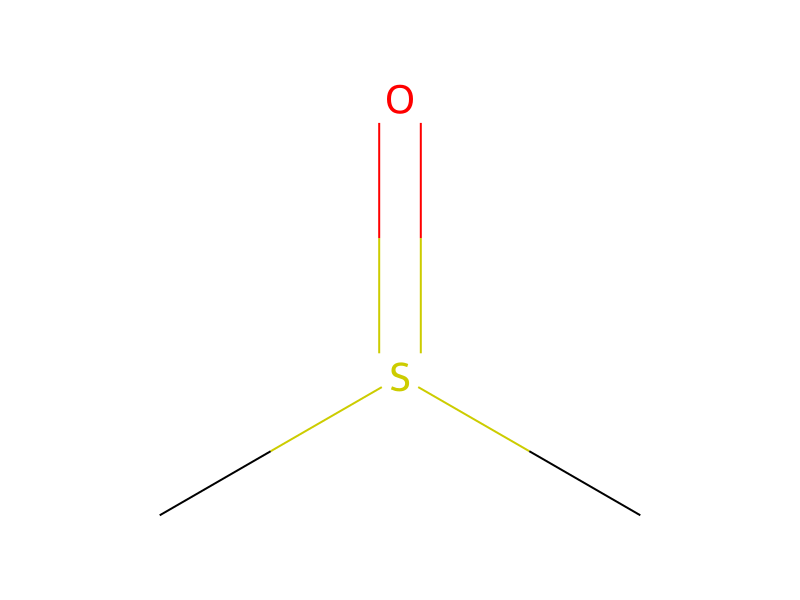
Br[C:2]1[C:7]([CH3:8])=[CH:6][C:5]([O:9][CH2:10][CH2:11][CH2:12][S:13]([CH3:16])(=[O:15])=[O:14])=[CH:4][C:3]=1[CH3:17].[CH:18]([C:20]1[CH:21]=[C:22](B(O)O)[CH:23]=[CH:24][CH:25]=1)=[O:19].P([O-])([O-])([O-])=O.[K+].[K+].[K+].O>CS(C)=O.[Pd].C1(P(C2C=CC=CC=2)[C-]2C=CC=C2)C=CC=CC=1.[C-]1(P(C2C=CC=CC=2)C2C=CC=CC=2)C=CC=C1.[Fe+2]>[CH3:17][C:3]1[CH:4]=[C:5]([O:9][CH2:10][CH2:11][CH2:12][S:13]([CH3:16])(=[O:15])=[O:14])[CH:6]=[C:7]([CH3:8])[C:2]=1[C:24]1[CH:23]=[CH:22][CH:21]=[C:20]([CH:18]=[O:19])[CH:25]=1Participants
Reactant 4



Product 1

Reagents, catalysts & solvents 3

Conditions
- Temperature
- 80 °C
Procedure
2-Bromo-1,3-dimethyl-5-[3-(methylsulfonyl)propoxy]benzene (50.0 g) and 3-formylphenylboronic acid (24.5 g, 1.05 eq) were dissolved in dimethyl sulfoxide (500 mL), and 10% Pd—C (PE type) (725.0 mg, 0.2 mol %) and 1,1′-bis(diphenylphosphino)ferrocene (171.6 mg, 0.2 mol %) were added. After nitrogen substitution, a deaeration operation was performed under reduced pressure three times, and the mixture was stirred at inside temperature 80±5° C. for 1 hr. A solution of separately prepared tripotassium phosphate (66.1 g, 2.0 eq)/water (250 mL) was added to the reaction mixture at the same temperature, and the mixture was stirred at the same temperature for 4.5 hr. After cooling to 60° C., the mixture was partitioned. To the organic layer was added ethyl acetate (175 mL), and the mixture was cooled to 30° C. Pd—C was filtered off, and washed with ethyl acetate (125 mL). To the filtrate was added 10% brine (300 mL), and the mixture was partitioned. The aqueous layer was extracted with ethyl acetate (300 mL). The organic layers were combined and washed with 10% brine (300 mL×2). The organic layer was concentrated to about 150 mL, to the residue was added ethyl acetate (300 mL), and the mixture was concentrated to about 150 mL. To the residue were added ethyl acetate (250 mL) and activated carbon Shirasagi A (5 g), and the mixture was stirred for 30 min. The activated carbon was filtered off, and washed with ethyl acetate (150 mL). The filtrate was concentrated to about 150 mL, and the mixture was stirred at room temperature overnight. Heptane (750 mL) was added dropwise, and the mixture was stirred at room temperature for 2 hr. The crystals were filtered, and washed with ethyl acetate/heptane (25 mL/125 mL). The crystal was dried at 60° C. under reduced pressure to give the title compound (46.9 g, yield 90.0%).
Workup
- stirringthe mixture was stirred at the same temperature for 4.5 hr
- temperatureAfter cooling to 60° C.
- customthe mixture was partitioned
- additionTo the organic layer was added ethyl acetate (175 mL)
- temperaturethe mixture was cooled to 30° C
- filtrationPd—C was filtered off
- washwashed with ethyl acetate (125 mL)
- additionTo the filtrate was added 10% brine (300 mL)
- customthe mixture was partitioned
- extractionThe aqueous layer was extracted with ethyl acetate (300 mL)
- washwashed with 10% brine (300 mL×2)
- concentrationThe organic layer was concentrated to about 150 mL, to the residue
- additionwas added ethyl acetate (300 mL)
- concentrationthe mixture was concentrated to about 150 mL
- additionTo the residue were added ethyl acetate (250 mL) and activated carbon Shirasagi A (5 g)
- stirringthe mixture was stirred for 30 min
- filtrationThe activated carbon was filtered off
- washwashed with ethyl acetate (150 mL)
- concentrationThe filtrate was concentrated to about 150 mL
- stirringthe mixture was stirred at room temperature overnight
- additionHeptane (750 mL) was added dropwise
- stirringthe mixture was stirred at room temperature for 2 hr
- filtrationThe crystals were filtered
- washwashed with ethyl acetate/heptane (25 mL/125 mL)
- customThe crystal was dried at 60° C. under reduced pressure
Sources & evidence
- ORD record
- ord-10e3e58b9bcb4c7ca68ea025e3108347
- Source dataset
- uspto-grants-2015_02
- Patent number
- US08952185B2
Reaction notes
Loading…