Benzaldehyde → 4-METHOXYBENZOIN
Reaction equation

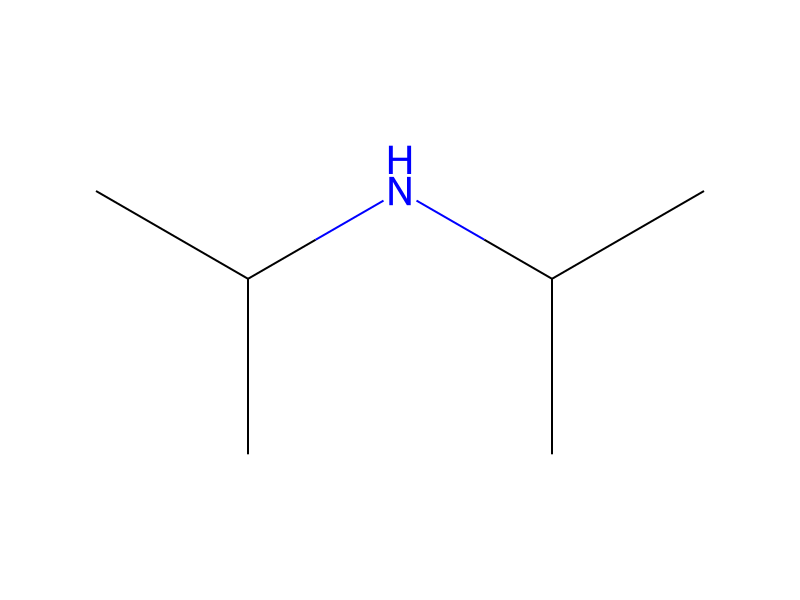
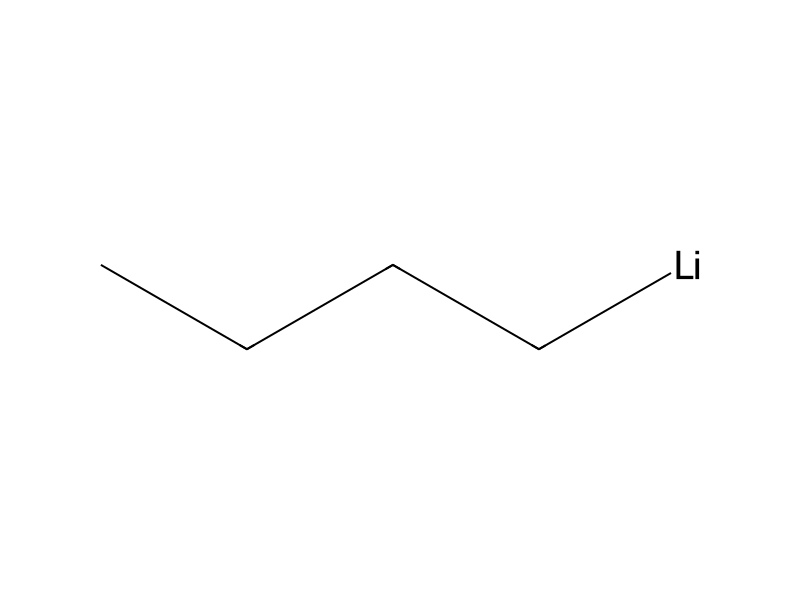
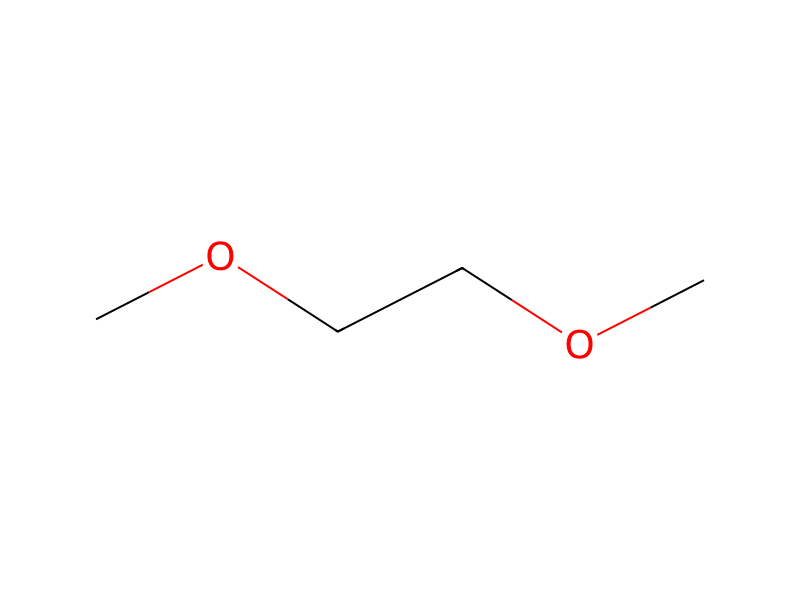
C(NC(C)C)(C)C.C([Li])CCC.[CH3:13][O:14][C:15]1[CH:20]=[CH:19][C:18]([CH:21]([O:24][Si](C)(C)C)C#N)=[CH:17][CH:16]=1.[CH:29](=[O:36])[C:30]1[CH:35]=[CH:34][CH:33]=[CH:32][CH:31]=1.[Cl-].[NH4+]>COCCOC>[OH:36][CH:29]([C:30]1[CH:35]=[CH:34][CH:33]=[CH:32][CH:31]=1)[C:21]([C:18]1[CH:19]=[CH:20][C:15]([O:14][CH3:13])=[CH:16][CH:17]=1)=[O:24]Participants
Reactant 5


Product 1

Reagents, catalysts & solvents 1
Conditions
- Temperature
- -78 °C
Procedure
According to the procedure in the literature [J. Chem. Soc. Perkin Trans. I, 1992, 2409-2417], dissolve 292 ml (2.08 mol) diisopropylamine in 3.6 litre 1,2-dimethoxyethane and cool to −78° C. Add 826 ml n-butyllithium solution (2.5 M in n-hexane, 2.066 mol) at a temperature below −60° C. Stir the mixture for a further 15 min at <−60° C. and then add a solution of 442 g (1.877 mol) (4-methoxyphenyl)[(trimethylsilyl)oxy]acetonitrile in 1.41 litre 1,2-dimethoxyethane dropwise at <−60° C. After further stirring for 30 min at −60° C., add a solution of 199.3 g (1.878 mol) benzaldehyde in 1.4 litre 1,2-dimethoxyethane in the space of 20 min at −60° C. Next, heat the reaction mixture slowly to RT in 4 h. Add 7 litre saturated ammonium chloride solution and extract with ethyl acetate. Wash the organic phase with saturated ammonium chloride solution, dry, and concentrate under vacuum. Take up the residue in 7 litre dioxan and 5 litre methanol, and add 6 litre 1 N hydrochloric acid. Stir the mixture for 3 h at RT, then add 3 litre saturated sodium chloride solution and extract the mixture with 6.5 litre ethyl acetate. Wash the organic phase with 1.0 litre 1 N sodium hydroxide solution and with saturated sodium chloride solution, dry, and concentrate under vacuum. Take up the residue in 2 litre diisopropyl ether, decant from the insoluble matter and seed with crystals. Stir the resultant suspension for 2 h at RT and then filter off the crystals with suction. Wash with 300 ml diisopropyl ether and petroleum ether and dry under vacuum. 236.8 g (47.8% of theor.) of the target compound is obtained.
Workup
- stirringAfter further stirring for 30 min at −60° C.
- temperatureNext, heat the reaction mixture slowly to RT in 4 h
- extractionextract with ethyl acetate
- washWash the organic phase with saturated ammonium chloride solution
- customdry
- concentrationconcentrate under vacuum
- addition5 litre methanol, and add 6 litre 1 N hydrochloric acid
- stirringStir the mixture for 3 h at RT
- additionadd 3 litre saturated sodium chloride solution
- extractionextract the mixture with 6.5 litre ethyl acetate
- washWash the organic phase with 1.0 litre 1 N sodium hydroxide solution
- dry with materialwith saturated sodium chloride solution, dry
- concentrationconcentrate under vacuum
- customdecant from the insoluble matter and seed with crystals
- stirringStir the resultant suspension for 2 h at RT
- filtrationfilter off the crystals with suction
- washWash with 300 ml diisopropyl ether and petroleum ether
- customdry under vacuum
- custom236.8 g (47.8% of theor.) of the target compound is obtained
Sources & evidence
- ORD record
- ord-efd03f6c65394ad08e4dba7da981934a
- Source dataset
- uspto-grants-2012_05
- Patent number
- US08183246B2
Reaction notes
Loading…