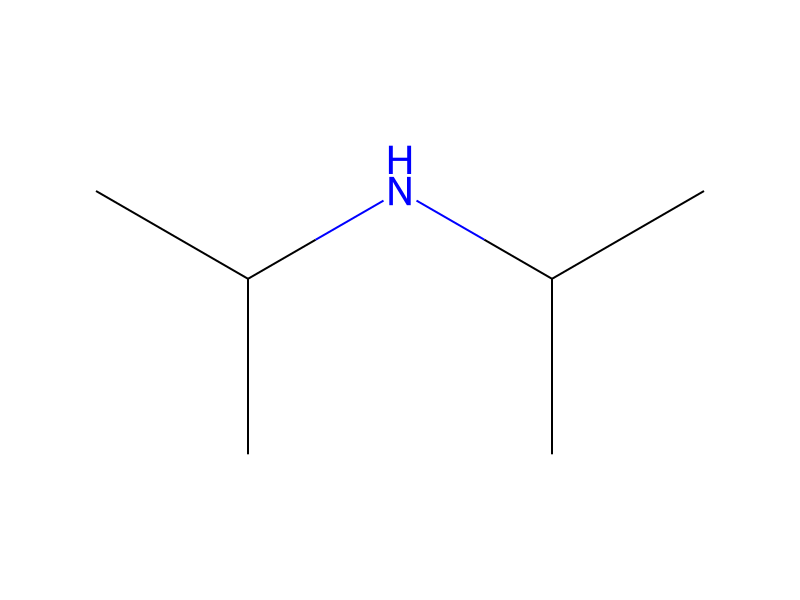
Diisopropylamine → HCID 86585362, compound
Reaction equation

[CH:1](NC(C)C)(C)C.C([Li])CCC.[CH2:13]([O:15][C:16]([C:18]1([C@@H:21]2[CH2:25][N:24]([C@H:26]([C:28]3[CH:33]=[CH:32][CH:31]=[CH:30][CH:29]=3)[CH3:27])[C:23](=[O:34])[C@H:22]2[F:35])[CH2:20][CH2:19]1)=[O:17])[CH3:14].C(C1C=CC=C(C(C)(C)C)C=1O)(C)(C)C.[Cl-].[NH4+]>O1CCCC1.CCCCCC>[CH2:13]([O:15][C:16]([C:18]1([C@H:21]2[CH2:25][N:24]([C@H:26]([C:28]3[CH:33]=[CH:32][CH:31]=[CH:30][CH:29]=3)[CH3:27])[C:23](=[O:34])[C@H:22]2[F:35])[CH2:20][CH2:1][CH2:19]1)=[O:17])[CH3:14]Participants




Conditions
- Temperature
- -78 °C
Procedure
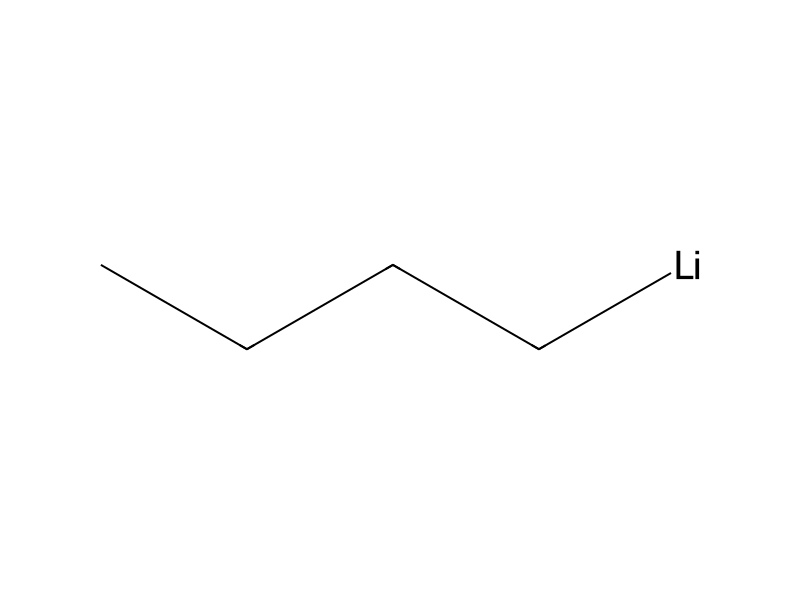
Under a nitrogen atmosphere, diisopropylamine (2.97 ml, 21.19 mmol) was dissolved in anhydrous tetrahydrofuran (30 ml) to which, after cooling to -78° C., was subsequently added dropwise an n-hexane solution of 1.63 M n-butyl lithium (10.8 ml, 17.60 mmol) over a period of 5 minutes. After 15 minutes of stirring at 0° C., the reaction solution was cooled to -78° C. and trans 4-(1-ethoxycarbonylcyclopropyl)-3-fluoro-1-[1-(S)-phenylethyl]-2-pyrrolidone (optical isomer B; 4.71 g, 14.13 mmol) dissolved in anhydrous tetrahydrofuran (30 ml) was added dropwise thereto over a period of 5 minutes. The reaction solution was stirred at -78° C. for 3 minutes, and 2,6-di-tert-butylphenol (4.37 g, 21.18 mmol) dissolved in anhydrous tetrahydrofuran (40 ml) was added dropwise thereto over a period of 5 minutes. The reaction solution was stirred at -78° C. for 10 minutes, mixed with saturated ammonium chloride aqueous solution (200 ml) and then warmed to room temperature. The organic layer was separated and then the aqueous layer was extracted with chloroform (100 ml×2). The organic layers were combined, washed with water (100 ml×2) and then dried over anhydrous sodium sulfate. After filtration, the filtrate was concentrated under reduced pressure, and the resulting residue was subjected to silica gel column chromatography and eluted with an eluant of n-hexane:ethyl acetate=2:1 to recover 1.96 g (42%) of the starting material and then with n-hexane:ethyl acetate=3:2 to obtain 1.79 g (38%) of the title compound in an oily form.
Workup
- temperaturethe reaction solution was cooled to -78° C.
- additionwas added dropwise
- waitover a period of 5 minutes
- stirringThe reaction solution was stirred at -78° C. for 3 minutes
- additionwas added dropwise
- waitover a period of 5 minutes
- stirringThe reaction solution was stirred at -78° C. for 10 minutes
- temperaturewarmed to room temperature
- customThe organic layer was separated
- extractionthe aqueous layer was extracted with chloroform (100 ml×2)
- washwashed with water (100 ml×2)
- dry with materialdried over anhydrous sodium sulfate
- filtrationAfter filtration
- concentrationthe filtrate was concentrated under reduced pressure
- washeluted with an eluant of n-hexane
- customethyl acetate=2:1 to recover 1.96 g (42%) of the starting material
Sources & evidence
- ORD record
- ord-9a1cad4cae31408dbce0fd8ba8e867af
- Source dataset
- uspto-grants-2000_09
- Patent number
- US06121285
Reaction notes
Loading…