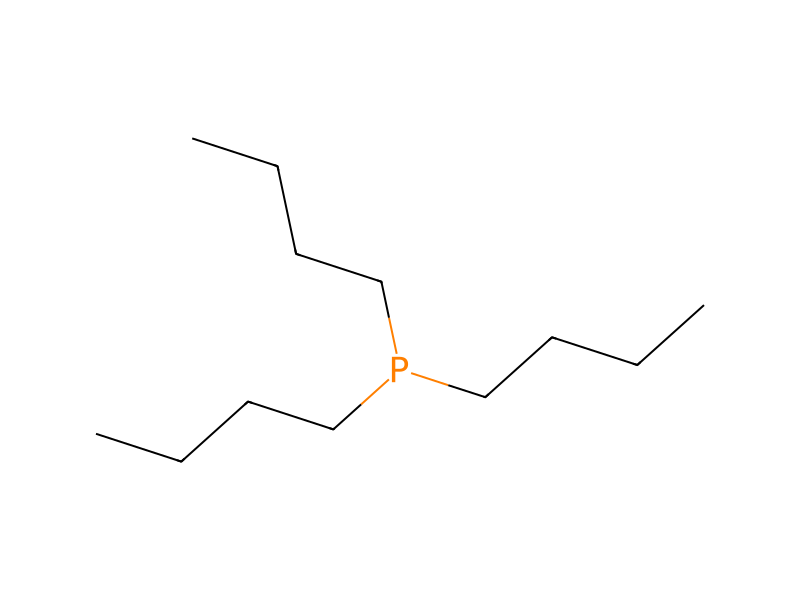
Tributylphosphine → HCID 68611425, compound
Reaction equation

N(C(N1CCCCC1)=O)=NC(N1CCCCC1)=O.C(P(CCCC)CCCC)CCC.[C:32]([O:36][C:37]([N:39]1[CH2:42][CH2:41][C@@H:40]1[CH2:43][OH:44])=[O:38])([CH3:35])([CH3:34])[CH3:33].[O:45]([CH2:52][C@H:53]1[CH2:55][C@@H:54]1[C:56]1[CH:57]=[C:58](O)[CH:59]=[N:60][CH:61]=1)[C:46]1[CH:51]=[CH:50][CH:49]=[CH:48][CH:47]=1>C1(C)C=CC=CC=1>[C:32]([O:36][C:37]([N:39]1[CH2:42][CH2:41][C@H:40]1[CH2:43][O:44][C:58]1[CH:59]=[N:60][CH:61]=[C:56]([C@H:54]2[CH2:55][C@@H:53]2[CH2:52][O:45][C:46]2[CH:51]=[CH:50][CH:49]=[CH:48][CH:47]=2)[CH:57]=1)=[O:38])([CH3:35])([CH3:34])[CH3:33]Participants


Procedure
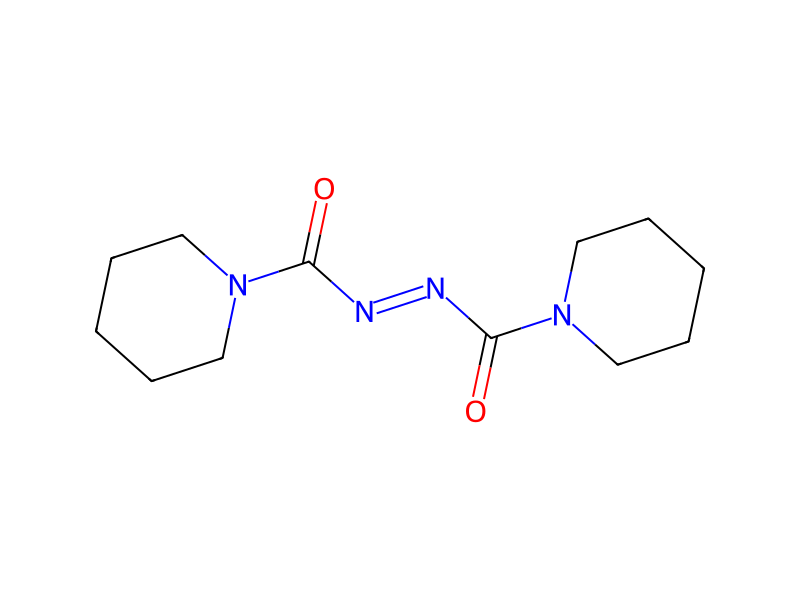
1,1′-(Azodicarbonyl)dipiperidine (253 mg, 1.00 mmol, 1.6 equiv.) was dissolved in 2 mL of toluene (dried with molecular sieve 3 Å) in a 10 mL round-bottom flask with a side neck. The flask was equipped with rubber septa and a magnetic stirrer. The atmosphere was exchanged with Ar (3 times), and the flask was cooled with an ice bath. Tributylphosphine (248 μL, 1.00 mmol, 1.6 equiv.) was added dropwise. The mixture was warmed to room temperature and was stirred for 10 min. The colorless solution formed was then transferred dropwise (via syringe) with ice cooling into a solution of 1-(tert-butoxycarbonyl)-2(R)-azetidinylmethanol (188 mg, 1.00 mmol, 1.6 equiv.) and 5-[(1S,2S)-2-(phenoxymethyl)cyclopropyl]-3-pyridinol (151 mg, 626 μmol) in a 25 mL round-bottom flask equipped with a magnetic stirrer and an Ar balloon. The resulting mixture was warmed to room temperature and stirred overnight. The reaction was quenched by addition of saturated NaHCO3 solution (10 mL). The product was extracted into EtOAc (3×20 mL). The combined organic phases were dried over Na2SO4 and concentrated. The residue was purified by CC on SiO2 (25×2.5 cm, EtOAc/hexanes 2:1) to give the crude product, which was further purified by preparative HPLC in two portions (Supelco Discovery C18, 250×21.2 mm, 5 μm particle size, UV detection at 270 nm, flow rate 12.5 mL/min, gradient from 20% to 100% CH3CN in water within 40 min, then 100% CH3CN for 20 min; tR 29.1-31.1 min) to give the title compound as a yellowish oil (231 mg, 90%). MS (EI) m/z 410 (M+, 0.1%).
Workup
- customThe flask was equipped with rubber septa and a magnetic stirrer
- temperaturethe flask was cooled with an ice bath
- customThe colorless solution formed
- customequipped with a magnetic stirrer
- temperatureThe resulting mixture was warmed to room temperature
- stirringstirred overnight
- customThe reaction was quenched by addition of saturated NaHCO3 solution (10 mL)
- extractionThe product was extracted into EtOAc (3×20 mL)
- dry with materialThe combined organic phases were dried over Na2SO4
- concentrationconcentrated
- customThe residue was purified by CC on SiO2 (25×2.5 cm, EtOAc/hexanes 2:1)
- customto give the crude product, which
- customwas further purified by preparative HPLC in two portions (Supelco Discovery C18, 250×21.2 mm, 5 μm particle size, UV detection at 270 nm, flow rate 12.5 mL/min, gradient from 20% to 100% CH3CN in water within 40 min
- wait100% CH3CN for 20 min
Sources & evidence
- ORD record
- ord-a49ce2be1006405aa82699c84ae42126
- Source dataset
- uspto-grants-2013_05
- Patent number
- US08445684B2
Reaction notes
Loading…