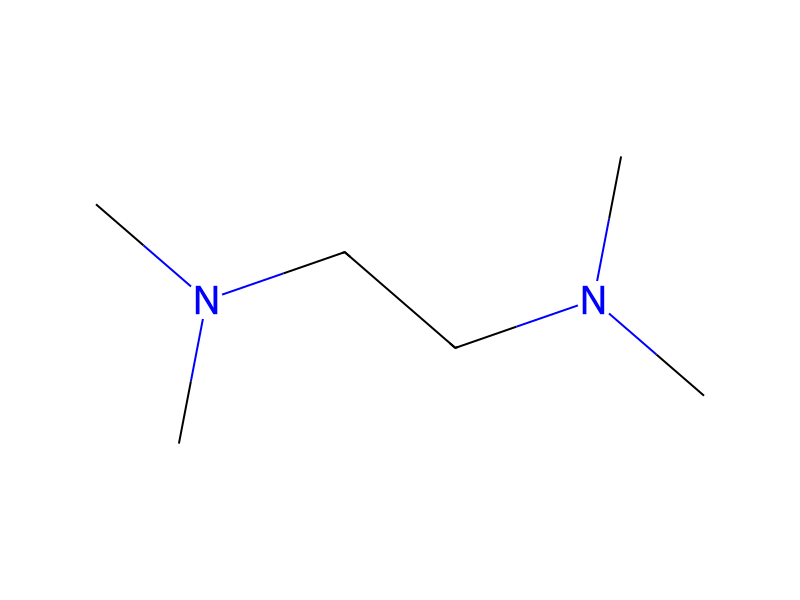
N,N,N',N'-Tetramethylethylenediamine → HCID 11808544, compound
Reaction equation

CN([CH2:4][CH2:5]N(C)C)C.[CH2:9]([S:16][C:17]#[CH:18])[C:10]1[CH:15]=[CH:14][CH:13]=[CH:12][CH:11]=1>Cl[Cu].CC(C)=O>[CH2:9]([S:16][C:17]#[C:18][C:10]#[C:9][S:16][CH2:17][C:5]1[CH:4]=[CH:14][CH:13]=[CH:12][CH:11]=1)[C:10]1[CH:15]=[CH:14][CH:13]=[CH:12][CH:11]=1Participants
Reactant 2

Product 1

Reagents, catalysts & solvents 2

Procedure
To a solution of trimethylsilylacetylene (10 mL, 71 mmol) in ether (100 mL) under argon at -78° C. was slowly added n-BuLi (2.5 M in hexanes, 28.3 mL, 71 mmol). The solution was warmed to -20° C. and then cooled to -70° C. Sulfur powder (2.3 g, 72 mmol), in a flask connected to the reaction vessel by Gooch tubing, was poured into the reaction mixture. The mixture was warmed to 10° C., becoming a pale yellow solution. Benzyl bromide (8.5 mL, 72 mmol) was added to quench the reaction. The resulting solution was stirred overnight. The solution was quenched by addition of saturated NH4Cl solution (30 mL), and the mixture was extracted with ether (3×50 mL), the combined ether layers washed with brine (1×30 mL) and water (1×30 mL). The organic phase was dried (MgSO4) and concentrated in vacuo affording a yellow oil (16.5 g; 100%): GC EI-MS: 220 (M+). Without further purification a solution of this yellow oil (16.5 g, 72 mmol) in THF (150 mL) was treated with water (10 mL) followed by THF solution (100 mL) of tetra-n-butylammonium fluoride (TBAF; 1.1 eq). The resulting reddish brown solution was stirred for 2.5 h, and then washed with brine (3×50 mL), dried (MgSO4, concentrated in vacuo, and distilled (39°-42° C./0.015 mmHg) affording benzylthioacetylene (2) as a yellow oil (10 g, 96%): 1H NMR (acetone-d6) δ 7.5-7.25 (m, 5 H) 4.05 (s, 2 H) 3.38 (s, 1 H); 13C NMR (acetone-d6 δ 137.89, 129.93, 129.38, 128.56, 85.43, 74.82, 40.07; IR 2041, 3286 cm-1. A mixture of CuCl (0.27 g), TMEDA (0.41 mL) and acetone (40 mL) were placed in a three-necked flask, and with stirring O2 was bubbled through the dark green solution for a few minutes. Compound 2 (8 g, 60 mmol) was added dropwise to the stirred green solution which was maintained at room temperature with a water bath. After 3 h, acetone was removed in vacuo and ice cold HCl solution (0.3 mL conc HCl diluted to 50 mL) was added to the concentrate. The suspension was extracted with ether (1×100, 2×50 mL). The combined ether layers were washed with brine (2×50 mL) and water (1×35 mL), dried (MgSO4), and concentrated in vacuo affording 3 as a yellow solid (7.7 g, 97%).
Workup
- customwere placed in a three-necked flask
- customwith stirring O2 was bubbled through the dark green solution for a few minutes
- customacetone was removed in vacuo and ice cold HCl solution (0.3 mL conc HCl diluted to 50 mL)
- additionwas added to the concentrate
- extractionThe suspension was extracted with ether (1×100, 2×50 mL)
- washThe combined ether layers were washed with brine (2×50 mL) and water (1×35 mL)
- dry with materialdried (MgSO4)
- concentrationconcentrated in vacuo
Sources & evidence
- ORD record
- ord-af039159f4254771a888142cd1a13320
- Source dataset
- uspto-grants-1995_12
- Patent number
- US05478959
Reaction notes
Loading…