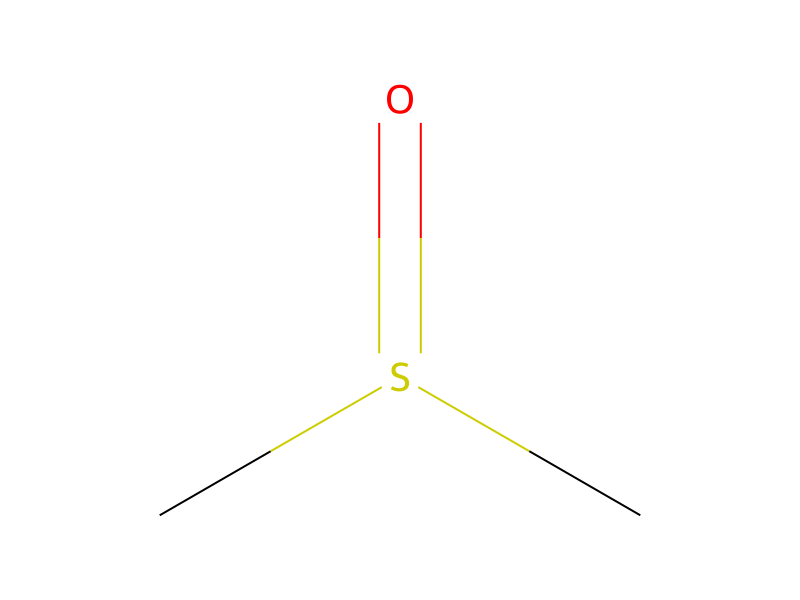
Dimethyl Sulfoxide → HCID 14179741, compound
Reaction equation

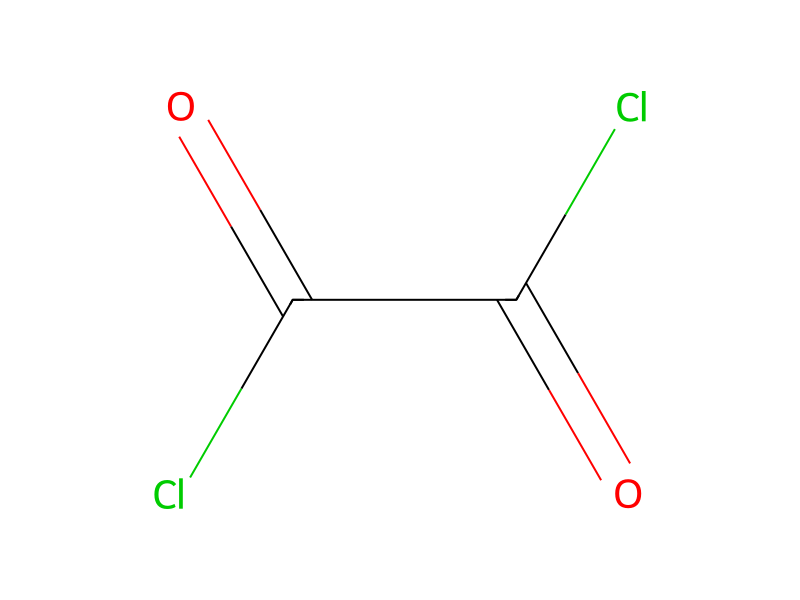
[C:1](Cl)(=O)[C:2](Cl)=O.[CH3:7]S(C)=O.[CH2:11]([O:18][CH:19]([CH2:22][O:23][CH2:24][CH2:25][CH2:26][CH2:27][CH2:28][CH2:29][CH2:30][CH2:31][CH2:32][CH2:33][CH2:34][CH2:35][CH2:36][CH2:37][CH2:38][CH3:39])[CH2:20][OH:21])[C:12]1[CH:17]=[CH:16][CH:15]=[CH:14][CH:13]=1.O>C(Cl)Cl>[CH2:11]([O:18][CH:19]([CH:20]([OH:21])[CH:1]([CH3:2])[CH3:7])[CH2:22][O:23][CH2:24][CH2:25][CH2:26][CH2:27][CH2:28][CH2:29][CH2:30][CH2:31][CH2:32][CH2:33][CH2:34][CH2:35][CH2:36][CH2:37][CH2:38][CH3:39])[C:12]1[CH:17]=[CH:16][CH:15]=[CH:14][CH:13]=1Participants
Reactant 4

Product 1

Reagents, catalysts & solvents 1
Conditions
- Temperature
- -50 °C
Procedure
To a solution of 3.05 g of oxalyl chloride in 40 ml of dry methylene chloride chilled with a dry ice- acetone bath was portionwise added under nitrogen atmosphere a solution of 3.75 g of dimethylsulfoxide in 10 ml of methylene chloride. The above addition was carried out keeping the oxalyl chloride solution at a temperature (inner temperature) below -60° C. To this solution was added a solution of 3.25 g of 2-benzyloxy-3-hexadecyloxy-1-propanol in 20 ml of methylene chloride, and then the dry ice-acetone bath was removed to allow the reaction solution to reach -10° C. The reaction solution was again chilled to -50° C. and, after addition of 5.28 g of triethylamine, allowed to reach 25° C. The reaction liquid was poured into 100 ml of water, and the separated organic layer was taken out. The organic layer was washed successively with 1N hydrochloric acid and aqueous saturated sodium hydrogencarbonate solution, and dried over anhydrous sodium sulfate. The solvent was removed under reduced pressure, and the residue was dissolved in 40 ml of dry tetrahydrofuran. Under nitrogen atmosphere, 8.0 ml of 2.0M isopropyl magnesium chloride-containing tetrahydrofuran was portionwise added to the above solution, keeping the inner temperature of the solution below 8° C. by chilling the solution with ice. The mixture was stirred overnight at room temperature. To the mixture (under chilling with ice) was portionwise added 16 ml of 1N hydrochloric acid, and further added ether. The organic layer was separated. The aqueous layer was then extracted with ether. The organic layer and the ether extract were combined, washed successively with an aqueous saturated sodium hydrogencarbonate solution and an aqueous saturated sodium chloride solution, and dried over anhydrous sodium sulfate. The solvent was distilled off, and the residue was purified by chromatography using silica gel (eluent: n-hexane/ethyl acetate=4/1) to give 2.16 g of the desired compound (mixture of isomers).
Workup
- additionThe above addition
- customat a temperature (inner temperature) below -60° C
- customthe dry ice-acetone bath was removed
- customto reach -10° C
- additionafter addition of 5.28 g of triethylamine
- customto reach 25° C
- customThe reaction liquid
- washThe organic layer was washed successively with 1N hydrochloric acid and aqueous saturated sodium hydrogencarbonate solution
- dry with materialdried over anhydrous sodium sulfate
- customThe solvent was removed under reduced pressure
- dissolutionthe residue was dissolved in 40 ml of dry tetrahydrofuran
- additionUnder nitrogen atmosphere, 8.0 ml of 2.0M isopropyl magnesium chloride-containing tetrahydrofuran
- additionwas portionwise added to the above solution
- customthe inner temperature of the solution below 8° C.
- temperatureby chilling the solution with ice
- temperatureTo the mixture (under chilling with ice)
- customThe organic layer was separated
- extractionThe aqueous layer was then extracted with ether
- extractionThe organic layer and the ether extract
- washwashed successively with an aqueous saturated sodium hydrogencarbonate solution
- dry with materialan aqueous saturated sodium chloride solution, and dried over anhydrous sodium sulfate
- distillationThe solvent was distilled off
- customthe residue was purified by chromatography
Sources & evidence
- ORD record
- ord-f8220b0147424f0a94a0bad99197830f
- Source dataset
- uspto-grants-1991_10
- Patent number
- US05053402
Reaction notes
Loading…