Water → 4-[4-(Chlorosulfonyl)phenoxy]-2-butynyl acetate
Reaction equation

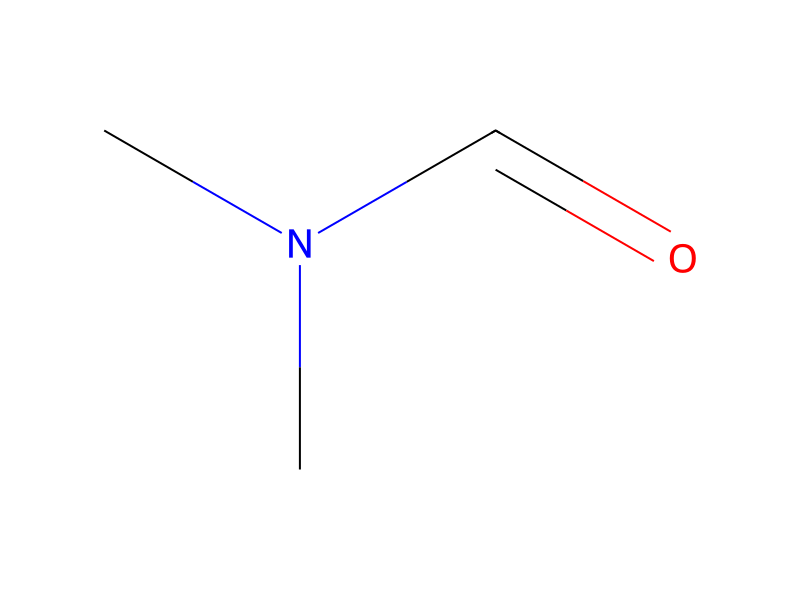
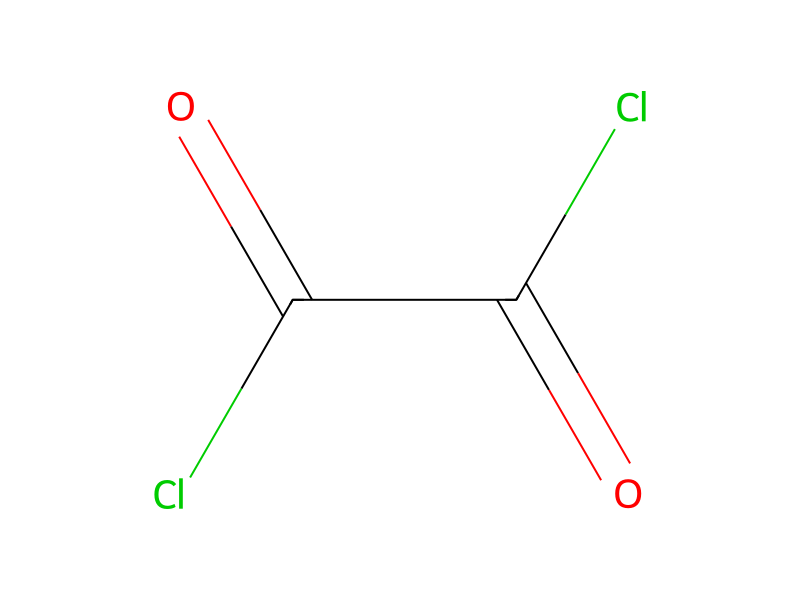
[C:1]([O:4][CH2:5][C:6]#[C:7][CH2:8][O:9][C:10]1[CH:15]=[CH:14][C:13]([S:16]([OH:19])(=O)=[O:17])=[CH:12][CH:11]=1)(=[O:3])[CH3:2].[Na].CN(C)C=O.O.C(Cl)(=O)C([Cl:30])=O>ClCCl.CCCCCCC>[C:1]([O:4][CH2:5][C:6]#[C:7][CH2:8][O:9][C:10]1[CH:15]=[CH:14][C:13]([S:16]([Cl:30])(=[O:19])=[O:17])=[CH:12][CH:11]=1)(=[O:3])[CH3:2]Participants
Reactant 5


Product 1

Reagents, catalysts & solvents 2

Conditions
- Temperature
- 12 °C
Procedure
To a suspension of 4-(4-acetoxy-but-2-ynyloxy)-benzenesulfonic acid, sodium salt from Example 6 (360 g, 1.18 mol) in 2.1 L of dichloromethane in a 5-L 4-neck flask, was added N,N-dimethylformamide (14 g, 0.19 mol) and water (4.5 g, 0.25 mol). The mixture was cooled to 12° C. Oxalyl chloride (165 mL, 240 g, 1.89 mol) was added over 70 min while the temperature was maintained at 10–20° C. The mixture was slowly warmed to 23° C. over 1 h and stirred at this temperature for 1 h. The reaction was monitored for completion by HPLC (S.M. <3%). The mixture was quenched into 2 L of water in a 12-L flask at 30° C. After 5 min of stirring, the phases were separated and the organic phase was washed with water (1 L). The mixture was held at 0–5° C. overnight, resulting in clear phase separation. Dichloromethane was distilled off to a residual volume of about 700 mL. t-Butyl methyl ether (1.5 L) was added. Distillation was continued to a residual volume of 850 mL. The solution was cooled to 18° C. and seeded with crystals. After 35 min of stirring, product crystals formed. Heptane (650 mL) was added over 40 min while the temperature was maintained at 15–20° C. The mixture was heated to 35° C. to dissolve the product before it was cooled back to 20° C. More heptane (350 mL) was added over 10 min. The mixture was stirred at 15–20° C. for 40 min. The crystals were filtered in a funnel and washed with heptane (400 mL). The product was dried in vacuo at 23° C. to a constant weight (316 g, 89% yield). 1H NMR (300 MHz, CDCl3) δ 8.01 (m, 2H), 7.12 (m, 2H), 4.86 (t, J=1.8 Hz, 2H), 4.72 (t, J=1.8 Hz, 2H), 2.10 (s, 3H); 13C NMR (75 MHz, CDCl3) ppm 170.5, 163.0, 137.2, 129.9, 116.0, 83.3, 80.2, 56.8, 52.3, 21.0.
Workup
- temperaturewas maintained at 10–20° C
- temperatureThe mixture was slowly warmed to 23° C. over 1 h
- customThe mixture was quenched into 2 L of water in a 12-L flask at 30° C
- stirringAfter 5 min of stirring
- customthe phases were separated
- washthe organic phase was washed with water (1 L)
- waitThe mixture was held at 0–5° C. overnight
- customresulting in clear phase separation
- distillationDichloromethane was distilled off to a residual volume of about 700 mL
- additiont-Butyl methyl ether (1.5 L) was added
- distillationDistillation
- temperatureThe solution was cooled to 18° C.
- stirringAfter 35 min of stirring
- customproduct crystals formed
- temperaturewas maintained at 15–20° C
- temperatureThe mixture was heated to 35° C.
- dissolutionto dissolve the product before it
- temperaturewas cooled back to 20° C
- stirringThe mixture was stirred at 15–20° C. for 40 min
- filtrationThe crystals were filtered in a funnel
- washwashed with heptane (400 mL)
- customThe product was dried in vacuo at 23° C. to a constant weight (316 g, 89% yield)
Sources & evidence
- ORD record
- ord-f2708ca6b50049c98338dc0515eb31fa
- Source dataset
- uspto-grants-2007_02
- Patent number
- US07179911B2
Reaction notes
Loading…