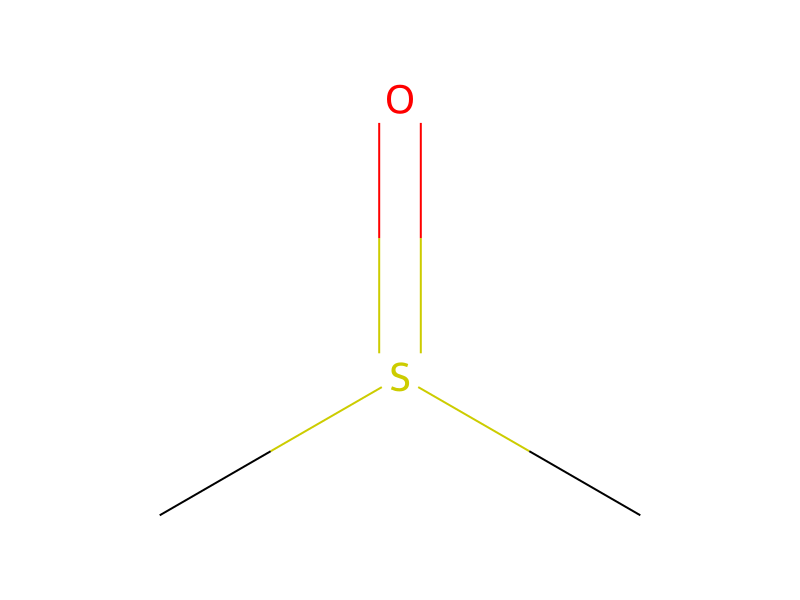
Dimethyl Sulfoxide → HCID 87822476, compound
Reaction equation

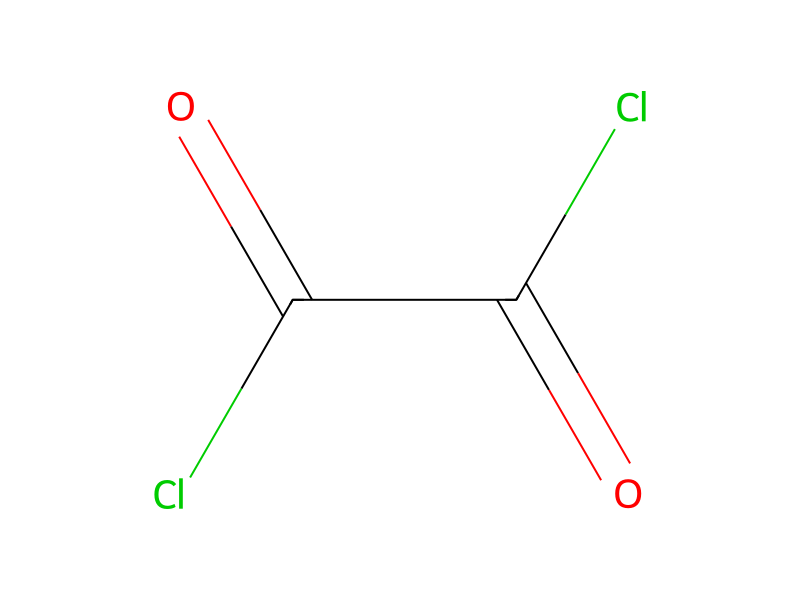
C(Cl)(=O)C(Cl)=O.C(=O)=O.C(Cl)(Cl)Cl.CS(C)=O.[C:18]([O:22][C:23]([N:25]([CH2:32][C:33]1[CH:38]=[CH:37][CH:36]=[C:35]([Br:39])[N:34]=1)[CH:26]([CH:29]([CH3:31])[CH3:30])[CH2:27][OH:28])=[O:24])([CH3:21])([CH3:20])[CH3:19]>C(Cl)Cl>[Br:39][C:35]1[N:34]=[C:33]([CH2:32][N:25]([CH:26]([CH:27]=[O:28])[CH:29]([CH3:30])[CH3:31])[C:23](=[O:24])[O:22][C:18]([CH3:19])([CH3:21])[CH3:20])[CH:38]=[CH:37][CH:36]=1Participants
Reactant 4


Product 1

Reagents, catalysts & solvents 1

Procedure
To a flame-dried flask was added oxalyl chloride (70 μL) followed by the addition of dry CH2Cl2 (2 mL). The resulting colorless solution was brought to −63° C. (dry ice/CHCl3) and a solution of DMSO (70 μL) in 0.5 mL dry CH2Cl2 was slowly added drop wise. The (tert-butoxy)-N-[(6-bromo(2-pyridyl))methyl]-N-[2-hydroxy-1-(methylethyl)ethyl]carboxamide (0.19 g, 0.51 mmol), previously dissolved in dry CH2Cl2 (2 mL), was added slowly dropwise. The resulting mixture was kept at −63° C., and stirred for 30 min followed by the slowly addition of a solution of TEA (0.3 mL) in dry CH2Cl2 (1 mL). The mixture was stirred at −63° C. until all the starting material was consumed (checked by MS)(1.5 h). The mixture was brought to −20° C., quenched with a saturated solution of NH4Cl (15 mL) and diluted with EtOAc (35 mL). The organic phase was separated and the aqueous phase was extracted with EtOAc (3×30 mL). The organic layers were combined, dried over MgSO4, filtered and concentrated by rotary evaporation without further purification to afford the title compound as a yellow-semi solid. EI-MS m/z 371 (M+H).
Workup
- customwas brought to −63° C.
- additionwas added slowly dropwise
- customwas kept at −63° C.
- stirringThe mixture was stirred at −63° C. until all the starting material
- customwas consumed (checked by MS)(1.5 h)
- customwas brought to −20° C.
- customquenched with a saturated solution of NH4Cl (15 mL)
- additiondiluted with EtOAc (35 mL)
- customThe organic phase was separated
- extractionthe aqueous phase was extracted with EtOAc (3×30 mL)
- dry with materialdried over MgSO4
- filtrationfiltered
- concentrationconcentrated by rotary evaporation without further purification
Sources & evidence
- ORD record
- ord-c226926615614cd8acedec433574f631
- Source dataset
- uspto-grants-2007_03
- Patent number
- US07196104B2
Reaction notes
Loading…