Phenol → HCID 23643972, compound
Reaction equation

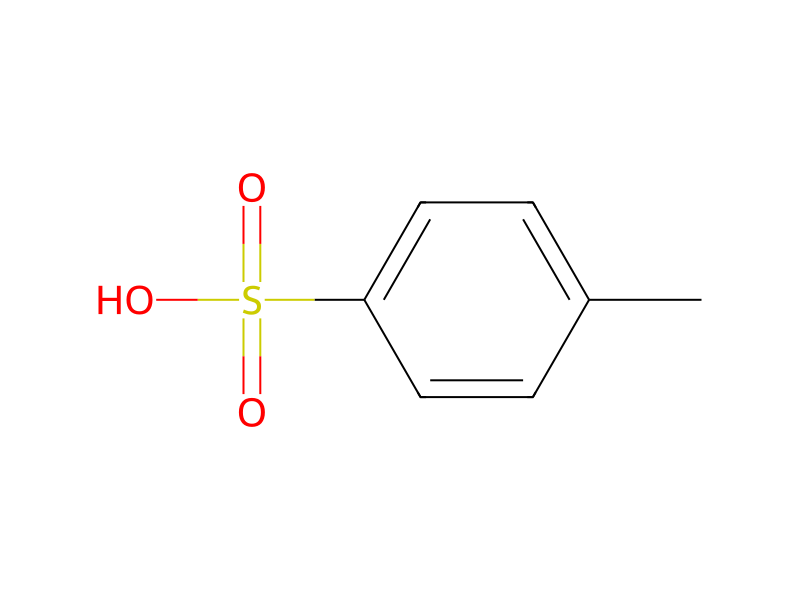
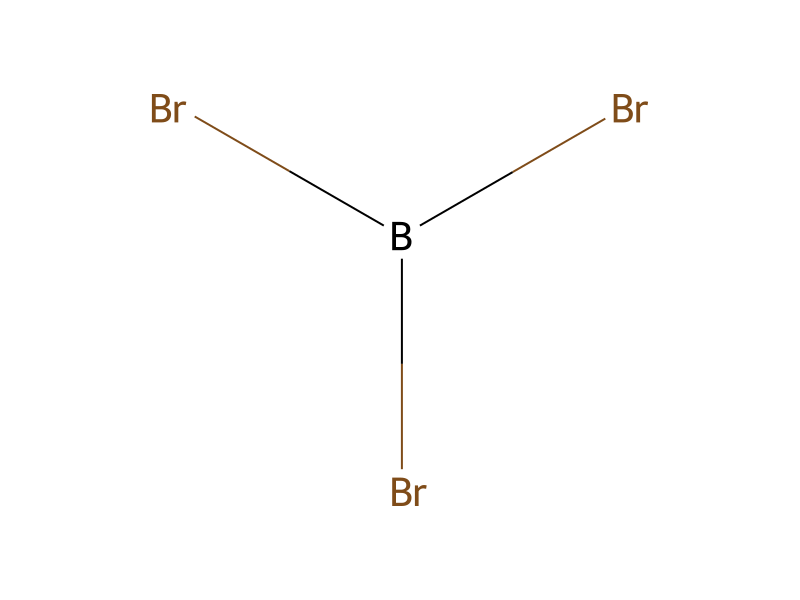
CC1C=CC(S(O)(=O)=[O:9])=CC=1.C[O:13][C:14]1[CH:19]=[CH:18][C:17](CS)=[CH:16][CH:15]=1.[CH:22]1[C:27](/[CH:28]=[C:29]2\[C@@H:30](C3C=C(O)C=C(O)C=3)[C@H:31](C3C=CC(O)=CC=3)[C:32]3[C:37]\2=[CH:36][C:35]([OH:38])=[CH:34][C:33]=3[OH:39])=[CH:26][CH:25]=[C:24]([OH:55])[CH:23]=1.[C:56]1([OH:62])[CH:61]=[CH:60][CH:59]=[CH:58][CH:57]=1.B(Br)(Br)Br>C(Cl)Cl>[CH:26]1[C:27]([CH2:28][C:29]2[C:37]3[C:32](=[C:33]([OH:39])[CH:34]=[C:35]([OH:38])[CH:36]=3)[C@H:31]([C:60]3[CH:61]=[C:56]([OH:62])[CH:57]=[C:58]([OH:9])[CH:59]=3)[C:30]=2[C:17]2[CH:16]=[CH:15][C:14]([OH:13])=[CH:19][CH:18]=2)=[CH:22][CH:23]=[C:24]([OH:55])[CH:25]=1Participants
Reactant 5



Product 1

Reagents, catalysts & solvents 1

Procedure
One of these compounds is biaryl alcohol 11 (FIG. 2), an adduct synthesized in 71% yield through an aldol reaction between the lithiated form of 9 (prepared in 4 steps in 80% overall yield from 3,5-dimethoxybenzaldehyde, (see Ref. 12) and 3,5-dimethoxybenzaldehyde (10). As shown in Scheme 1, when this key intermediate was treated with a stoichiometric amount of TFA under carefully controlled conditions (−30→—20° C.) in CH2Cl2, a cascade sequence featuring cation generation, regio- and stereoselective cyclization (in relative terms), and stereoselective cation capture, smoothly afforded intermediate 14 after 4 hours. A terminating quench under basic conditions (K2CO3, MeOH) then completed the one-pot synthesis of intermediate 15 from 9 in 75% yield, a compound that proved to be just two steps away from paucifloral F (7). Those operations, alcohol oxidation using Dess-Martin periodinane and BBr3-induced global demethylation in CH2Cl2 at 0° C., [13] proceeded smoothly in 84% overall yield. However, if 11 was exposed to a different proton source, one with a non-nucleophilic counterion such as that possessed by p-TsOH, then it proved possible to arrest the sequence at cation 13 prior to β-hydride elimination and access entirely different cyclic products. Indeed, if a nucleophile such as p-methoxy-α-toluenethiol was added at −30° C. after 11 had been exposed to p-TsOH for 5 hours and the reaction media was then concentrated to near dryness, sulfide 16 was obtained in 57% overall yield. Sulfide 16 could also be accessed in 82% yield from alcohol 15 upon its treatment with p-TsOH and p-methoxy-α-toluenethiol in CH2Cl2 at 25° C. This new tetra-aryl intermediate could then be converted into the natural product ampelopsin D (2) through a highly selective Ramberg-Backlund reaction [14] under Meyer's modified conditions [15] that afforded permethylated ampelopsin D along with its chromatographically separable Z-olefin isomer in a 5:1 ratio (40% and 7% yield over two steps, respectively), followed by Lewis acid-mediated phenol deprotection using BBr3. This final deprotection step produced a 5:1 mixture of both ampelopsin D (2) and isoampelopsin D (17) which were obtained in pure form in near quantitative yield by treating the product mixture with Ac2O, chromatographically separating the resultant acetates, and using KCN in MeOH to then effect ester hydrolysis. Subsequent treatment of 2 with 5 equivalents of HCl in MeOH at 80° C. then provided the means to effect, in near quantitative yield, the olefin isomerization necessary to access isoampelopsin D (17) [16].
Workup
- customafforded
- customThis final deprotection step produced
Sources & evidence
- ORD record
- ord-3794d2c02b9e4462820b338e495c0307
- Source dataset
- uspto-grants-2015_11
- Patent number
- US09185909B2
Reaction notes
Loading…