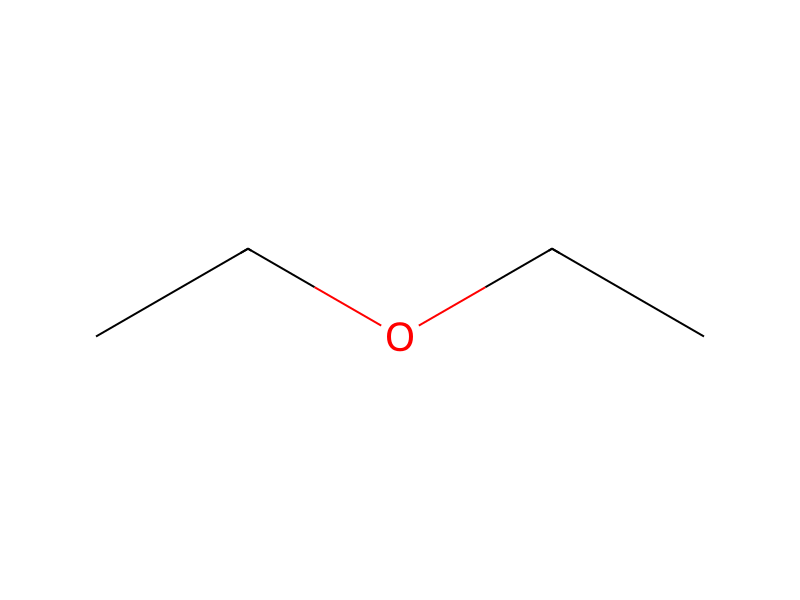
Diethyl Ether → 660417-38-9
Reaction equation

[Cl:1][C:2]1[CH:3]=[C:4]([C:8]2[O:12][N:11]=[C:10]([CH:13](O)O)[N:9]=2)[CH:5]=[CH:6][CH:7]=1.C(O[CH2:19][CH3:20])C>Cl>[Cl-:1].[Cl:1][C:2]1[CH:3]=[C:4]([C:8]2[O:12][N:11]=[C:10]([CH2:13][N+:9]3[CH:20]=[CH:19][CH:3]=[CH:4][CH:8]=3)[N:9]=2)[CH:5]=[CH:6][CH:7]=1Participants
Reactant 2

Product 1

Reagents, catalysts & solvents 1
Procedure
The title compound was synthesized according to the method described in Palazzo et al. J. Heterocycl. Chem. (1979) 16:1469. 1-[5-(3-Chloro-phenyl)-[1,2,4]oxadiazol-3-ylmethyl]-pyridinium chloride (1.81 g, 5.87 mmol) was dissolved in water (20 ml). To this solution, 4-nitroso-N,N-dimethylanilin (0.88 g, 5.86 mmol) dissolved in ethanol (50 ml) was added, followed by slow addition of 1 molar aq. sodium hydroxide (5.9 ml, 5.9 mmol) over a 3 min period. After 1 h the formed precipitate was filtered, washed with water and air-dried to give the title compound (2.08 g, wet) which was used immediately in the next step MS (ESI) m/z 344 (M+1). Step 2: [5-(3-Chloro-phenyl)-[1,2,4]oxadiazol-3-yl]-methanediol: N-{4-[(Z)-{[5-(3-Chlorophenyl)-1,2,4-oxadiazol-3-yl]methylene}(oxido)amino]phenyl}-N,N-dimethylamine (2.08 g wet) was suspended in diethyl ether (30 ml), followed by the addition of 1 molar aqueous hydrochloric acid. The mixture was stirred vigorously for 20 min, transferred to a separation funnel and diluted with diethyl ether and 1 molar aqueous hydrochloric acid. After extraction, the aqueous layer was extracted two more times with diethyl ether. Combining the organic layers, drying over magnesium sulfate, followed by evaporation to dryness and drying in vacuo gave the title compound as crude (0.56 g, 42% from 1-[5-(3-chloro-phenyl)-[1,2,4]oxadiazol-3-ylmethyl]-pyridinium chloride). MS (ESI) m/z 227 (M+1). Step 3: [5-(3-Chloro-phenyl)-[1,2,4]oxadiazol-3-yl]-methanol: Step 3: [5-(3-Chloro-phenyl)-[1,2,4]oxadiazol-3-yl]-methanol: 1-[5-(3-Chloro-phenyl)-[1,2,4]oxadiazol-3-ylmethyl]-pyridinium chloride (99.3 mg, 0.44 mmol) was dissolved in methanol (4 ml) followed by the addition of sodium borohydride (32 mg, 0.84 mmol). More sodium borohydride was added after 2 h and the reaction was allowed to run over night. The reaction mixture was diluted with dichloromethane and aq. ammonium chloride and stirred vigorously. After separation of the layers and washing of the organic layer with brine, followed by evaporation to dryness, crude product was obtained. This was purified by flash chromatography using heptane/ethyl acetate which gave the title compound (32.0 mg, 32%). 1H NMR (CDCl3), δ (ppm): 8.11 (s, 1H), 8.00 (apparent d, 1H), 7.56 (apparent d, 1H), 7.46 (apparent t, 1H), 4.87 (d, 2H), 2.91 (t, 1H).
Workup
- customtransferred to a separation funnel
- extractionAfter extraction
- extractionthe aqueous layer was extracted two more times with diethyl ether
- dry with materialdrying over magnesium sulfate
- customfollowed by evaporation to dryness
- customdrying in vacuo
Sources & evidence
- ORD record
- ord-fb1045f68c3845f58126112590461485
- Source dataset
- uspto-grants-2009_09
- Patent number
- US07585881B2
Reaction notes
Loading…