3-Oxo-azetidine-1-carboxylic acid tert-butyl ester → 1-(tert-Butoxycarbonyl)-3-(cyanomethylene)azetidine
Reaction equation

P(O[CH2:10][C:11]#[N:12])(OCC)(OCC)=O.CC(C)([O-])C.[K+].[C:19]([O:23][C:24]([N:26]1[CH2:29][C:28](=O)[CH2:27]1)=[O:25])([CH3:22])([CH3:21])[CH3:20]>O1CCCC1.O.[Cl-].[Na+]>[C:11]([CH:10]=[C:28]1[CH2:29][N:26]([C:24]([O:23][C:19]([CH3:22])([CH3:21])[CH3:20])=[O:25])[CH2:27]1)#[N:12]Participants
Reactant 3


Product 1

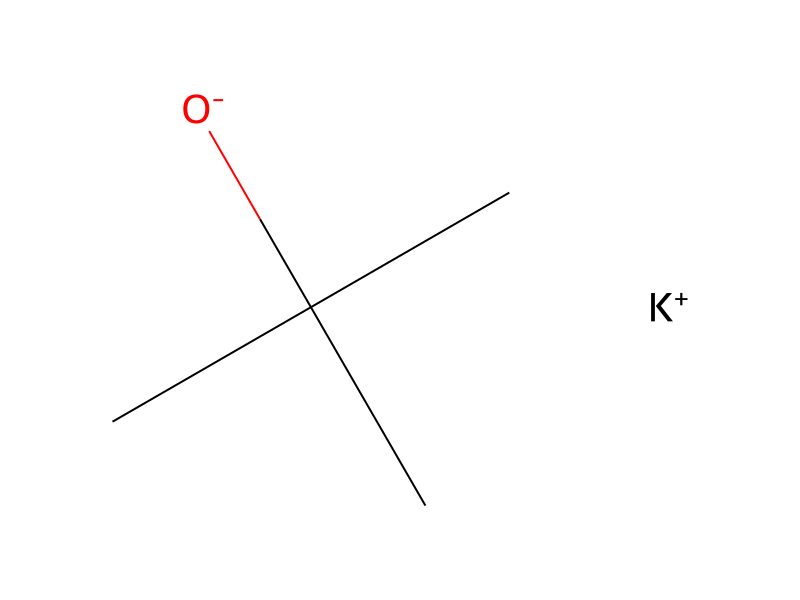
Reagents, catalysts & solvents 3


Conditions
- Temperature
- -14 °C
Procedure
Diethyl cyanomethyl phosphate (745 g, 4.20 mol, 1.20 equiv) and anhydrous tetrahydrofuran (THF, 9 L) were added to a four-neck flask equipped with a thermowell, an addition funnel and the nitrogen protection tube at room temperature. The solution was cooled with an ice-methanol bath to −14° C. and a 1.0 M solution of potassium tert-butoxide (t-BuOK) in anhydrous tetrahydrofuran (THF, 3.85 L, 3.85 mol, 1.1 equiv) was added over 20 min keeping the reaction temperature below −5° C. The resulting reaction mixture was stirred for 3 h at −10° C. and a solution of 1-tert-butoxycarbonyl-3-azetidinone (600 g, 3.50 mol) in anhydrous tetrahydrofuran (THF, 2 L) was added over 2 h keeping the internal temperature below −5° C. The reaction mixture was stirred at −5 to −10° C. over 1 h and then slowly warmed up to room temperature and stirred at room temperature for overnight. The reaction mixture was then diluted with water (4.5 L) and saturated aqueous sodium chloride solution (NaCl, 4.5 L) and extracted with ethyl acetate (EtOAc, 2×9 L). The combined organic layers were washed with brine (6 L) and dried over anhydrous sodium sulfate (Na2SO4). The solvent was removed under reduced pressure and the residue was diluted with dichloromethane (CH2Cl2, 4 L) before being absorbed onto silica gel (SiO2, 1.5 Kg). The crude product, which was absorbed on silica gel, was purified by flash column chromatography (SiO2, 3.5 Kg, 0-25% EtOAc/hexanes gradient elution) to afford tert-butyl 3-(cyanomethylene)azetidine-1-carboxylate (2, 414.7 g, 679.8 g theoretical, 61% yield) as a white solid. For 2: 1H NMR (300 MHz, CDCl3) δ 5.40 (m, 1H), 4.70 (m, 2H), 4.61 (m, 2H), 1.46 (s, 9H) ppm; C10H14N2O2 (MW, 194.23), LCMS (EI) m/e 217 (M++Na).
Workup
- customequipped with a thermowell, an addition funnel
- customat room temperature
- customthe reaction temperature below −5° C
- customThe resulting reaction mixture
- customthe internal temperature below −5° C
- stirringThe reaction mixture was stirred at −5 to −10° C. over 1 h
- temperatureslowly warmed up to room temperature
- stirringstirred at room temperature for overnight
- extractionextracted with ethyl acetate (EtOAc, 2×9 L)
- washThe combined organic layers were washed with brine (6 L)
- dry with materialdried over anhydrous sodium sulfate (Na2SO4)
- customThe solvent was removed under reduced pressure
- additionthe residue was diluted with dichloromethane (CH2Cl2, 4 L)
- custombefore being absorbed onto silica gel (SiO2, 1.5 Kg)
- customThe crude product, which was absorbed on silica gel
- customwas purified by flash column chromatography (SiO2, 3.5 Kg, 0-25% EtOAc/hexanes gradient elution)
Sources & evidence
- ORD record
- ord-f0efbe0b7efe48e994884a9f96741041
- Source dataset
- uspto-grants-2015_03
- Patent number
- US08987443B2
Reaction notes
Loading…