Sulfuryl chloride → HCID 23394558, compound
Reaction equation

S(Cl)(Cl)(=O)=O.C(SSC(C)C)(C)C.[CH:14]([S:17]Cl)([CH3:16])[CH3:15].[C:19]1([NH:29][C:30]2[CH:35]=[CH:34][C:33]([CH2:36][C@H:37]([NH:43][C:44]3[C:47]4([CH2:52][CH2:51][O:50][CH2:49][CH2:48]4)[C:46](=[O:53])[CH:45]=3)[C:38]([O:40][CH2:41][CH3:42])=[O:39])=[CH:32][CH:31]=2)[C:28]2[C:23](=[CH:24][CH:25]=[N:26][CH:27]=2)[CH:22]=[CH:21][N:20]=1>O1CCCC1>[CH:14]([S:17][C:45]1[C:46](=[O:53])[C:47]2([CH2:52][CH2:51][O:50][CH2:49][CH2:48]2)[C:44]=1[NH:43][C@@H:37]([CH2:36][C:33]1[CH:32]=[CH:31][C:30]([NH:29][C:19]2[C:28]3[C:23](=[CH:24][CH:25]=[N:26][CH:27]=3)[CH:22]=[CH:21][N:20]=2)=[CH:35][CH:34]=1)[C:38]([O:40][CH2:41][CH3:42])=[O:39])([CH3:16])[CH3:15]Participants
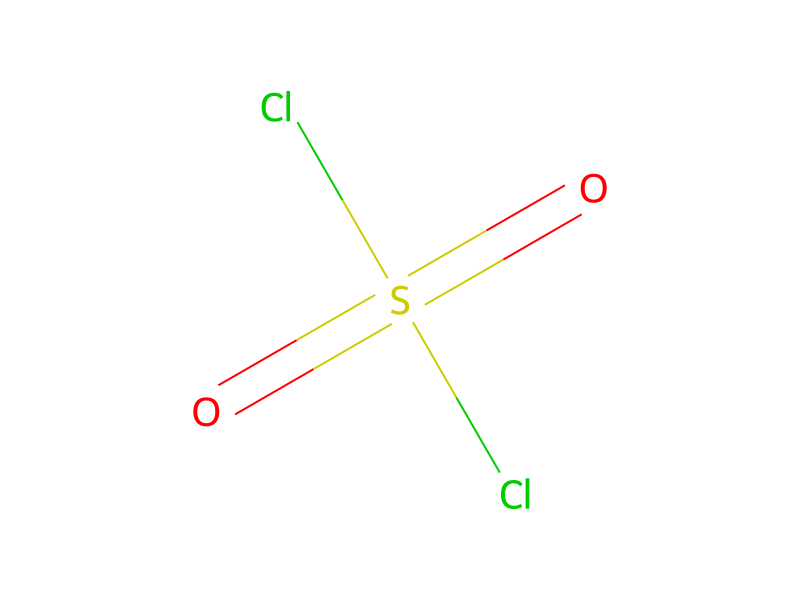



Procedure
Sulphuryl chloride (13.6 g) was added drop-wise to diisopropyl disulphide (25.0 g) in tetrahydrofuran (THF, 150 mL) at 0-5° C. under nitrogen. After stirring out for 30 minutes, the resulting isopropyl sulphenyl chloride preparation was introduced slowly, from a graduated dropping funnel, to a solution of Example 9 (30.0 g) in THF (500 mL), held at the same temperature. Conversion to product was complete after 85 mL of the stock solution had been charged. The reaction was quenched with 10% sodium bicarbonate solution (175 mL) and the layers separated. The aqueous phase was extracted with ethyl acetate (100 mL), and the combined organic solutions then washed with saturated brine (100 mL). The isolated organic phase was solvent swapped to ethanol (650 mL), by distillation at atmospheric pressure, from which the product crystallised on cooling. This suspension was filtered at 20° C. and the title compound washed with ethanol (2×30 mL) prior to drying in vacuo at 50-60° C. (26.9 g; 77.5% yield, off-white powder m.p. 221° C.). δH (DMSO d6, 390K) 9.83 (1H, s), 9.52 (1H, s), 8.94 (1H, d, J 9.5 Hz), 8.65 (1H, d, J 5.6 Hz), 8.15 (1H, d, J 5.7 Hz), 7.78 (2H, d, J 8.5 Hz), 7.68 (1H, d, J 5.6 Hz), 7.23 (2H, d, J 8.5 Hz), 7.12 (1H, d, J 5.7 Hz), 5.26 (1H, m), 4.19 (2H, q, J 7.1 Hz), 3.81-3.76 (2H, m), 3.64-3.55 (2H, m), 3.20 (1H, dd, J 13.8, 4.3 Hz), 2.96 (1H, dd, J 13.8, 10.3 Hz), 2.81-2.74 (1H, m), 2.06-1.93 (2H, m), 1.50-1.47 (1H, m), 1.32-1.28 (1H, m), 1.23 (3H, t, J 7.1 Hz), 1.07 (3H, d, J 6.6 Hz), 1.05 (3H, d, J 6.6 Hz); m/z (ES+, 70V) 547.2 (MH+).
Workup
- additionhad been charged
- customThe reaction was quenched with 10% sodium bicarbonate solution (175 mL)
- customthe layers separated
- extractionThe aqueous phase was extracted with ethyl acetate (100 mL)
- washthe combined organic solutions then washed with saturated brine (100 mL)
- distillationswapped to ethanol (650 mL), by distillation at atmospheric pressure, from which the product
- customcrystallised
- temperatureon cooling
- filtrationThis suspension was filtered at 20° C.
- washthe title compound washed with ethanol (2×30 mL)
- dry with materialto drying in vacuo at 50-60° C. (26.9 g; 77.5% yield, off-white powder m.p. 221° C.)
Sources & evidence
- ORD record
- ord-c3bff0e26e4a440b823b620e4dc18f38
- Source dataset
- uspto-grants-2005_10
- Patent number
- US06951943B2
Reaction notes
Loading…