Nitric Acid → Methyl (4-methoxy-3-nitrophenyl)acetate
Reaction equation

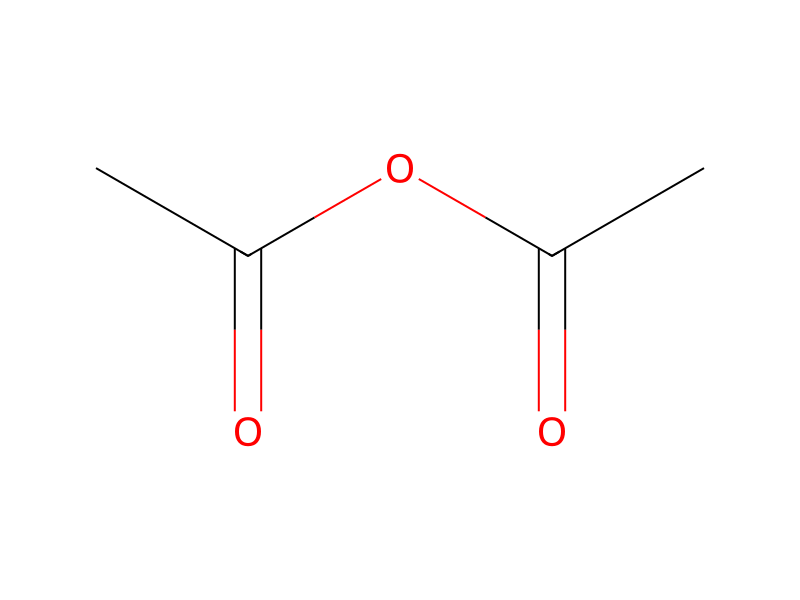
[C:1]([O:4][C:5](=[O:7])[CH3:6])(=O)C.[N+:8]([O-:11])(O)=[O:9].[Cl-].[Cl-].[Ca+2].[CH3:15][O:16][C:17]1[CH:22]=[CH:21][C:20](CC(O)=O)=[CH:19][CH:18]=1.[OH-].[Na+]>>[N+:8]([C:18]1[CH:19]=[C:20]([CH2:6][C:5]([O:4][CH3:1])=[O:7])[CH:21]=[CH:22][C:17]=1[O:16][CH3:15])([O-:11])=[O:9]Participants



Conditions
- Temperature
- -78 °C
Procedure
An oven-dried 2-L, 3-neck round bottom flask, equipped with a mechanical stir motor, a low-temperature thermometer and an equalizing dropping funnel, was charged with acetic anhydride (631 mL) and subsequently cooled to −78° C. Fuming nitric acid (Baker, 90%, 27 mL) was added dropwise via the dropping funnel protected with a drying tube filled with CaCl2. After addition was completed, the reaction temperature was allowed to warm to 20° C. over 1 h. The reaction mixture was cooled to −78° C. again and added 4-methoxyphenylacetic acid (50 g, 0.28 mol) dropwise via the dropping funnel. After stirring at −50° C. for 1 h., the reaction mixture was allowed to warm to −30° C. over 20 min. and then cooled to −50° C. again. The reaction mixture was quenched with H2O (500 mL) at −50° C. and warmed up to room temperature and stirred for 0.5 h. The reaction mixture was partitioned between CH2Cl2 (500 mL) and H2O. The aqueous layer was extracted with CH2Cl2 (3×500 mL). The combined CH2Cl2 extracts were concentrated in vacuo to give a yellow oil. This was added slowly to a 2 M solution of NaOH (2 L) cooled at 0° C. and stirred at room temperature overnight. The reaction mixture was partitioned between CH2Cl2 (500 mL) and H2O. The aqueous layer was extracted with CH2Cl2 (3×500 mL). The combined CH2Cl2 extracts were stirred with 2 M NaOH solution (1 L) for 1 h. The layers were separated and the organic layer was washed with H2O (500 mL), brine (500 mL), dried over Na2SO4 and filtered. The solvents were removed in vacuo to afford crude product as a light yellow solid (56 g). Purification by recrystallization from MeOH (600 mL) gave product. Yield 48 g (77%).
Workup
- customprotected with a drying tube
- additionAfter addition
- temperatureto warm to 20° C. over 1 h
- temperatureThe reaction mixture was cooled to −78° C. again
- temperatureto warm to −30° C. over 20 min.
- temperaturecooled to −50° C. again
- customThe reaction mixture was quenched with H2O (500 mL) at −50° C.
- temperaturewarmed up to room temperature
- stirringstirred for 0.5 h
- customThe reaction mixture was partitioned between CH2Cl2 (500 mL) and H2O
- extractionThe aqueous layer was extracted with CH2Cl2 (3×500 mL)
- concentrationThe combined CH2Cl2 extracts were concentrated in vacuo
- customto give a yellow oil
- temperaturecooled at 0° C.
- stirringstirred at room temperature overnight
- customThe reaction mixture was partitioned between CH2Cl2 (500 mL) and H2O
- extractionThe aqueous layer was extracted with CH2Cl2 (3×500 mL)
- stirringThe combined CH2Cl2 extracts were stirred with 2 M NaOH solution (1 L) for 1 h
- customThe layers were separated
- washthe organic layer was washed with H2O (500 mL), brine (500 mL)
- dry with materialdried over Na2SO4
- filtrationfiltered
- customThe solvents were removed in vacuo
Sources & evidence
- ORD record
- ord-7843942161f640df9b06d8df5e03577d
- Source dataset
- uspto-grants-2003_10
- Patent number
- US06630496B1
Reaction notes
Loading…