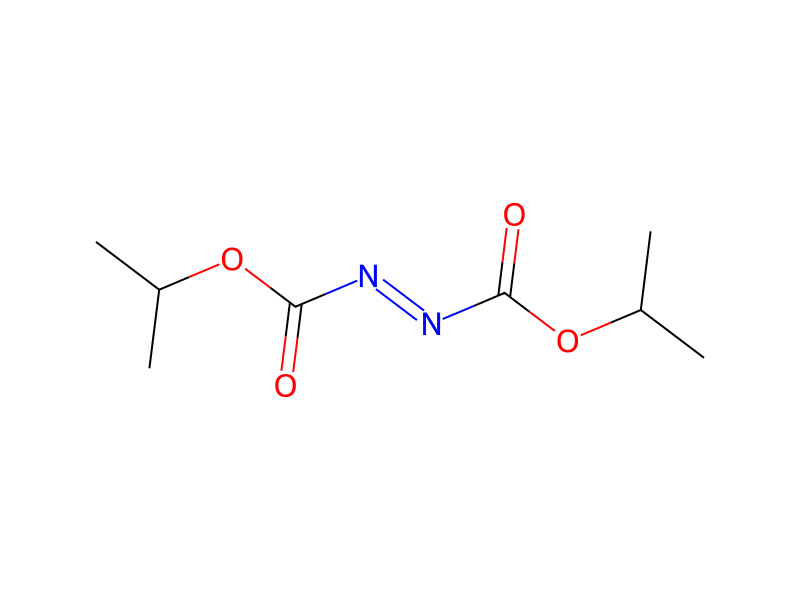
[Cl:1][C:2]1[CH:3]=[C:4]([CH2:9][N:10]2[C:14]3[CH:15](O)[CH2:16][CH2:17][CH2:18][CH2:19][C:13]=3[N:12]=[C:11]2[CH:21]([CH3:23])[CH3:22])[CH:5]=[CH:6][C:7]=1[Cl:8].[CH2:24]([O:26][C:27]([CH:29]([C:35]([O:37][CH2:38][CH3:39])=[O:36])[C:30]([O:32][CH2:33][CH3:34])=[O:31])=[O:28])[CH3:25].CP(C)C.N(C(OC(C)C)=O)=NC(OC(C)C)=O>C1(C)C=CC=CC=1.CC1CCCO1>[Cl:1][C:2]1[CH:3]=[C:4]([CH2:9][N:10]2[C:14]3[CH:15]([C:29]([C:35]([O:37][CH2:38][CH3:39])=[O:36])([C:27]([O:26][CH2:24][CH3:25])=[O:28])[C:30]([O:32][CH2:33][CH3:34])=[O:31])[CH2:16][CH2:17][CH2:18][CH2:19][C:13]=3[N:12]=[C:11]2[CH:21]([CH3:23])[CH3:22])[CH:5]=[CH:6][C:7]=1[Cl:8]To a suspension of Intermediate 8 (270 mg) and triethylmethanetricarboxylate (0.324 ml) in toluene (4 ml) and methyltetrahydrofuran (4.00 ml), stirred at RT under an inert atmosphere of nitrogen, was added trimethylphosphine, 1M in toluene (1.529 ml). The resulting solution was cooled to −78° C. and diisopropyl azodicarboxylate (0.301 ml) was added dropwise via syringe and the reaction was stirred at −78° C. for 45 mins. The reaction mixture was removed from the cold bath and allowed to warm to RT and stirred under N2 atmosphere overnight. Reaction mixture concentrated slowly in vacuo using the rotavapor with the water bath at 50° C. The residue was taken up in DCM (20 ml) and washed with water (20 mL). The 2 phases were separated; the aqueous was extracted again with DCM (20 mL). The organic extracts were combined, dried (hydrophobic frit) and concentrated under vacuum. The residue was purified by silica using a cyclohexane:ethyl acetate 0-100% gradient. The desired fractions were combined and concentrated under vacuum to give the title compound (435 mg).