HomeMCPWorkbenchSkills
HomeWorkbenchNotificationsMe
llms.txt · MCP GuidePrivacy Policy · Terms of Service · Supportv1.8.3
  1. Home
  2. /Reaction

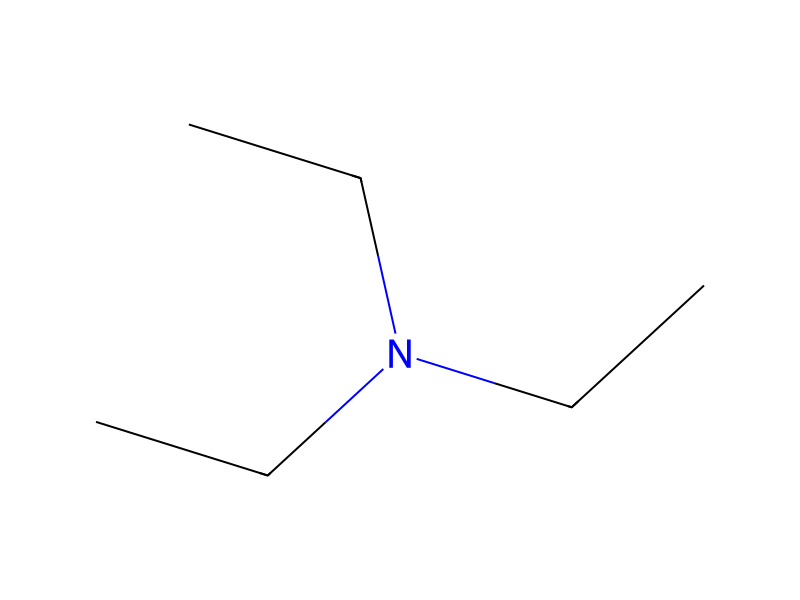
Triethylamine → tert-Butyl 2-{7-[4-(4-cyanophenyl)butyl]-9-oxa-3,7-diazabicyclo[3.3.1]non-3-yl}ethylcarbamate

HRID 1374121
Print

Reaction equation

HRID 1374121 reaction equation
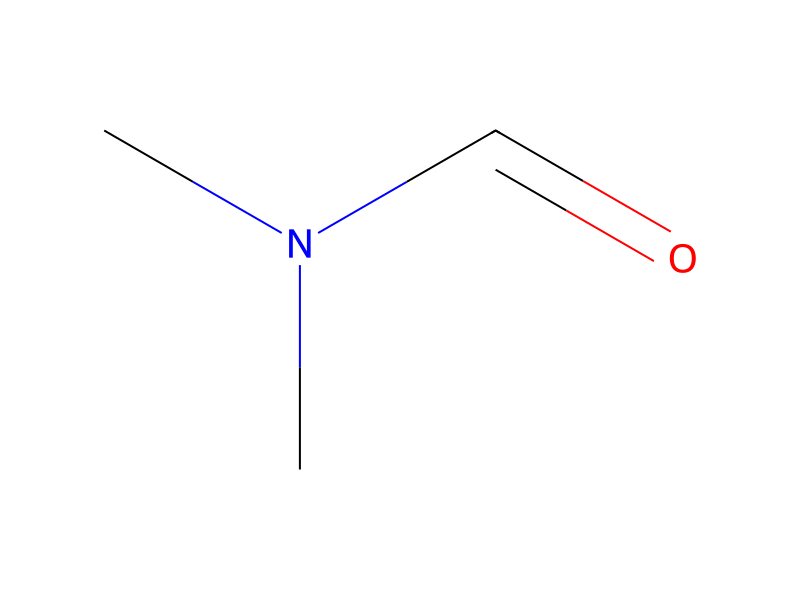
C(N(CC)CC)C.Br[CH2:9][CH2:10][NH:11][C:12](=[O:18])[O:13][C:14]([CH3:17])([CH3:16])[CH3:15].[CH:19]12[O:27][CH:23]([CH2:24][NH:25][CH2:26]1)[CH2:22][N:21]([CH2:28][CH2:29][CH2:30][CH2:31][C:32]1[CH:39]=[CH:38][C:35]([C:36]#[N:37])=[CH:34][CH:33]=1)[CH2:20]2>CN(C=O)C>[C:36]([C:35]1[CH:34]=[CH:33][C:32]([CH2:31][CH2:30][CH2:29][CH2:28][N:21]2[CH2:22][CH:23]3[O:27][CH:19]([CH2:26][N:25]([CH2:9][CH2:10][NH:11][C:12](=[O:18])[O:13][C:14]([CH3:17])([CH3:16])[CH3:15])[CH2:24]3)[CH2:20]2)=[CH:39][CH:38]=1)#[N:37]
ParticipantsConditionsProcedureWorkupSources & evidenceReaction notes

Participants

5

Reactant 3

HCID 8471TriethylamineHCID 4103526tert-Butyl N-(2-bromoethyl)carbamateHCID 179653304-[4-(9-Oxa-3,7-diazabicyclo[3.3.1]non-3-yl)butyl]benzonitrile

Product 1

HCID 9888837tert-Butyl 2-{7-[4-(4-cyanophenyl)butyl]-9-oxa-3,7-diazabicyclo[3.3.1]non-3-yl}ethylcarbamate

Reagents, catalysts & solvents 1

HCID 6228
Dimethylformamide
Solvent

Conditions

Temperature
54 °C

Procedure

Triethylamine (2.2 mL, 0.016 mol) and tert-butyl 2-bromoethylcarbamate (see Preparation R above, 2.83 g, 0.013 mol) were added to a solution of 4-[4-(9-oxa-3,7-diazabicyclo[3.3.1]non-3-yl)butyl]benzonitrile (see Preparation G above, 3.0 g, 0.011 mol) in DMF (50 mL). The mixture was stirred for 24 h at 54° C., cooled to 25° C., and concentrated in vacuo. The residue was dissolved in chloroform and washed with saturated sodium chloride. The aqueous layer was separated and extracted with chloroform (2×150 mL). The combined organic layers were dried (Na2SO4) and concentrated in vacuo. The resulting material was chromatographed on silica gel, eluting first with chloroform:acetonitrile:conc. ammonium hydroxide (9:1:0.02) until the higher Rf impurities were removed. Then the eluent was switched to chloroform:methanol:conc. ammonium hydroxide (9:1:0.02). This afforded 2.76 g (61%) of the title compound.

Workup

  1. temperaturecooled to 25° C.
  2. concentrationconcentrated in vacuo
  3. dissolutionThe residue was dissolved in chloroform
  4. washwashed with saturated sodium chloride
  5. customThe aqueous layer was separated
  6. extractionextracted with chloroform (2×150 mL)
  7. dry with materialThe combined organic layers were dried (Na2SO4)
  8. concentrationconcentrated in vacuo
  9. customThe resulting material was chromatographed on silica gel
  10. washeluting first with chloroform
  11. customammonium hydroxide (9:1:0.02) until the higher Rf impurities were removed

Sources & evidence

ORD record
ord-af550c2ebfca4880920ffe4ddb7dc129
Source dataset
uspto-grants-2007_01
DOI
10.6084/m9.figshare.5104873.v1
Patent number
US07164017B2

Reaction notes

On this pageParticipantsConditionsProcedureWorkupSources & evidenceReaction notes