反応式

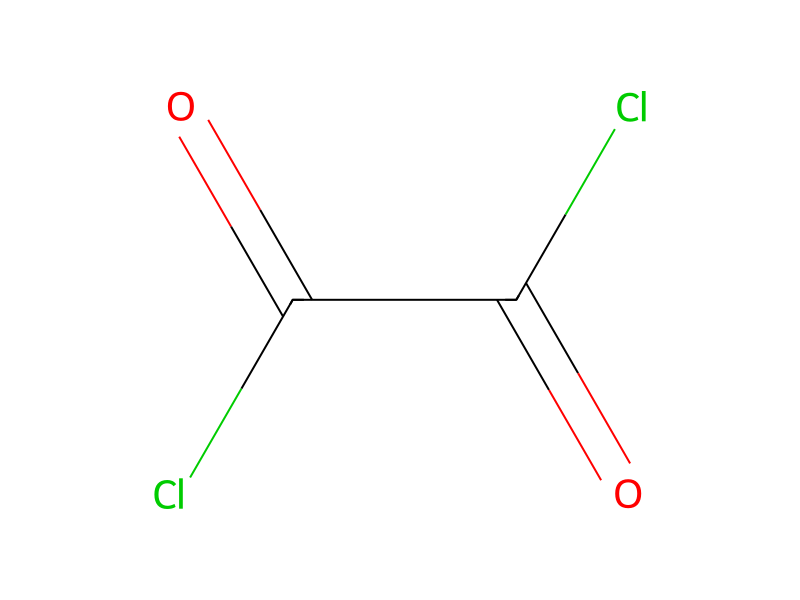
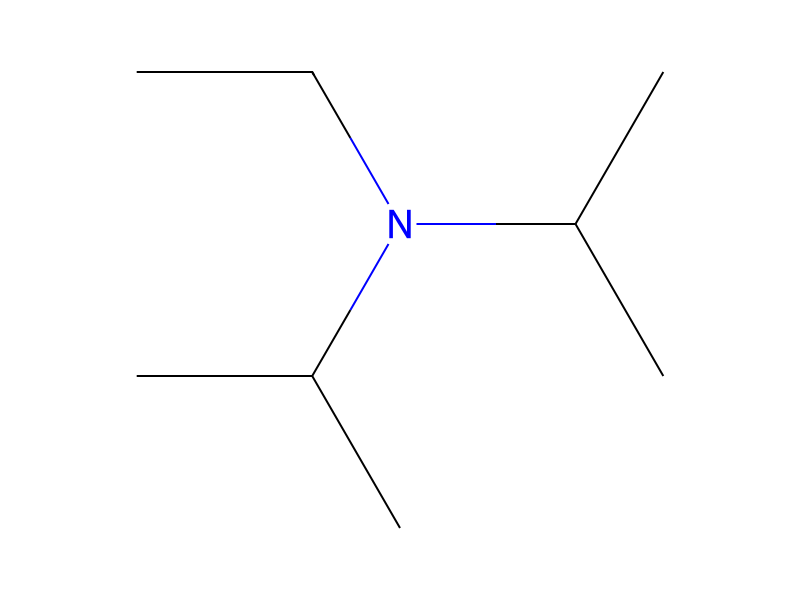
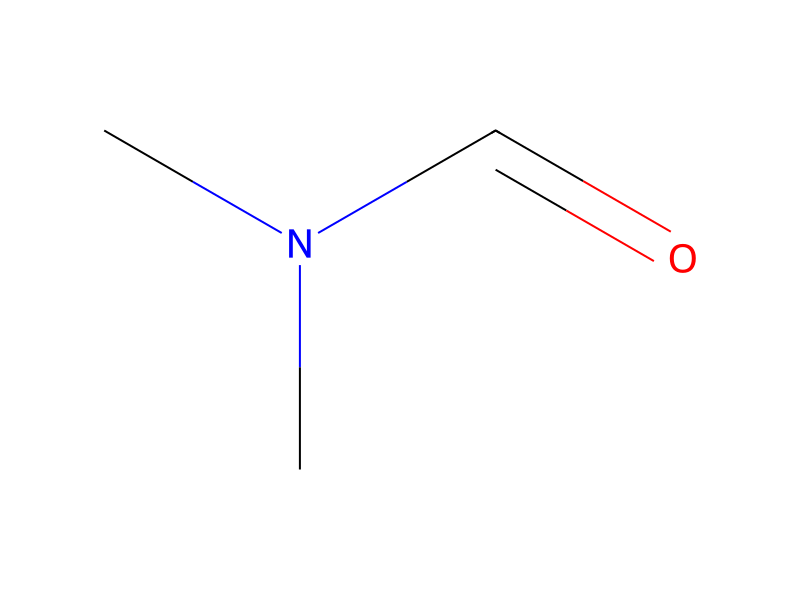
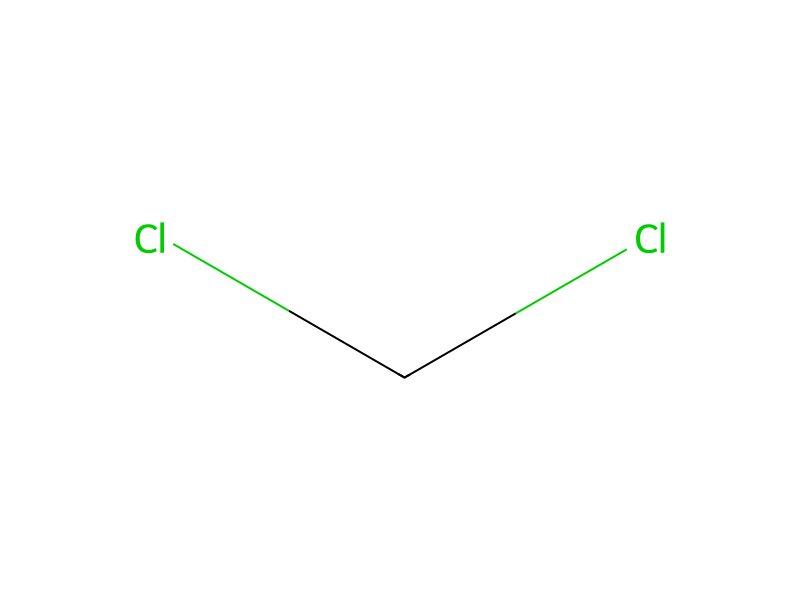
[Cl:1][C:2]1[CH:7]=[CH:6][CH:5]=[CH:4][C:3]=1[C:8]1[C:9]2[N:17]([CH3:18])[C:16](=[O:19])[C:15]([C:20]3[CH:21]=[C:22]([CH:26]=[CH:27][C:28]=3[CH3:29])[C:23](O)=[O:24])=[CH:14][C:10]=2[CH:11]=[N:12][N:13]=1.C(Cl)(=O)C(Cl)=O.[NH2:36][C:37]1[CH:41]=[CH:40][O:39][N:38]=1.C(N(CC)C(C)C)(C)C>C(Cl)Cl.CN(C=O)C>[Cl:1][C:2]1[CH:7]=[CH:6][CH:5]=[CH:4][C:3]=1[C:8]1[C:9]2[N:17]([CH3:18])[C:16](=[O:19])[C:15]([C:20]3[CH:21]=[C:22]([CH:26]=[CH:27][C:28]=3[CH3:29])[C:23]([NH:36][C:37]3[CH:41]=[CH:40][O:39][N:38]=3)=[O:24])=[CH:14][C:10]=2[CH:11]=[N:12][N:13]=1参加物
反応物 4


生成物 1

試薬・触媒・溶媒 2
反応条件
- 温度
- 0 °C
実験手順
To a suspension of 3-(8-(2-chlorophenyl)-1-methyl-2-oxo-1,2-dihydropyrido[3,2-d]pyridazin-3-yl)-4-methylbenzoic acid (100 mg, 0.25 mmol; prepared by the general method shown in scheme 2) in DCM at 0° C. was added oxalyl chloride (43.7 μl, 0.49 mmol) followed by a drop of DMF. The reaction was stirred at 0° C. for 15 min then warmed to RT for 1 h. The volatile solvents were removed under reduced pressure and the residue was dissolved in DCM. To the reaction mixture was added 3-aminoisoxazole (54.6 μl, 0.74 mmol) and N,N-diisopropylethylamine (42.9 μl, 0.25 mmol). After the reaction mixture was stirred at RT for 3 h, the solvent was then removed in vacuo. The residual crude material was purified with Reverse phase-HPLC [elution gradient of 10-50% of (0.1% TFA in acetonitrile) in (0.1% of TFA) in water] and the fractions containing the product were combined and concentrated in vacuo. The remaining crude mixture was stirred with EtOAc and sat. Na2CO3. The organic layer was separated and washed with brine, separated, then dried with sodium sulfate and concentrated to give 3-(8-(2-chlorophenyl)-1-methyl-2-oxo-1,2-dihydropyrido[3,2-d]pyridazin-3-yl)-N-(isoxazol-3-yl)-4-methylbenzamide (33.0 mg) as a white solid. 1H NMR (400 MHz, DMSO-d6) δ ppm 11.43 (1H, s), 9.48 (1H, s), 8.81 (1H, s), 8.24 (1H, s), 7.95-8.10 (2H, m), 7.76-7.86 (1H, m), 7.56-7.74 (3H, m), 7.46 (1H, d, J=8.0 Hz), 7.05 (1H, s), 3.12 (3H, s), 2.24 (3H, s); Mass Found: LC-MS (ESI): m/e=472/474
後処理
- customprepared by the general method
- customat 0° C.
- temperaturethen warmed to RT for 1 h
- customThe volatile solvents were removed under reduced pressure
- dissolutionthe residue was dissolved in DCM
- stirringAfter the reaction mixture was stirred at RT for 3 h
- customthe solvent was then removed in vacuo
- customThe residual crude material was purified with Reverse phase-HPLC [elution gradient of 10-50% of (0.1% TFA in acetonitrile) in (0.1% of TFA) in water]
- additionthe fractions containing the product
- concentrationconcentrated in vacuo
- stirringThe remaining crude mixture was stirred with EtOAc and sat. Na2CO3
- customThe organic layer was separated
- washwashed with brine
- customseparated
- dry with materialdried with sodium sulfate
- concentrationconcentrated
出典と根拠
- ORDレコード
- ord-20b768b1b01f4ab9a1c39efe082d21f6
- 出典データセット
- uspto-grants-2013_06
- 特許番号
- US08455495B2
関連ノート
読み込み中…