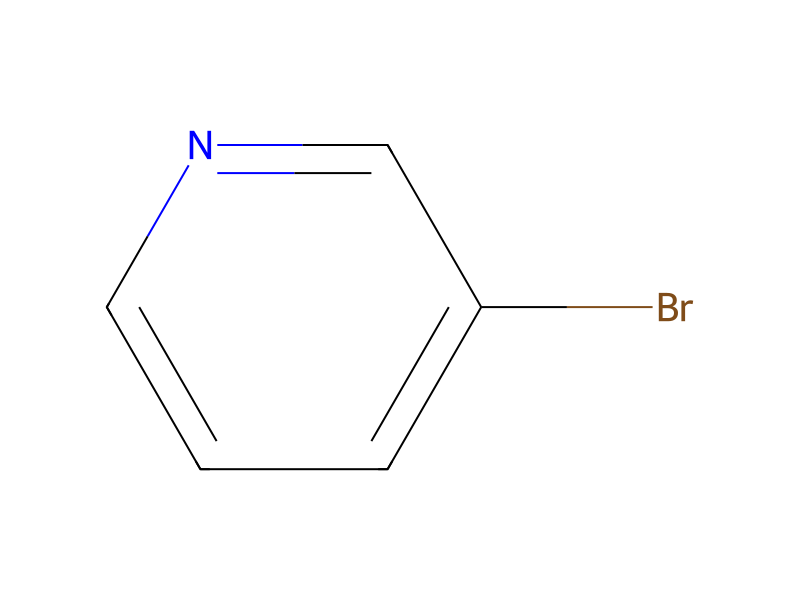
3-Bromopyridine → 2-(7-Fluoro-2,4-dioxo-5-pyridin-3-yl-2,3,4,5-tetrahydro-benzo[b][1,4]diazepin-1-yl)-N-isopropyl-N-(4-methoxy-phenyl)-acetamide
反応式

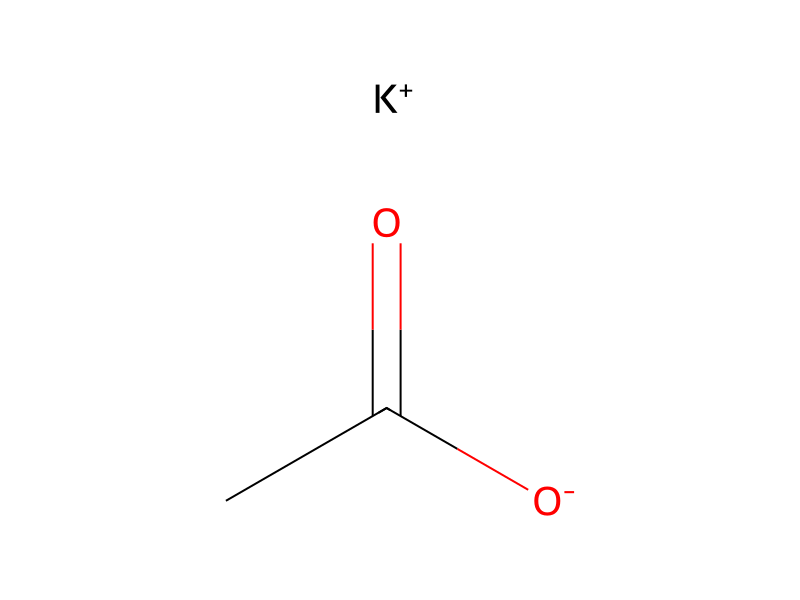
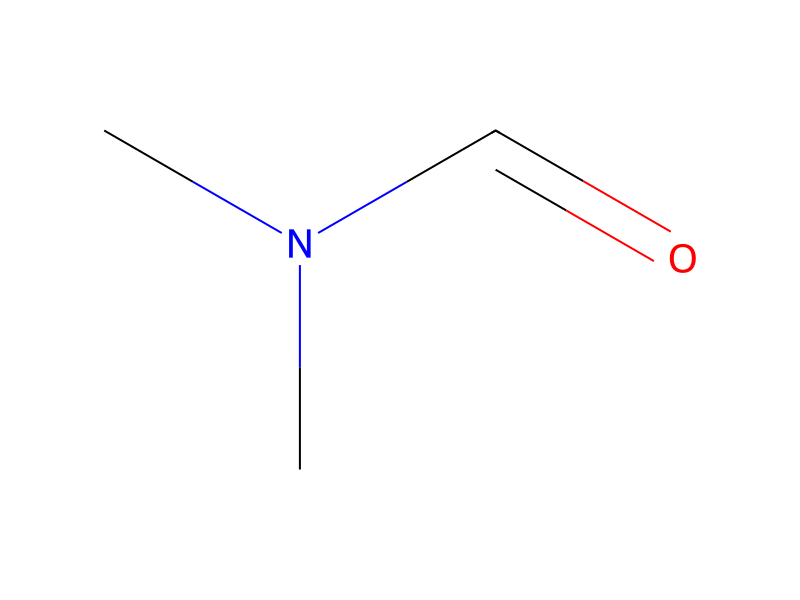
[F:1][C:2]1[CH:29]=[CH:28][C:5]2[N:6]([CH2:13][C:14]([N:16]([CH:25]([CH3:27])[CH3:26])[C:17]3[CH:22]=[CH:21][C:20]([O:23][CH3:24])=[CH:19][CH:18]=3)=[O:15])[C:7](=[O:12])[CH2:8][C:9](=[O:11])[NH:10][C:4]=2[CH:3]=1.C([O-])(=O)C.[K+].Br[C:36]1[CH:37]=[N:38][CH:39]=[CH:40][CH:41]=1>CN(C)C=O.[Cu]>[F:1][C:2]1[CH:29]=[CH:28][C:5]2[N:6]([CH2:13][C:14]([N:16]([CH:25]([CH3:27])[CH3:26])[C:17]3[CH:22]=[CH:21][C:20]([O:23][CH3:24])=[CH:19][CH:18]=3)=[O:15])[C:7](=[O:12])[CH2:8][C:9](=[O:11])[N:10]([C:36]3[CH:37]=[N:38][CH:39]=[CH:40][CH:41]=3)[C:4]=2[CH:3]=1参加物
反応物 3

生成物 1

試薬・触媒・溶媒 2
反応条件
- 温度
- 100 °C
実験手順
A mixture of 0.880 g of 2-(7-Fluoro-2,4-dioxo-2,3,4,5-tetrahydrobenzo[b][1,4]diazepin-1-yl)-N-isopropyl-N-(4-methoxy-phenyl)-acetamide (2.20 mmol), 420 mg of copper (bronze, 6.61 mmol, 3 equiv), 476 mg of potassium acetate (4.85 mmol, 2.2 equiv), and 0.290 mL of 3-bromopyridine (4.85 mmol, 2.2 equiv) in 10 mL of dimethylformamide was heated at 100° C. under nitrogen for 3 hrs. An additional 0.132 mL of 3-bromopyridine (2.43 mmol, 1.1 equiv) was added and the reaction was maintained for an additional 2 hrs. The reaction mixture was cooled to ambient temperature, filtered through a sintered glass funnel and then evaporated in vacuo to a residue. The residue was partitioned between ethyl acetate and aqueous ammonium hydroxide (5 mL conc. diluted to 100 mL). After separating the layers, the organic layer was washed with aqueous ammonium hydroxide (5 mL conc. diluted to 100 mL), and then saturated aqueous brine. The organic phase was then extracted three times with aqueous HCl (1N). The acid layers were combined and neutralized with aqueous sodium hydroxide (1N). The neutralized mixture was extracted twice with methylene chloride. The methylene chloride layers were combined, dried over anhydrous sodium sulfate, filtered and evaporated in vacuo to a residue. The residue was triturated with hexane and then concentrated in vacuo to provide 0.705 g of the title compound as a tan solid. H NMR (300 MHz, CDCl3) d 8.60 (m, 2H), 8.08 (b, 1H), 7.54 (b, 1H), 7.39 (m, 1H), 7.15 (m, 2H), 7.00 (m, 3H), 6.57 (dd, 1H, J=2.7, 9.2), 4.95 (m, 1H), 4.32 (d, 1H, J=17.9), 4.14 (d, 1H, J=17.8), 3.85 (s, 3H), 3.61 (d, 1H, J=12.1), 3.52 (d, 1H, J=12.1), 1.06 (d, 6H, J=6.8); low resolution MS (FAB)m/e 477 (MH+).
後処理
- temperaturethe reaction was maintained for an additional 2 hrs
- temperatureThe reaction mixture was cooled to ambient temperature
- filtrationfiltered through a sintered glass funnel
- customevaporated in vacuo to a residue
- customThe residue was partitioned between ethyl acetate and aqueous ammonium hydroxide (5 mL conc. diluted to 100 mL)
- customAfter separating the layers
- washthe organic layer was washed with aqueous ammonium hydroxide (5 mL conc. diluted to 100 mL)
- extractionThe organic phase was then extracted three times with aqueous HCl (1N)
- extractionThe neutralized mixture was extracted twice with methylene chloride
- dry with materialdried over anhydrous sodium sulfate
- filtrationfiltered
- customevaporated in vacuo to a residue
- customThe residue was triturated with hexane
- concentrationconcentrated in vacuo
出典と根拠
- ORDレコード
- ord-e4966b9511754bc0a64cc6785a0becc5
- 出典データセット
- uspto-grants-1998_04
- 特許番号
- US05739129
反応ノート
読み込み中…