反応式

[Mg].[CH:2]1(Br)[CH2:8][CH2:7][CH2:6][CH2:5][CH2:4][CH2:3]1.[Cl-].[Li+].[Cu](C#N)C#N.C1([Mg]Br)CCCCCC1.[C:26]([O:30][CH3:31])(=[O:29])[C:27]#[CH:28].[I:32]I>O1CCCC1.BrCCBr>[CH3:31][O:30][C:26](=[O:29])/[C:27](/[I:32])=[CH:28]\[CH:2]1[CH2:8][CH2:7][CH2:6][CH2:5][CH2:4][CH2:3]1参加物
反応物 7
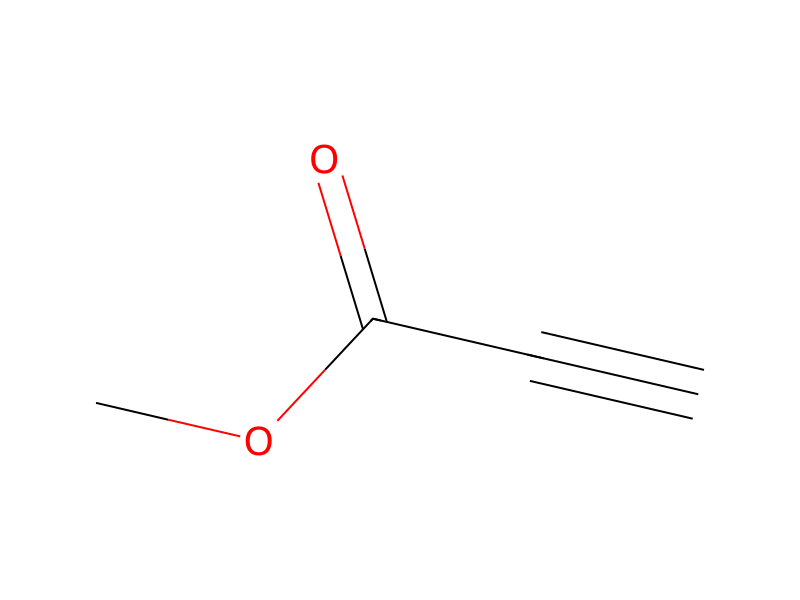

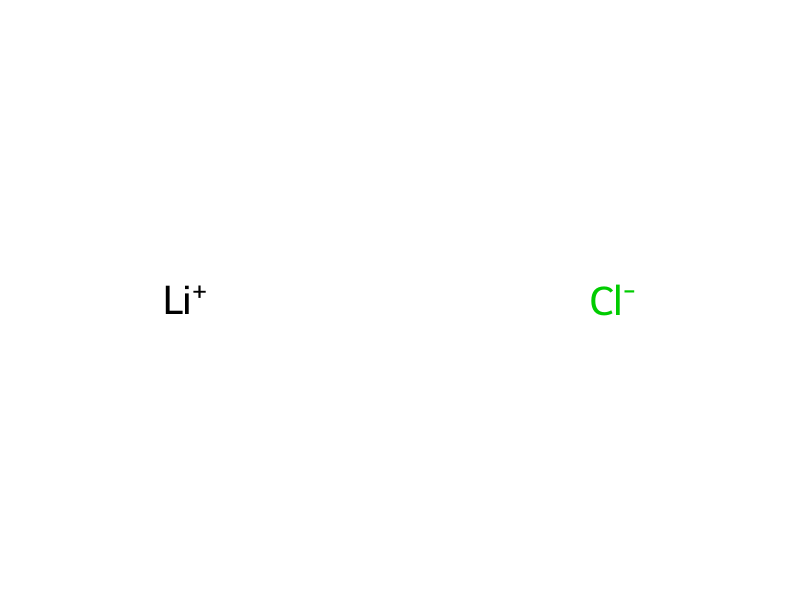
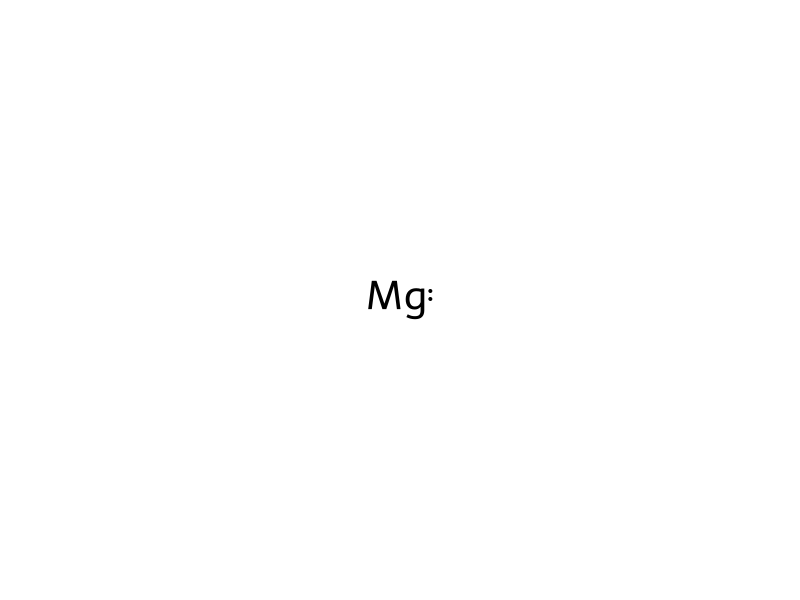


生成物 1

試薬・触媒・溶媒 2

反応条件
- 温度
- -70 °C
実験手順
A mixture of magnesium metal (4.81 g, 200 mmol) and dry tetrahydrofuran (10 mL) under argon was treated with a solution of 1,2-dibromoethane (0.94 g, 5 mmol) in dry tetrahydrofuran (5 mL). The resulting reaction mixture was stirred for 10 min to activate the magnesium metal. The reaction mixture was then treated dropwise with a solution of cycloheptyl bromide (17.7 g, 100 mmol) in dry tetrahydrofuran (30 mL), one-fifth portion over a period of 5 min. The resulting reaction mixture was stirred for 5-10 min to initiate the exothermic reaction. The remaining portion of the cycloheptyl bromide solution was then added dropwise while controlling the inside temperature below 50° C. After complete addition, the solution was stirred for 1 h and then diluted with dry tetrahydrofuran (80 mL). In a separate reaction flask, a mixture of lithium chloride (8.48 g, 200 mmol, predried at 130° C. under high vacuum for 3 h) and copper cyanide (8.96 g, 100 mmol) in dry tetrahydrofuran (110 mL) was stirred at 25° C. under argon for 10 min to obtain a clear solution. The reaction mixture was cooled to −70° C. and then slowly treated with the freshly prepared cycloheptylmagnesium bromide. After the addition, the reaction mixture was allowed to warm to −10° C. where it was stirred for 5 min. The resulting reaction mixture was again cooled back to −70° C. and then treated with methyl propiolate (7.57 g, 90 mmol). The reaction mixture was stirred for 15 h at −70° C. to −50° C. and then slowly treated with a solution of iodine (34.3 g, 135 mmol) in dry tetrahydrofuran (30 mL), with the temperature kept at −70° C. to −60° C. After addition of the iodine solution, the cooling bath was removed, and the reaction mixture was allowed to warm to 25° C. where it was stirred for 2 h. The reaction mixture was then poured into a solution consisting of a saturated aqueous ammonium chloride solution (400 mL) and ammonium hydroxide (100 mL), and the organic compound was extracted into ethyl acetate (3×200 mL). The combined organic extracts were successively washed with a saturated aqueous sodium thiosulfate solution (1×400 mL) and a saturated aqueous sodium chloride solution (1×400 mL). The organic layer was then dried over anhydrous magnesium sulfate, filtered, and concentrated in vacuo. Flash chromatography (Merck Silica gel 60, 230-400 mesh, 20/1 to 10/1 hexanes/diethyl ether) afforded (E)-3-cycloheptyl-2-iodo-acrylic acid methyl ester (17.86 g, 64%) as a colorless oil: EI-IIRMS m/e calcd for C11H17IO2 (M+) 308.0273, found 308.0273.
後処理
- customThe resulting reaction mixture
- customThe resulting reaction mixture
- stirringwas stirred for 5-10 min
- customthe exothermic reaction
- customthe inside temperature below 50° C
- additionAfter complete addition
- stirringthe solution was stirred for 1 h
- customIn a separate reaction flask
- stirringwas stirred at 25° C. under argon for 10 min
- customto obtain a clear solution
- additionAfter the addition
- temperatureto warm to −10° C. where it
- stirringwas stirred for 5 min
- customThe resulting reaction mixture
- temperaturewas again cooled back to −70° C.
- stirringThe reaction mixture was stirred for 15 h at −70° C. to −50° C.
- customkept at −70° C. to −60° C
- customthe cooling bath was removed
- temperatureto warm to 25° C. where it
- stirringwas stirred for 2 h
- additionThe reaction mixture was then poured into a solution
- extractionthe organic compound was extracted into ethyl acetate (3×200 mL)
- washThe combined organic extracts were successively washed with a saturated aqueous sodium thiosulfate solution (1×400 mL)
- dry with materialThe organic layer was then dried over anhydrous magnesium sulfate
- filtrationfiltered
- concentrationconcentrated in vacuo
出典と根拠
- ORDレコード
- ord-c2d366eb401742cab9ce98b0eda8da0e
- 出典データセット
- uspto-grants-2002_03
- 特許番号
- US06353111B1
関連ノート
読み込み中…