Hydrochloric Acid → 3-Chloro-4,5-dihydroxybenzaldehyde
Reaction equation

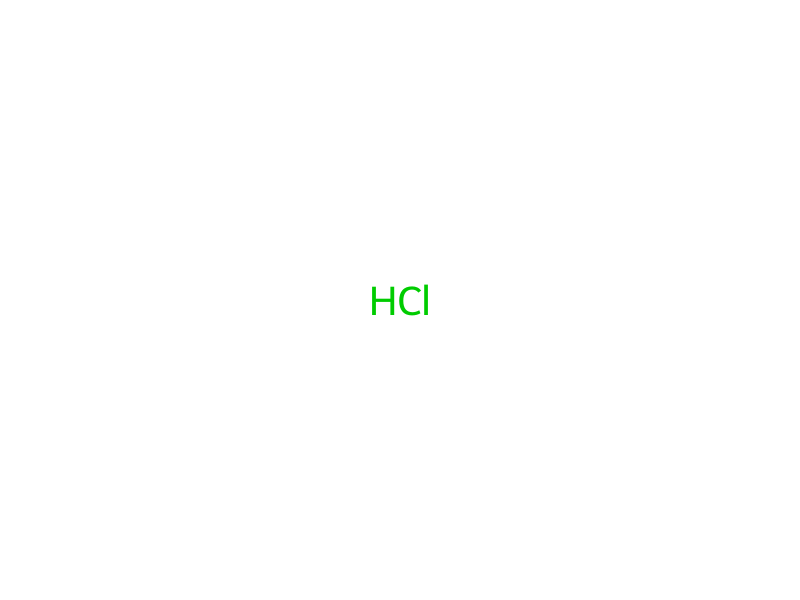
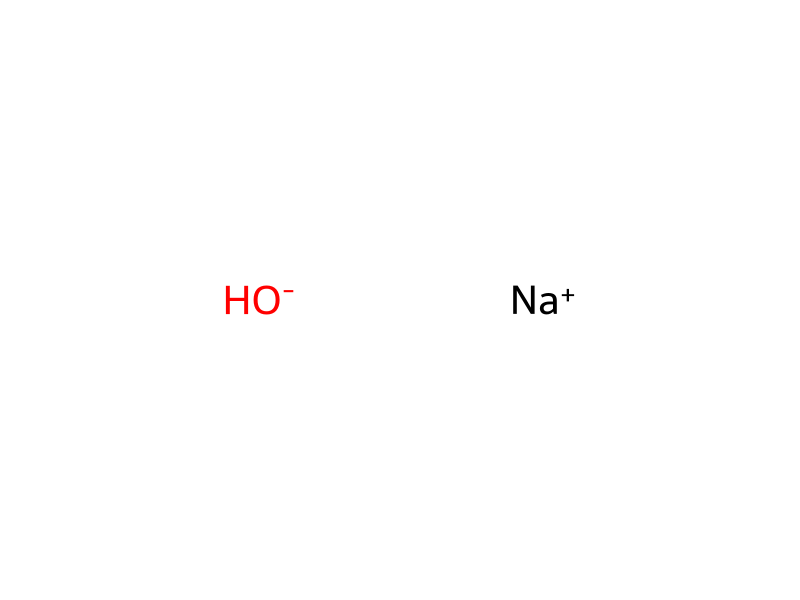
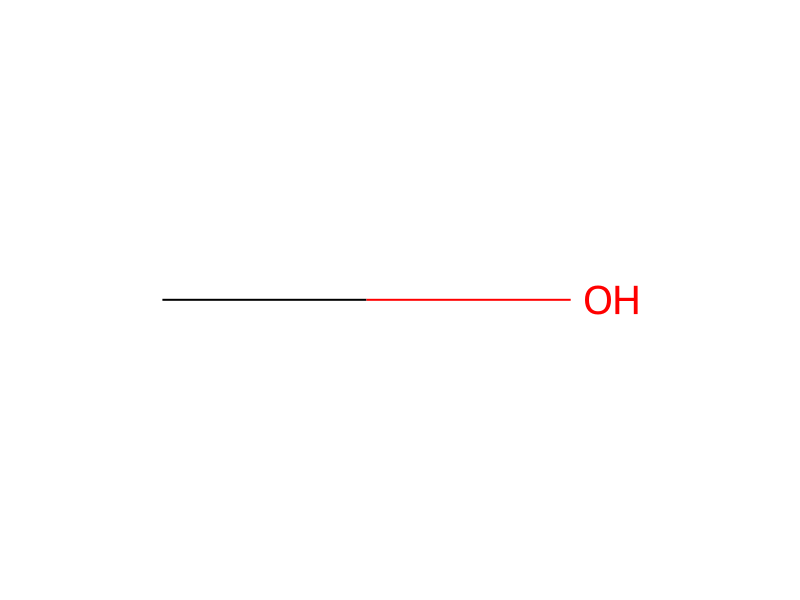
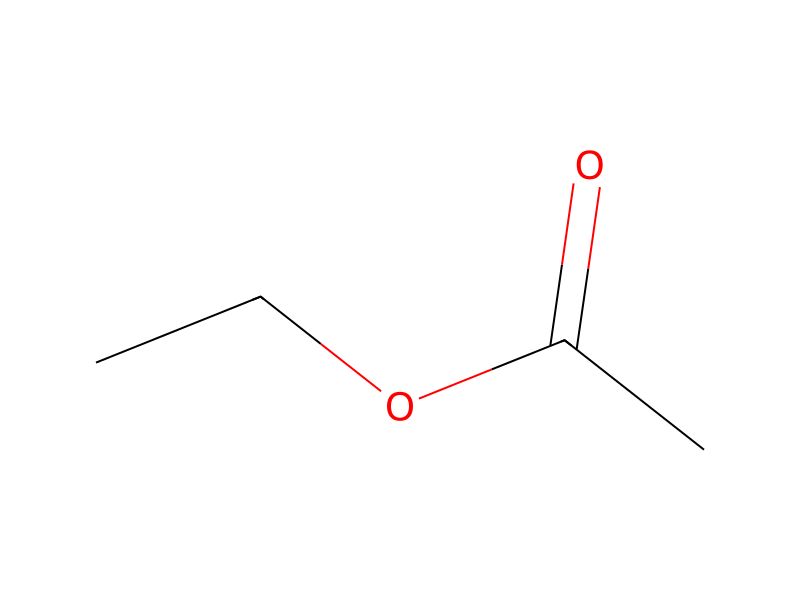
C([O:5][C:6]1[C:16]([Cl:17])=[CH:15][C:14]([CH2:18]N(C=O)O)=[CH:13][C:7]=1[O:8]CC(O)=O)CCC.C([O:25]C(=O)COC1C=C(CN(C=O)O)C=C(Cl)C=1OCCCC)C.[OH-].[Na+].Cl>CO.O.C(OCC)(=O)C>[Cl:17][C:16]1[CH:15]=[C:14]([CH:13]=[C:7]([OH:8])[C:6]=1[OH:5])[CH:18]=[O:25]Participants
Reactant 4


Product 1

Reagents, catalysts & solvents 3

Procedure
To a solution of 3-Chloro-4-hydroxy-5-methoxybenzaldehyde (1.0 g, 5.4 mmol) in dichloromethane (150 ml) stirred at −78° C. was added dropwise a solution of boron tribromide in dichloromethane (5.4 ml of 1M solution, 5.4 mmol). Upon completion of the addition, the resulting solution was stirred 18 h at room temperature. The reaction was quenched by addition of methanol then evaporated twice from methanol and the residue purified by reversed-phase preparative HPLC to afford the title compound (0.66 g, 72%). 1H NMR (400 MHz, CD3OD) δ 9.69 (s, 1H), 7.42 (s, 1H), 7.25 (s, 1H). M+1=173. b) 4-Butoxy-3-chloro-5-hydroxybenzaldehyde A solution consisting of 3-chloro-4,5-dihydroxybenzaldehyde (3.0 g, 17.4 mmol), n-butanol (1.42 g, 19.1 mmol), triphenylphosphine (5.02 g, 19.4 mmol), and diethylazodicarboxylate (3.32 g, 19.1 mmol) in THF (60 ml) was stirred at room temperature for 24 h. All volatiles were removed in-vacuo and the residue purified by preparative HPLC to afford the title compound (2.7 g, 68%). 1H NMR (400 MHz, CDCl3) δ 9.84 (s, 1H), 7.46 (d, j=1.8 Hz, 1H), 7.38 (d, j=1.8 Hz, 1H), 5.91 (s, 1H), 4.18 (t, j=6.7 Hz, 2H), 1.83 (m, 2H), 1.51 (m, 2H), 1.00 (t, j=7.4). M+1=299. c) (2-Butoxy-3-chloro-5-formylphenoxy)acetic acid ethyl ester Sodium hydride (84 mg 60% in mineral oil, 2.09 mmol) was added in portions to a solution of 4-butoxy-3-chloro-5-hydroxybenzaldehyde (400 mg, 1.74 mmol) in DMF (5 ml). The resultant mixture was stirred at room temperature for 10 min then ethyl bromoacetate (583 mg, 3.5 mmol) was added slowly. After 1.5 h at room temperature, HPLC indicated complete reaction. The mixture was partitioned between ethyl acetate and water, the organic solution was washed with water then brine and dried with magnesium sulfate. Removal of all volatiles in-vacuo afforded the title compound (100% crude yield). 1H NMR (400 MHz, CDCl3) δ 9.82 (s, 1H), 7.55 (d, j=1.8 Hz, 1H), 7.25 (d, j=1.8 Hz, 1H), 4.73 (s, 2H), 4.21-4.30 (m, 4H) 1.79-1.60 (m, 2H), 1.52-1.60 (m, 2H), 1.31 (t, j=7.1 Hz, 3H), 0.98 (t, j=7.1 Hz, 3H). M+1=315. d) 2-Butoxy-3-chloro-5-[(formylhydroxyamino)methyl]phenoxyacetic acid ethyl ester 2-Butoxy-3-chloro-5-formylphenoxyacetic acid ethyl ester was converted to the title compound using the methods described in example 2d. 1H NMR (400 MHz, CDCl3) δ 8.38* 8.00 (s, 1H), 7.01* 6.98 (s, 1H), 6.70 (s, 1H), 4.68 (s, 2H), 4.60* 4.54 (s, 2H), 4.25 (q, j=7.1 Hz, 2H), 4.07 (t, j=6.6 Hz, 2H) 1.75-1.82 (m, 2H), 1.50-1.56 (m, 2H), 1.29 (t, j=7.1 Hz, 3H), 0.97 (t, j=7.3 Hz, 3H). M+1=360. * signals due to minor rotamer e) 2-Butoxy-3-chloro-5-[(formylhydroxyamino)methyl]phenoxyacetic acid A solution of 2-butoxy-3-chloro-5-[(formylhydroxyamino)methyl]phenoxyacetic acid ethyl ester (0.45 mmol crude) in methanol (5 ml) with 10% aqueous NaOH solution (0.5 ml) was stirred for 1 h. To the resulting solution was added ethyl acetate (20 ml) and water (20 ml). The pH was brought to 6 with 3 N HCl and the layers separated. The organic extract was dried with magnesium sulfate and all volatiles removed. The residue was purified by preparative HPLC to afford the title compound (29 mg, 20% for four steps). 1H NMR (400 MHz, CD3OD) δ 8.37* 8.18 (s, 1H), 7.04* 7.02 (s, 1H), 6.90* 6.88 (s, 1H), 4.75* 4.74 (s, 2H), 4.63* 4.10 (m, 2H), 1.76-1.79 (m, 2H), 1.53-1.59 (m, 2H), 0.98-1.02 (m, 3H). M+1=332. * signals due to rotamer f) 2-{2-Butoxy-3-chloro-5-[(formylhydroxyamino)methyl]-phenoxy}-N-hydroxyformamide To a solution of 2-butoxy-3-chloro-5-[(formylhydroxyamino)methyl]phenoxyacetic acid (crude 0.5 mmol) in THF (4 ml) with TEA (0.167 ml, 1.2 mmol) was added ethyl chloroformate (0.115 ml, 1.2 mmol). This was stirred 15 min then methyl amine (0.21 ml of 40% aqueous solution) was added and stirring continued for an additional 30 min. The mixture was partitioned between ethyl acetate and water aqueous layer adjusted to pH 5 with 1 N HCl). The organic extract was dried with magnesium sulfate and all volatiles removed in vacuo. Purification by preparative HPLC afforded the title compound (10 mg). 1H NMR (400 MHz, CD3OD) δ 8.38* 8.18 (s, 1H), 7.11* 7.08 (s, 1H), 6.98* 6.95 (s, 1H), 4.59-4.65 (m, 4H), 2.84 (s, 3H), 1.76-1.80 (m, 2H), 1.55-1.60 (m, 2H), 1.01 (t, j=7.4 Hz, 3H). M+1=345. * signals due to rotamer
Workup
- customthe layers separated
- extractionThe organic extract
- dry with materialwas dried with magnesium sulfate
- customall volatiles removed
- customThe residue was purified by preparative HPLC
Sources & evidence
- ORD record
- ord-b4ba68c9c5054f03a1dfac76b5eaf877
- Source dataset
- uspto-grants-2005_11
- Patent number
- US06967220B2
Reaction notes
Loading…