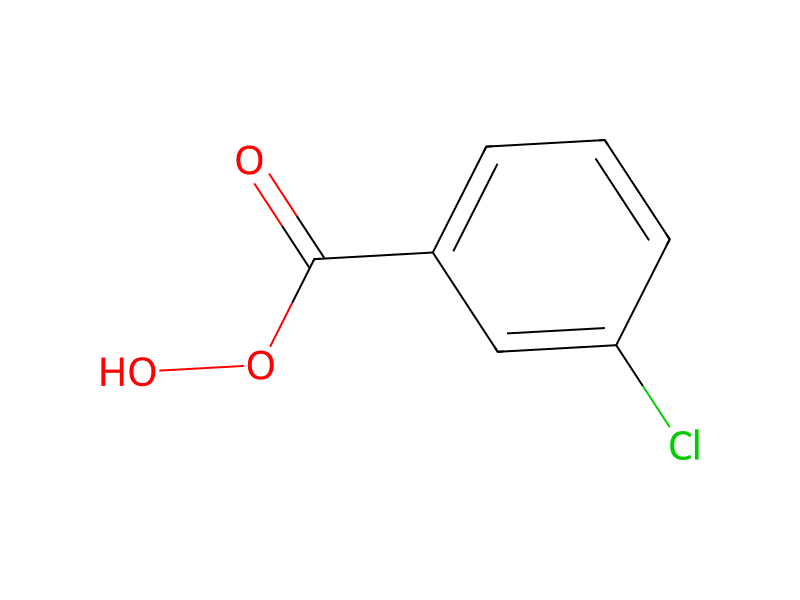
m-Chloroperbenzoic acid → HCID 53629045, compound
Reaction equation

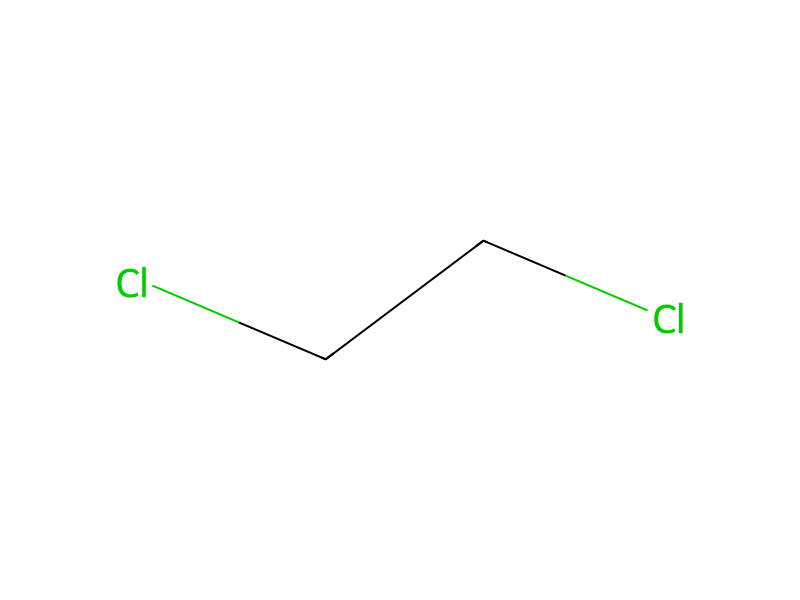
[Cl:1][C:2]1[C:3]2[N:4]([C:8]([CH:27]3[CH2:30][CH2:29][CH2:28]3)=[N:9][C:10]=2[C:11]2[CH:20]=[C:19]3[C:14]([CH:15]=[CH:16][C:17]([C:21]4[CH:26]=[CH:25][CH:24]=[CH:23][CH:22]=4)=[N:18]3)=[CH:13][CH:12]=2)[CH:5]=[CH:6][N:7]=1.C1C=C(Cl)C=C(C(OO)=[O:39])C=1>C(Cl)CCl>[Cl:1][C:2]1[C:3]2[N:4]([C:8]([CH:27]3[CH2:30][CH2:29][CH2:28]3)=[N:9][C:10]=2[C:11]2[CH:20]=[C:19]3[C:14]([CH:15]=[CH:16][C:17]([C:21]4[CH:26]=[CH:25][CH:24]=[CH:23][CH:22]=4)=[N+:18]3[O-:39])=[CH:13][CH:12]=2)[CH:5]=[CH:6][N:7]=1Participants
Reactant 2

Product 1

Reagents, catalysts & solvents 1
Procedure
To a cooled (ice-H2O) solution of 7-(8-chloro-3-cyclobutylimidazo[1,5-a]pyrazin-1-yl)-2-phenylquinoline (197 mg, 0.48 mmol) in ClCH2CH2Cl (20 mL) was added mCPBA (97 mg, max 0.43 mmol, max. 77% Aldrich) in one portion. The solution was stirred at the temperature for 30 min and then allowed to warm to rt by removing the cooling bath and stirred at rt (2 h). The reaction mixture was again cooled (ice-H2O) and treated with another portion of mCPBA (107 mg, max 0.48 mmol), stirred for 30 min at the temperature and then overnight at rt (15 h). After that time the crude mixture was filtered through hydromatrix (25 mL) pretreated with 2 M aq NaOH (10 mL). The hydromatrix column was washed with DCM (˜100 mL) and the filtrate was concentrated under reduced pressure. The resultant yellow residue was purified by flash chromatography on silica gel (70 g cartridge, 0→0.75→4% MeOH in DCM) to yield 7-(8-chloro-3-cyclobutylimidazo[1,5-a]pyrazin-1-yl)-2-phenylquinoline 1-oxide as a yellow gum. A cooled (−10° C.) i-PrOH (15 mL) solution of 7-(8-chloro-3-cyclobutylimidazo[1,5-a]pyrazin-1-yl)-2-phenylquinoline 1-oxide (50 mg) in a Parr bomb was saturated with NH3(g) for 3 min. The vessel was sealed and heated at 100-110° C. (bath temperature) for 2 d. The reaction mixture was then cooled to rt, concentrated under reduced pressure and purified by flash chromatography on silica gel (0-4% MeOH+2%˜6 M NH3 in MeOH. A trituration with hexanes (3×) provided the title material as a bright yellow solid; 1H NMR (400 MHz, CDCl3) δ 9.12 (s, 1H), 8.16 (dd, J=8.2 Hz, 1.8 Hz, 1H), 8.04-7.96 (m, 3H), 7.79 (d, J=8.8 Hz, 1H), 7.57-7.43 (m, 4H), 7.16 (d, J=5.2 Hz, 1H), 7.13 (d, J=4.8 Hz, 1H), 5.30 (s, 2H), 3.85 (quintet, J=8.2 Hz, 1H), 2.69-2.60 (m, 2H), 2.55-2.50 (m, 2H), 2.30-2.15 (m, 1H), 2.15-2.00 (m, 1H); MS (ES+): m/z 408.13 (100) [MH+]; HPLC: tR=2.14 min (OpenLynx, polar—5 min).
Workup
- customby removing the cooling bath
- stirringstirred at rt (2 h)
- temperatureThe reaction mixture was again cooled (ice-H2O)
- stirringstirred for 30 min at the temperature
- customovernight
- customat rt
- custom(15 h)
- filtrationAfter that time the crude mixture was filtered through hydromatrix (25 mL)
- washThe hydromatrix column was washed with DCM (˜100 mL)
- concentrationthe filtrate was concentrated under reduced pressure
- customThe resultant yellow residue was purified by flash chromatography on silica gel (70 g cartridge, 0→0.75→4% MeOH in DCM)
Sources & evidence
- ORD record
- ord-dd63db389cca416abeadad502db4e6ac
- Source dataset
- uspto-grants-2010_10
- Patent number
- US07820662B2
Reaction notes
Loading…