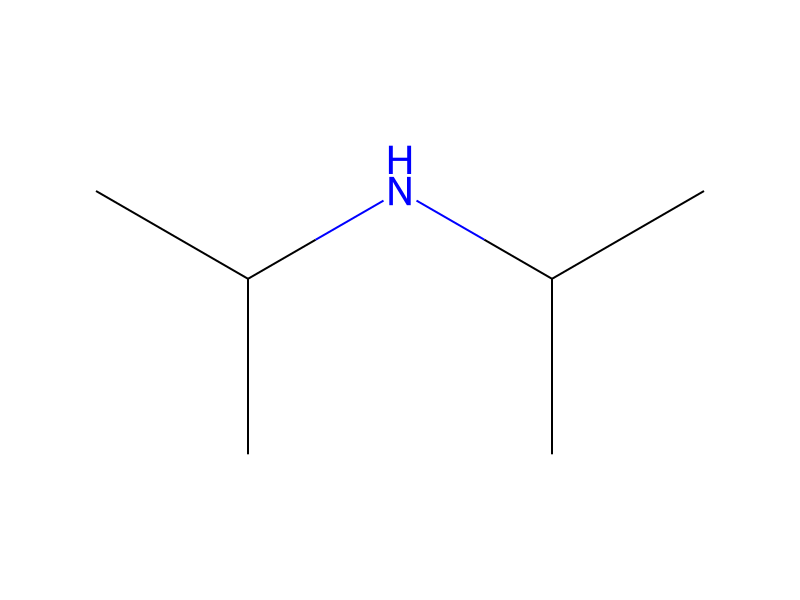
Diisopropylamine → Lithium diisopropylamide
Reaction equation

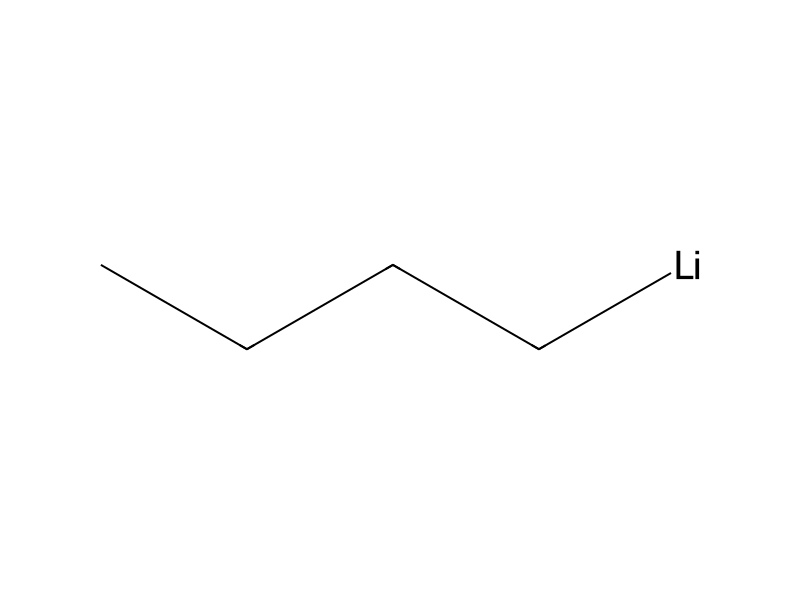
[CH:1]([NH:4][CH:5]([CH3:7])[CH3:6])([CH3:3])[CH3:2].C([Li:12])CCC.[CH2:13]([C:15]1[CH:20]=[CH:19][N:18]=[CH:17][CH:16]=1)[CH3:14].[F:21][C:22]1[CH:27]=[C:26]([F:28])[CH:25]=[CH:24][C:23]=1[C:29](=[O:36])[CH2:30][N:31]1[CH:35]=[N:34][CH:33]=[N:32]1>CCCCCC.O1CCCC1.O.C(O)(=O)C>[CH:1]([N-:4][CH:5]([CH3:7])[CH3:6])([CH3:3])[CH3:2].[Li+:12].[F:21][C:22]1[CH:27]=[C:26]([F:28])[CH:25]=[CH:24][C:23]=1[C:29]([OH:36])([CH:13]([C:15]1[CH:20]=[CH:19][N:18]=[CH:17][CH:16]=1)[CH3:14])[CH2:30][N:31]1[CH:35]=[N:34][CH:33]=[N:32]1Participants
Reactant 4


Product 2


Reagents, catalysts & solvents 4


Procedure
A solution of lithium diisopropylamide was prepared as described in Example 1(i) from diisopropylamine (40.4 g) and n-butyllithium (160 ml of a 2.5M solution in hexane) in dry tetrahydrofuran (800 ml) under an atmosphere of dry nitrogen. To this solution at -70° was added 4-ethylpyridine (42.8 g), dropwise with stirring over 0.17 hour. The solution was stirred at -70° for 0.33 hour and then a solution of 1-(2,4-difluorophenyl)-2-(1H-1,2,4-triazol-1-yl)ethanone (89.2 g) in dry tetrahydrofuran (350 ml) was added over 0.33 hour. The solution was stirred at -70° for a further 0.75 hour and then acetic acid (40 ml) was added dropwise. The solution was allowed to reach room temperature and was diluted with water. The mixture was extracted three times with ether and the combined extracts were washed with water. The aqueous washings were extracted once with ethyl acetate and the organic extracts were combined, dried (Na2SO4) and evaporated. The residue was dissolved in boiling dichloromethane, an equal volume of ether was added and then the solution was allowed to cool. The precipitated solid was filtered off to give recovered ketone starting material (17.5 g). The filtrate was evaporated and the residue was chromatographed on silica gel. Initial elution with ethyl acetate/hexane (1:1) gave further recovered ketone starting material. Further elution with ethyl acetate gave fractions containing the title compound, diastereoisomeric pair A (not treated further). The solvent was then changed to ethyl acetate/methanol (19:1) and elution was continued until pure fractions containing the title compound, diastereoisomeric pair B, were obtained. These fractions were combined, evaporated and the residue was crystallised from dichloromethane/ether to give the title compound, diastereoisomeric pair B, (20.5 g), m.p. 155°-157° (N.M.R. [300 MHz] spectrum identical with that obtained for a sample of diastereoisomeric pair B prepared as described in Method A, part (ii)).
Workup
- stirringThe solution was stirred at -70° for 0.33 hour
- stirringThe solution was stirred at -70° for a further 0.75 hour
- customto reach room temperature
- extractionThe mixture was extracted three times with ether
- washthe combined extracts were washed with water
- extractionThe aqueous washings were extracted once with ethyl acetate
- dry with materialdried (Na2SO4)
- customevaporated
- dissolutionThe residue was dissolved
- additionan equal volume of ether was added
- temperatureto cool
- filtrationThe precipitated solid was filtered off
- customto give
- customrecovered ketone starting material (17.5 g)
- customThe filtrate was evaporated
- customthe residue was chromatographed on silica gel
- washInitial elution with ethyl acetate/hexane (1:1)
- customgave further
- customrecovered ketone starting material
- washFurther elution with ethyl acetate
- customgave fractions
- additioncontaining the title compound, diastereoisomeric pair A (not
- additiontreated further)
- washThe solvent was then changed to ethyl acetate/methanol (19:1) and elution
- additioncontaining the title compound, diastereoisomeric pair B
- customwere obtained
- customevaporated
- customthe residue was crystallised from dichloromethane/ether
Sources & evidence
- ORD record
- ord-a520c89e147f478383d7d04008be560e
- Source dataset
- uspto-grants-1992_05
- Patent number
- US05116844
Reaction notes
Loading…