Nitric Acid → Isosorbide Mononitrate
Reaction equation

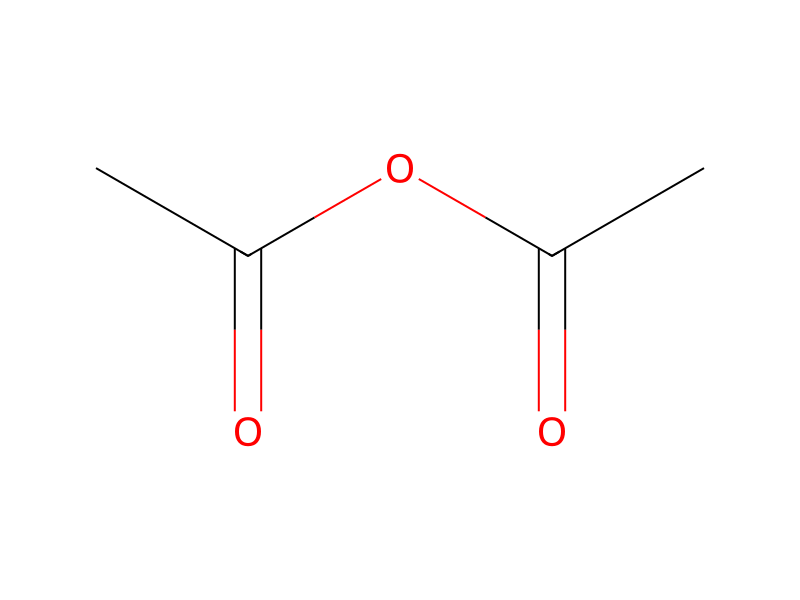
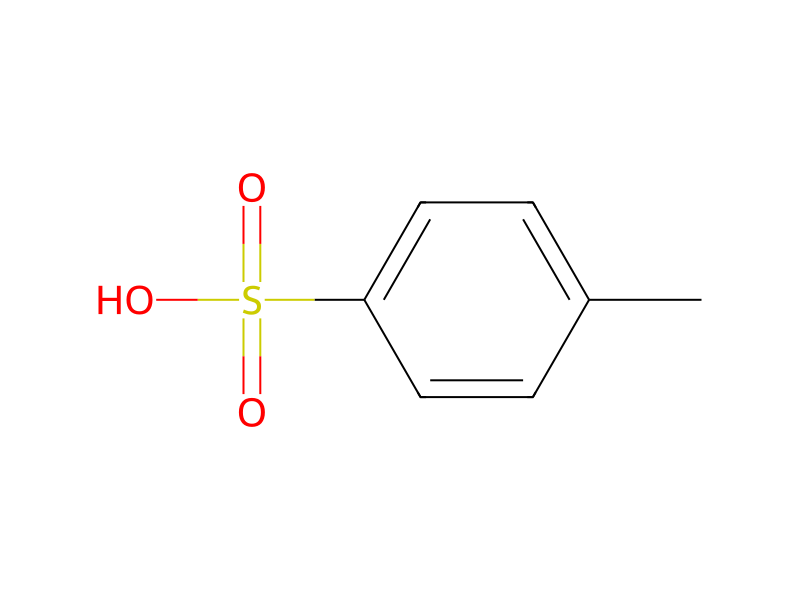
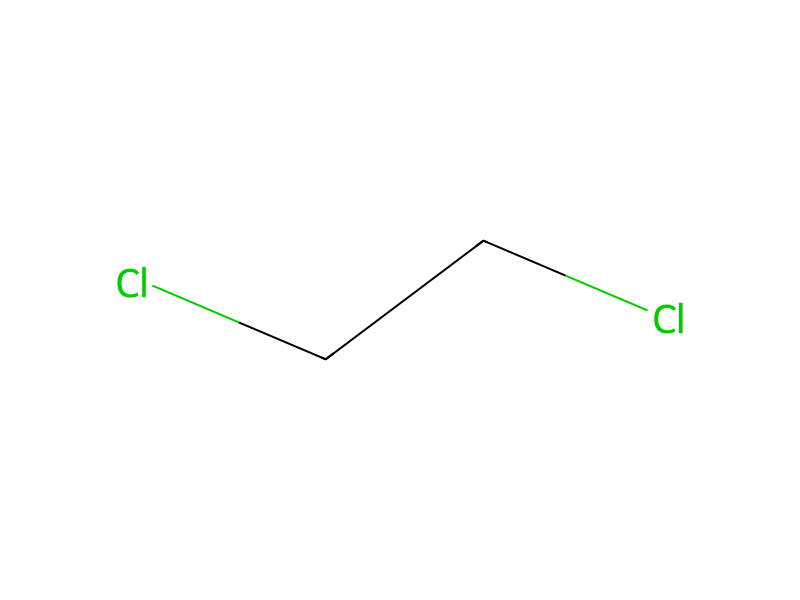
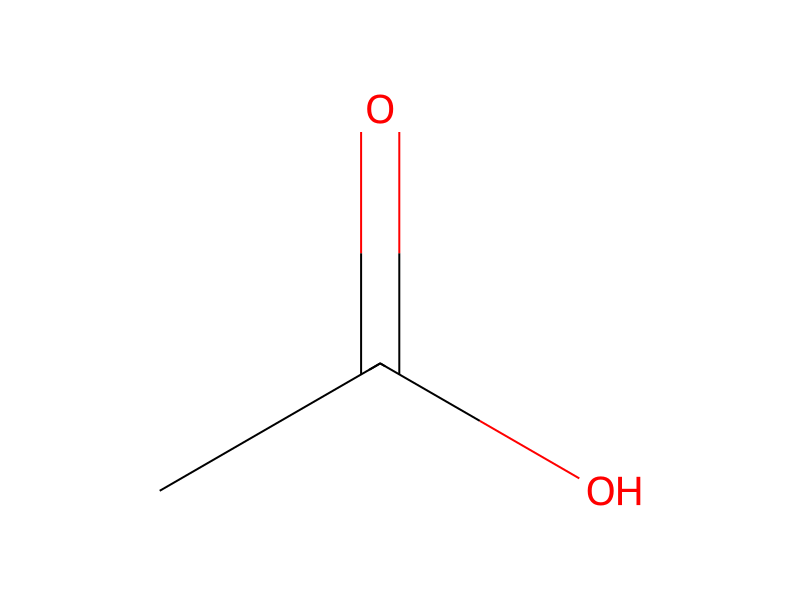
[CH2:1]1[O:5][C@@H:4]2[C@@H:6]([OH:9])[CH2:7][O:8][C@@H:3]2[C@@H:2]1[OH:10].[N+:11]([O-])([OH:13])=[O:12].C(OC(=O)C)(=O)C>C(Cl)CCl.C1(C)C=CC(S(O)(=O)=O)=CC=1.C(O)(=O)C.O>[CH2:1]1[O:5][C@@H:4]2[C@H:6]([O:9][N+:11]([O-:13])=[O:12])[CH2:7][O:8][C@@H:3]2[C@H:2]1[OH:10]Participants




Conditions
- Temperature
- 10 °C
Procedure
An admixture of 146 g of isosorbide, 60 g of glacial acetic acid and 5 g of p-toluenesulphonic acid in 700 ml of ethylene chloride are boiled under reflux, with the interposition of a water separator, until no more water is split off. The reaction mixture is then concentrated; the residue is taken up in 400 ml of methylene chloride and extracted by shaking twice with 100 ml of aqueous concentrated sodium bicarbonate solution. The aqueous phases are extracted by shaking twice with 200 ml of methylene chloride. The methylene chloride phases are combined and dried with sodium sulphate. The methylene chloride solution is then cooled to approximately 10° C. The nitration mixture--prepared by adding 61 g of 100% strength nitric acid dropwise at from -5° to 0° C. to acetic anhydride (108 g) and glacial acetic acid (64 g) which have been cooled to -5° C.--is pre-cooled to -5° C. and is then added dropwise, while stirring, the internal temperature being kept below 20° C. After stirring for a further 2 hours at 20° C., the mixture is poured into approximately 1.2 l of ice water, the organic phase is separated off, the aqueous phase is extracted by shaking once with 200 ml of methylene chloride, and the combined organic phases are extracted by shaking with 300 ml of water and are concentrated (thin-layer chromatography confirms that no explosive isosorbide dinitrate is present). The residue is dissolved in 700 ml of ethanol and warmed to 50° C., a solution of 10 g of potassium hydroxide in 100 ml of ethanol is added, and the mixture is stirred for 10 minutes at 50° C. After cooling to 20° C., the mixture is neutralized with dilute hydrochloric acid and concentrated. The residue is taken up in 500 ml of water and extracted by shaking four times with 100 ml of methylene chloride. The methylene chloride phases are combined, dried over sodium sulphate and concentrated. The residue is recrystallized from a mixture of 3 parts of chloroform and 1 part of carbon tetrachloride and gives 80.2 g of isosorbide-5-nitrate in the form of colorless, felted needles of melting point: 90° to 91° C.
Workup
- customis split off
- concentrationThe reaction mixture is then concentrated
- extractionextracted
- extractionThe aqueous phases are extracted
- stirringby shaking twice with 200 ml of methylene chloride
- dry with materialdried with sodium sulphate
- customThe nitration mixture--prepared
- temperaturehave been cooled to -5° C.
- temperatureis pre-cooled to -5° C.
- additionis then added dropwise
- stirringwhile stirring
- custombeing kept below 20° C
- stirringAfter stirring for a further 2 hours at 20° C.
- customthe organic phase is separated off
- extractionthe aqueous phase is extracted
- stirringby shaking once with 200 ml of methylene chloride
- extractionthe combined organic phases are extracted
- stirringby shaking with 300 ml of water
- concentrationare concentrated (thin-layer chromatography
- dissolutionThe residue is dissolved in 700 ml of ethanol
- temperaturewarmed to 50° C.
- additiona solution of 10 g of potassium hydroxide in 100 ml of ethanol is added
- stirringthe mixture is stirred for 10 minutes at 50° C
- temperatureAfter cooling to 20° C.
- concentrationconcentrated
- extractionextracted
- stirringby shaking four times with 100 ml of methylene chloride
- dry with materialdried over sodium sulphate
- concentrationconcentrated
- customThe residue is recrystallized from a mixture of 3 parts of chloroform and 1 part of carbon tetrachloride
Sources & evidence
- ORD record
- ord-c599885efa6c488ca64c063c8755619d
- Source dataset
- uspto-grants-1984_02
- Patent number
- US04431830
Reaction notes
Loading…