Trimethylsilyl triflate → HCID 67960584, compound
Reaction equation

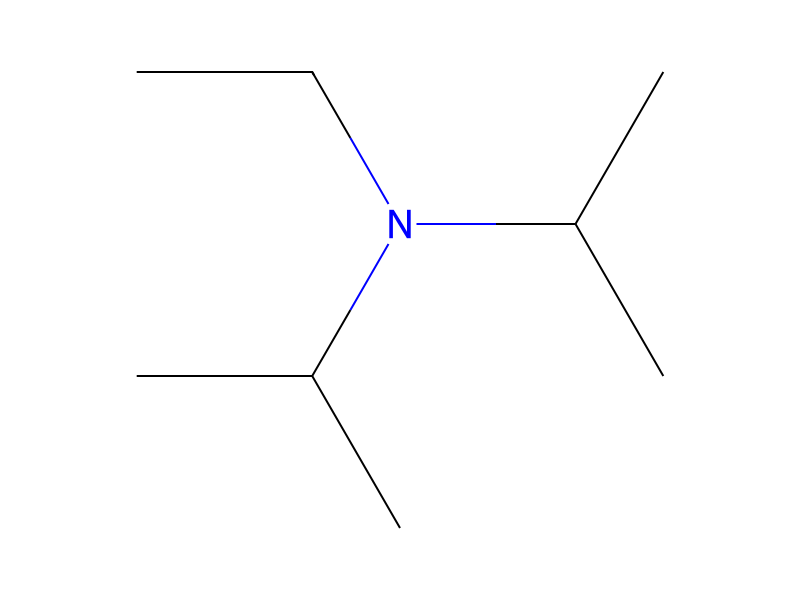
[Cl:1][C:2]([Cl:47])([Cl:46])[CH2:3][O:4][C:5]([C@@H:7]1[CH2:12][CH2:11][CH2:10][N:9]([C:13](=[O:45])[C@@H:14]([NH:30][C:31](=[O:44])[C@@H:32]([NH:36]C(OC(C)(C)C)=O)[CH:33]([CH3:35])[CH3:34])[CH2:15][C:16]2[CH:21]=[CH:20][CH:19]=[C:18]([O:22][Si:23]([C:26]([CH3:29])([CH3:28])[CH3:27])([CH3:25])[CH3:24])[CH:17]=2)[NH:8]1)=[O:6].FC(F)(F)S(O[Si](C)(C)C)(=O)=O.C(N(CC)C(C)C)(C)C>ClCCl>[Cl:47][C:2]([Cl:1])([Cl:46])[CH2:3][O:4][C:5]([C@@H:7]1[CH2:12][CH2:11][CH2:10][N:9]([C:13](=[O:45])[C@@H:14]([NH:30][C:31](=[O:44])[C@@H:32]([NH2:36])[CH:33]([CH3:35])[CH3:34])[CH2:15][C:16]2[CH:21]=[CH:20][CH:19]=[C:18]([O:22][Si:23]([C:26]([CH3:27])([CH3:28])[CH3:29])([CH3:25])[CH3:24])[CH:17]=2)[NH:8]1)=[O:6]Participants
Reactant 3


Product 1

Reagents, catalysts & solvents 1

Conditions
- Temperature
- 0 °C
Procedure
A solution of (S)-1-{(S)-2-((S)-2-tert-butoxycarbonylamino-3-methyl-butyrylamino)-3-[3-(tert-butyl-dimethyl-silanyloxy)-phenyl]-propionyl}-hexahydro-pyridazine-3-carboxylic acid 2,2,2-trichloro-ethyl ester (2.21 g, 3 mmol) in dichloromethane (30 mL) was stirred at 0° C. under nitrogen. Trimethylsilyl trifluoromethanesulfonate (1.00 g, 0.82 mL, 4.5 mmol) was added and the reaction mixture was stirred at 0° C. for 40 minutes. N,N-Diisopropylethylamine (1.55 g, 2.1 mL, 12 mmol) was added and the solvent was evaporated to afford (S)-1-{(S)-2-((S)-2-amino-3-methyl-butyrylamino)-3-[3-(tert-butyl-dimethyl-silanyloxy)-phenyl]-propionyl}-hexahydro-pyridazine-3-carboxylic acid 2,2,2-trichloro-ethyl ester (3 mmol), which was used crude in the next step. A mixture of crude (S)-1-{(S)-2-((S)-2-amino-3-methyl-butyrylamino)-3-[3-(tert-butyl-dimethyl-silanyloxy)-phenyl]-propionyl}-hexahydro-pyridazine-3-carboxylic acid 2,2,2-trichloro-ethyl ester (3 mmol) in acetonitrile (30 mL) was stirred at 0° C. under nitrogen. 4-Pentenoic acid (330 mg, 3.3 mmol) and N,N-diisopropylethylamine (1.55 g, 2.1 mL, 12 mmol) was added followed by 2-(1H-7-azabenzotriazol-1-yl)-1,1,3,3-tetramethyl uronium hexafluorophosphate methanaminium (1.6 g, 4.2 mmol) and the reaction mixture was stirred at room temperature for 3 hours. The solvent was evaporated and the residue was partitioned between ethyl acetate and water. The organic extract was separated, washed with brine, dried over anhydrous sodium sulfate and evaporated. The residue was purified by silica gel chromatography using a gradient of iso-hexanes/ethyl acetate 3:1 to 1:3 to afford the title compound (1.46 g, 68%) as a gum. 1H NMR (300 MHz, CDCl3) δ0.20 (s, 6H), 0.92 (d, J=6.2 Hz, 3H), 0.94 (d, J=6.3 Hz, 3H), 0.99 (s, 9H), 1.50-2.50 (m, 10H), 2.80-3.02 (m, 3H), 3.50 (d, J=10.7 Hz, 1H), 4.20-4.26 (m, 1H), 4.31 (dd, J=8.5, 6.0 Hz, 1H), 4.62 (d, J=12.0 Hz, 1H), 4.92 (d, J=12.0 Hz, 1H), 5.03 (dd, J=10.2, 1.5 Hz, 1H), 5.10 (dd, J=17.2, 1.5 Hz, 1H), 5.70-5.90 (m, 2H), 6.10 (d, J=8.5 Hz, 1H), 6.43 (d, J=8.3 Hz, 1H), 6.68-6.74 (m, 2H), 6.82 (d, J=7.6 Hz, 1H), 7.16 (t, J=7.8 Hz, 1H). LCMS (m/z) 719/721 [M+H], Tr=3.89 min.
Workup
- customthe solvent was evaporated
Sources & evidence
- ORD record
- ord-a2ea22d13d8849999d5bb1fff97cb1c3
- Source dataset
- uspto-grants-2015_01
- Patent number
- US08933015B2
Reaction notes
Loading…