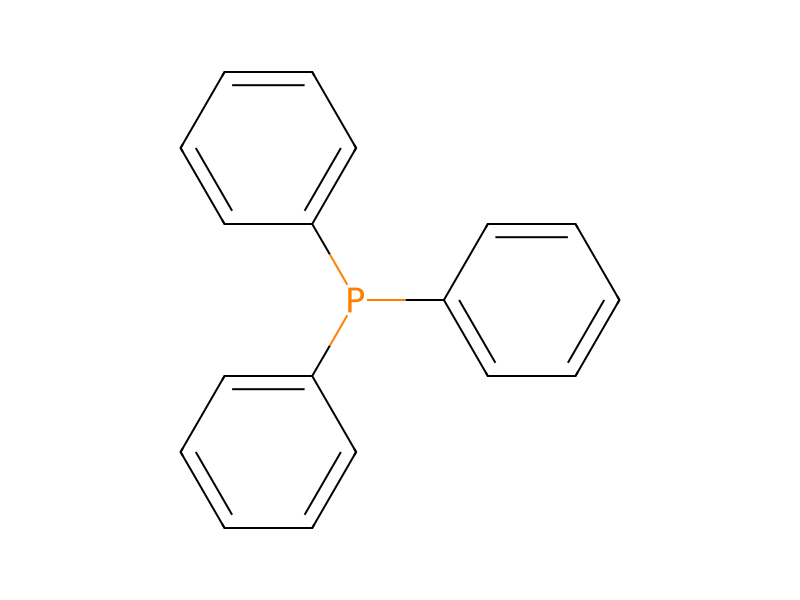
Triphenylphosphine → Triphenylphosphane dibromide
Reaction equation

[Br:1]Br.[C:3]1([P:9]([C:16]2[CH:21]=[CH:20][CH:19]=[CH:18][CH:17]=2)[C:10]2[CH:15]=[CH:14][CH:13]=[CH:12][CH:11]=2)[CH:8]=[CH:7][CH:6]=[CH:5][CH:4]=1>CC(N(C)C)=O>[Br-:1].[Br-:1].[C:16]1([P:9]([C:3]2[CH:4]=[CH:5][CH:6]=[CH:7][CH:8]=2)[C:10]2[CH:15]=[CH:14][CH:13]=[CH:12][CH:11]=2)[CH:17]=[CH:18][CH:19]=[CH:20][CH:21]=1Participants
Reactant 2
Product 1

Reagents, catalysts & solvents 1

Procedure
Bromine (59.6 g, 0.373 mole) was added drop wise over a 30-min period to a stirred solution of triphenylphosphine (97.7 g, 0.373 mole) in anhydrous 486 ml of dimethylacetamide (DMAC) kept at about 10° C. (ice bath) and protected from atmospheric moisture. (Bromine remaining in the funnel was rinsed with 10 ml of DMAC). A smooth suspension containing finely divided, crystalline triphenylphosphine dibromide resulted. The 2,4-diamino-6-pteridinemethanol.HBr (2) (25.4 g, 0.093 mole) described above was added in one portion through a powder funnel (with the aid of 10 ml DMAC). The ice bath was removed, and the stirred mixture was allowed to warm to 20-25° C. After about 1 hr, complete solution had occurred. The solution, which gradually developed a dark-red color, was kept at 20-25° C. for 1 hr longer and was then chilled (ice bath) before it was treated with EtOH (72 ml). After overnight refrigeration, the solvents were removed by evaporation in vacuo. The dark, semisolid residue was stirred with two 300-ml portions of benzene (to remove triphenylphosphine oxide), and each portion was removed from the benzene-insoluble product by decantation. The solid that remained was dissolved with stirring in glacial AcOH (660 ml) which had been preheated to 80° C. The mixture was kept in a bath at 80° C. until solution was complete. A tan crystalline solid separated as the dark solution was allowed to cool. Overnight refrigeration caused the AcOH to partially freeze. When it had thawed, the solid was collected, washed with chilled AcOH followed by Et2O, and dried in vacuo (over P2O5 and NaOH pellets) at successive temperatures of 25° C., 56° C., and 110° C. (The higher temperature was necessary for complete removal of AcOH). The yield was 15.3 g (49%). (Some runs afforded 60% yield). This sample was further purified by reprecipitation from MeOH solution (Norit) by addition of Et2O followed by drying in vacuo (25° C., P2O5), yield 13.0 g (42%) of pale-yellow solid. Spectral data: λmax, nm (ε×10−3), 0.1 N HCl, 249 (17.3), 339 (10.5), 353 (sh); pH 7, 258 (21.2), 370 (6.87); 0.1 N NaOH, 258 (21.5), 370 (6.94); 1H NMR (CF3CO2D), δ 4.70 (s, 2, CH2) and δ 9.08 (s, 1, C7—H); estimated proportion relative to the methyl-substituted contaminant, 25:1. The preparation of 2 described above is typical of several runs that gave similar yields of material whose 1H NMR spectra differed only slightly in the estimated proportion of 2 with respect to the methyl-substituted contaminant. The proportions usually ranged from 16:1 to 25:1, which corresponds to a percentage of 2 of 94 to 96%.
Workup
- customkept at about 10° C.
- wash(Bromine remaining in the funnel was rinsed with 10 ml of DMAC)
- additionA smooth suspension containing finely
Sources & evidence
- ORD record
- ord-4223c37d866c44e1963001af17cffdc6
- Source dataset
- uspto-grants-2009_11
- Patent number
- US07612071B2
Reaction notes
Loading…