Hydrochloric Acid → t-Butyl 3-(1-carboxycyclopentyl)propanoate
Reaction equation

C(NC(C)C)(C)C.C([Li])CCC.[CH:13]1([C:18]([OH:20])=[O:19])[CH2:17][CH2:16][CH2:15][CH2:14]1.Br[CH2:22][CH2:23][C:24]([O:26][C:27]([CH3:30])([CH3:29])[CH3:28])=[O:25].Cl>O1CCCC1.CCCCCC.C(OCC)C>[C:27]([O:26][C:24]([CH2:23][CH2:22][C:13]1([C:18]([OH:20])=[O:19])[CH2:17][CH2:16][CH2:15][CH2:14]1)=[O:25])([CH3:30])([CH3:29])[CH3:28]Participants
Reactant 5
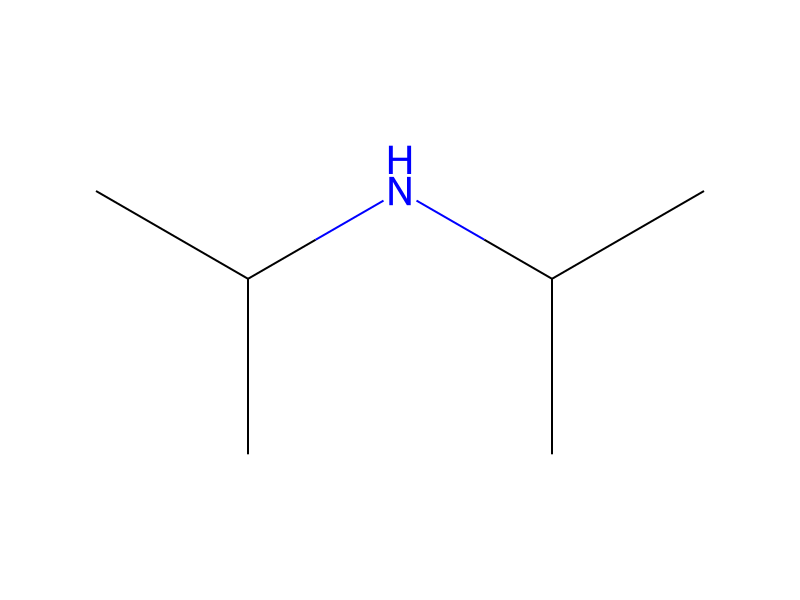


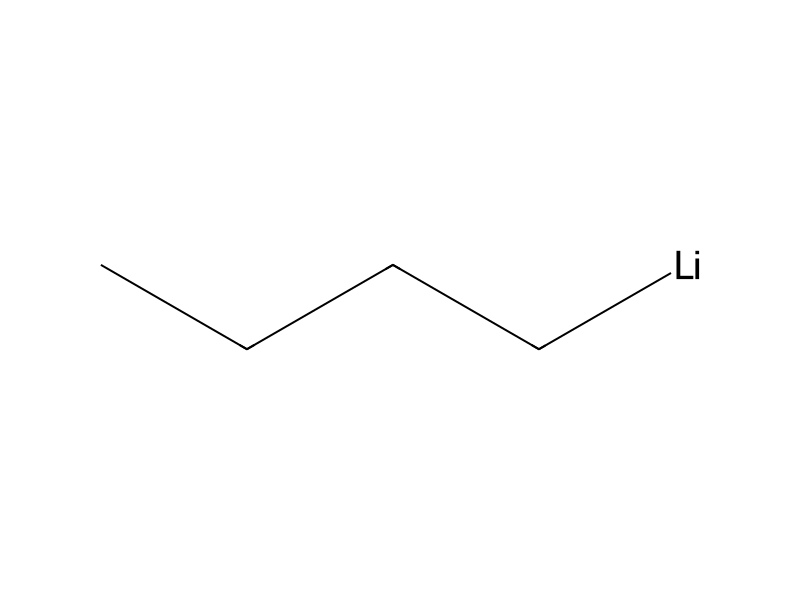
Product 1

Reagents, catalysts & solvents 3

Conditions
- Temperature
- -70 °C
Procedure
50.4 ml of diisopropylamine were dissolved in 300 ml of absolute tetrahydrofuran under a nitrogen atmosphere, and the solution was cooled to -70° C. At this temperature, 200 ml of a 1.6 molar solution of butyllithium in n-hexane were slowly added dropwise to the solution. The reaction mixture was allowed to warm to 0° C., stirred at this temperature for 30 minutes and again cooled to -20° C. At this temperature, a solution of 16.2 ml of cyclopentanecarboxylic acid in 30 ml of absolute tetrahydrofuran was added dropwise. The reaction mixture was then stirred at room temperature for 2 hours. The mixture was subsequently cooled to -10° C. and slowly added dropwise to a solution, cooled to -10° C., of 35 g of tert-butyl 3-bromopropionate in 100 ml of tetrahydrofuran. The reaction mixture was stirred at room temperature for several hours. For working up, it was acidified with dilute aqueous hydrochloric acid solution and diluted with 375 ml of diethyl ether. The organic phase was separated, and the aqueous phase was extracted three more times with 100 ml of diethyl ether each time. The combined organic extracts were washed with aqueous sodium chloride solution, dried over sodium sulfate and evaporated under reduced pressure. The remaining residue was dissolved in 300 ml of diethyl ether. The solution was shaken six times with aqueous sodium bicarbonate solution and subsequently four times with 10% strength aqueous sodium carbonate solution. The combined sodium carbonate solutions were acidified while cooling in ice and extracted three times with 150 ml of ether each time. These ether extracts were combined with the ethereal solution, and the resulting solution was washed with aqueous sodium chloride solution, dried over sodium sulfate and concentrated under reduced pressure. The resulting crude product was crystallized from ice-cold n-hexane. 7.7 g of pure 1-[2-(tert-butoxycarbonyl)ethyl]-1-cyclopentanecarboxylic acid were obtained with a melting point of 78° to 81° C.
Workup
- temperatureto warm to 0° C.
- temperatureagain cooled to -20° C
- stirringThe reaction mixture was then stirred at room temperature for 2 hours
- temperatureThe mixture was subsequently cooled to -10° C.
- additionslowly added dropwise to a solution
- stirringThe reaction mixture was stirred at room temperature for several hours
- customThe organic phase was separated
- extractionthe aqueous phase was extracted three more times with 100 ml of diethyl ether each time
- washThe combined organic extracts were washed with aqueous sodium chloride solution
- dry with materialdried over sodium sulfate
- customevaporated under reduced pressure
- dissolutionThe remaining residue was dissolved in 300 ml of diethyl ether
- stirringThe solution was shaken six times with aqueous sodium bicarbonate solution and subsequently four times with 10% strength aqueous sodium carbonate solution
- temperaturewhile cooling in ice
- extractionextracted three times with 150 ml of ether each time
- washthe resulting solution was washed with aqueous sodium chloride solution
- dry with materialdried over sodium sulfate
- concentrationconcentrated under reduced pressure
- customThe resulting crude product was crystallized from ice-cold n-hexane
Sources & evidence
- ORD record
- ord-38af31bf3784440b91151a582192a788
- Source dataset
- uspto-grants-1997_10
- Patent number
- US05677297
Reaction notes
Loading…