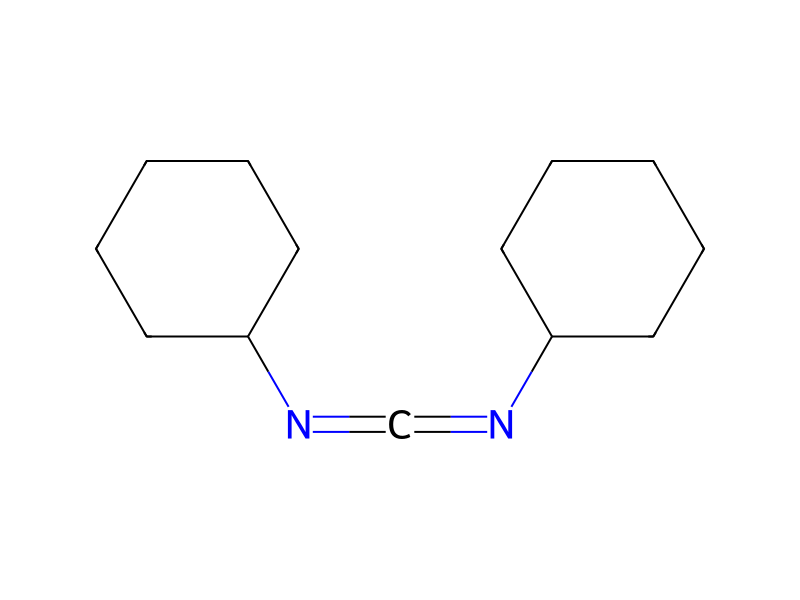
Dicyclohexylcarbodiimide → HCID 23729488, compound
Reaction equation

[OH:1][CH2:2][CH2:3][N:4]([CH3:24])[CH2:5][CH2:6][CH2:7][CH2:8][CH2:9][CH2:10][CH2:11][C:12]([NH:14][C:15]1[CH:23]=[CH:22][C:18]([C:19]([OH:21])=O)=[CH:17][CH:16]=1)=[O:13].C(N(CC)C(C)C)(C)C.C1(N=C=NC2CCCCC2)CCCCC1.O.ON1C2C=CC=CC=2N=N1.[NH2:60][CH2:61][C:62]1[C:67]([CH2:68][CH3:69])=[N:66][C:65]2[N:70]([CH2:73][CH3:74])[N:71]=[CH:72][C:64]=2[C:63]=1[NH:75][CH:76]1[CH2:81][CH2:80][O:79][CH2:78][CH2:77]1>CN(C)C=O>[CH2:73]([N:70]1[C:65]2=[N:66][C:67]([CH2:68][CH3:69])=[C:62]([CH2:61][NH:60][C:19](=[O:21])[C:18]3[CH:17]=[CH:16][C:15]([NH:14][C:12](=[O:13])[CH2:11][CH2:10][CH2:9][CH2:8][CH2:7][CH2:6][CH2:5][N:4]([CH2:3][CH2:2][OH:1])[CH3:24])=[CH:23][CH:22]=3)[C:63]([NH:75][CH:76]3[CH2:77][CH2:78][O:79][CH2:80][CH2:81]3)=[C:64]2[CH:72]=[N:71]1)[CH3:74]Participants
Reactant 5


Product 1

Reagents, catalysts & solvents 1
Procedure
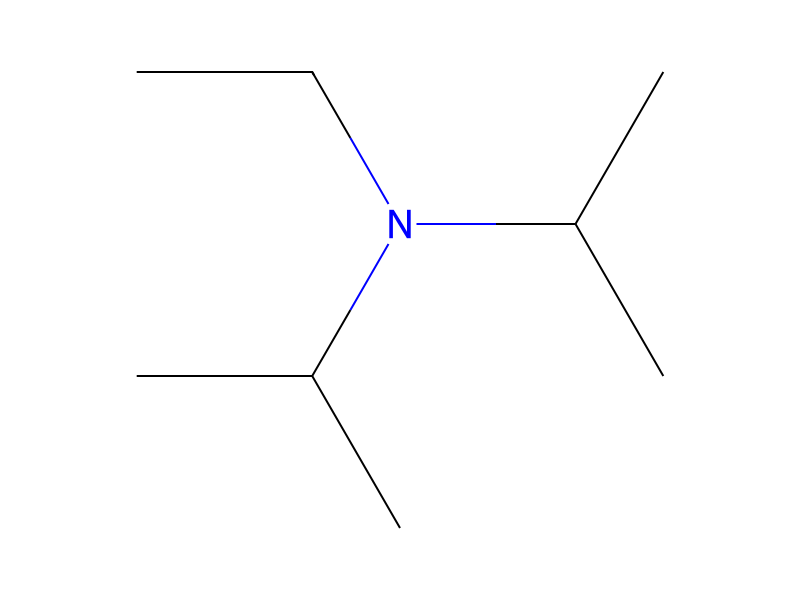
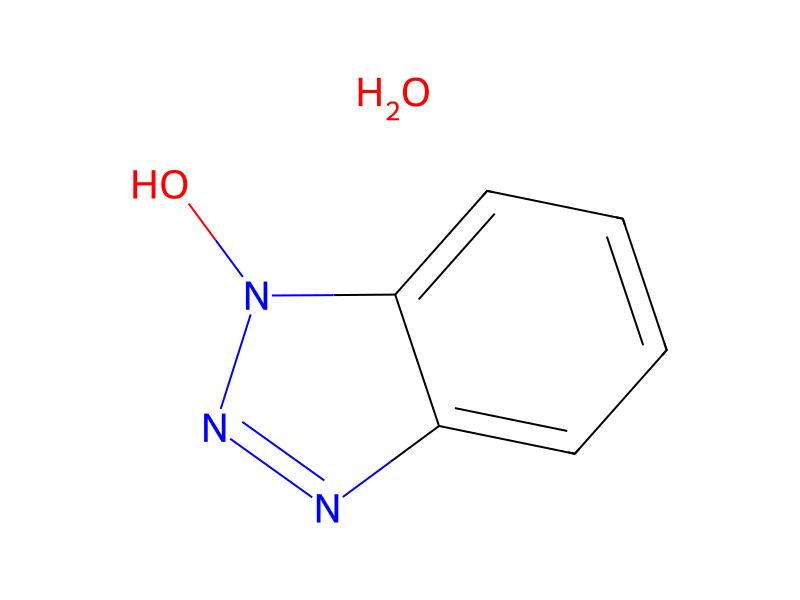
A suspension of 4-({8-[(2-hydroxyethyl)(methyl)amino]octanoyl}amino)benzoic acid (7.38 g, 19.8 mmol, 1.2 equivalents, e.g. which can be as prepared in Intermediate 39) in N,N-dimethylformamide (75 ml, about 10 volumes cf. the weight of the acid) was treated with N,N-diisopropylethylamine (4.89 ml, 3.62 g, 28.1 mmol, 1.7 equivalents) resulting in a clear yellow solution. N,N′-Dicyclohexylcarbodiimide (4.42 g, 21.45 mmol, 1.3 equivalents) and then 1-hydroxybenzotriazole hydrate (3.28 g, 21.45 mmol, 1.3 equivalents) were added, and the mixture was stirred at room temperature for 15 minutes. 5-(Aminomethyl)-1,6-diethyl-N-(tetrahydro-2H-pyran-4-yl)-1H-pyrazolo[3,4-b]pyridin-4-amine (5 g, 16.5 mmol, 1 equivalent, e.g. which can optionally be as prepared in Intermediate 7 including alternative preparation no. 1 thereof) was then added, resulting in a cloudy yellow/orange solution which was allowed to stir at room temperature overnight. As much N,N-dimethylformamide as possible was evaporated off, the residue was partitioned between dichloromethane and sodium bicarbonate solution, and then the separated aqueous phase was extracted with more dichloromethane. The combined organic extracts were washed with lithium chloride solution, water and brine, were dried (MgSO4), and were concentrated by evaporation, to give the crude product (15.5 g). 1.73 g of this crude product was removed. The remainder of the crude product was purified by column chromatography on silica gel (about 600 g), eluting with 10% methanol in dichloromethane, followed by 20% methanol in dichloromethane, 20% methanol in dichloromethane containing triethylamine, and finally 30% methanol in dichloromethane containing triethylamine. The majority of the fractions containing the product were combined and evaporated to give the title compound (6.68 g) as a white foam. LCMS showed MH+=622; TRET=2.21 min. The apparently-more-pure fractions containing the product were combined and evaporated to give more of the title compound (0.48 g).
Workup
- customresulting in a clear yellow solution
- addition1 thereof) was then added
- customresulting in a cloudy yellow/orange solution which
- stirringto stir at room temperature overnight
- customAs much N,N-dimethylformamide as possible was evaporated off
- customthe residue was partitioned between dichloromethane and sodium bicarbonate solution
- extractionthe separated aqueous phase was extracted with more dichloromethane
- washThe combined organic extracts were washed with lithium chloride solution, water and brine
- dry with materialwere dried (MgSO4)
- concentrationwere concentrated by evaporation
- customto give the crude product (15.5 g)
- custom1.73 g of this crude product was removed
- customThe remainder of the crude product was purified by column chromatography on silica gel (about 600 g)
- washeluting with 10% methanol in dichloromethane
- additioncontaining triethylamine, and finally 30% methanol in dichloromethane
- additioncontaining triethylamine
- additionThe majority of the fractions containing the product
- customevaporated
Sources & evidence
- ORD record
- ord-0af897c072934735a06b9ce4dabb3be3
- Source dataset
- uspto-grants-2011_08
- Patent number
- US08003663B2
Reaction notes
Loading…