Diisopropylamine → (S)-methyl 2-((S)-1-hydroxyethyl)-5-methylhex-4-enoate
Reaction equation

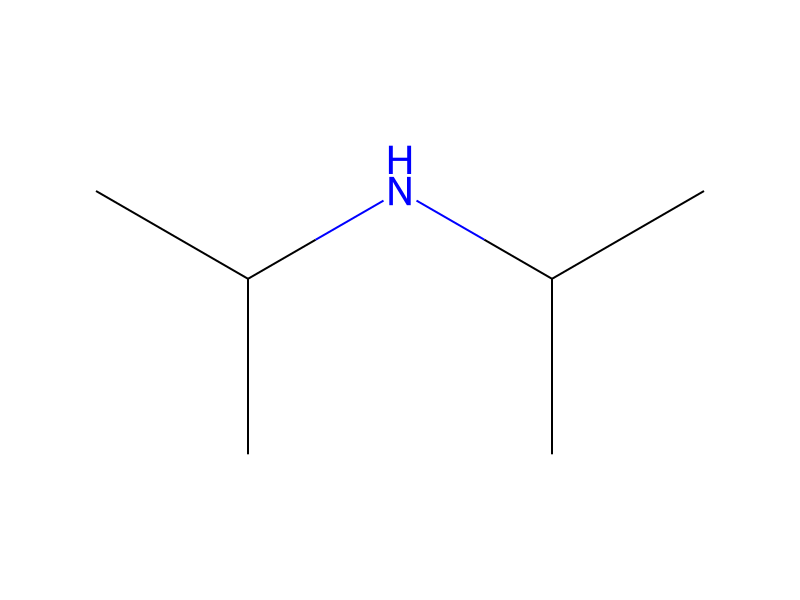
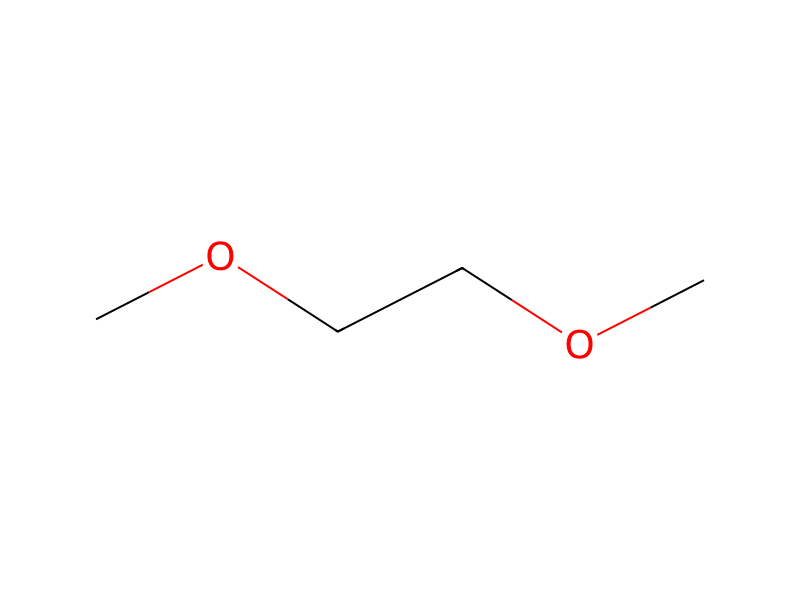
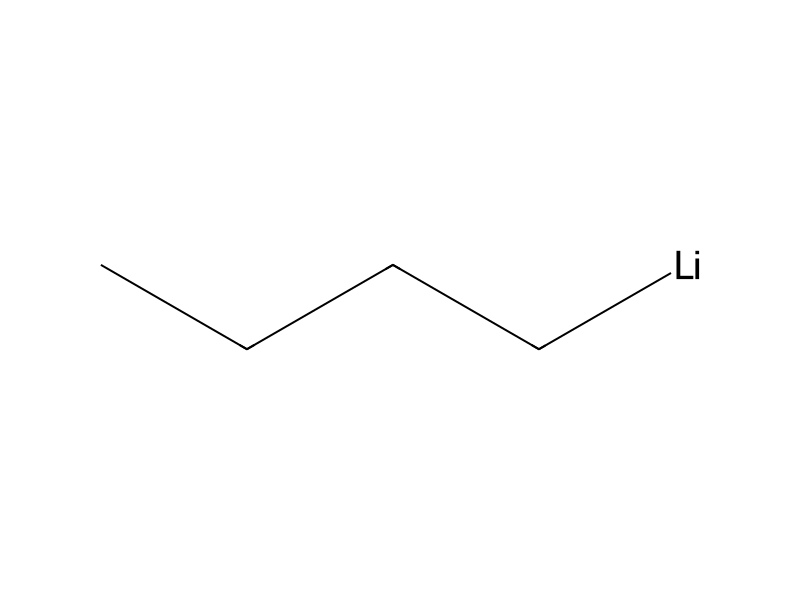
C(N[CH:5]([CH3:7])[CH3:6])(C)C.C(=O)=O.[CH3:11][C:12](C)=O.C([Li])CCC.C([N-]C(C)C)(C)C.[Li+].[OH:28][C@@H:29]([CH3:35])[CH2:30][C:31]([O:33][CH3:34])=[O:32].BrCC=C(C)C.COCCOC>C1COCC1>[OH:28][C@H:29]([C@H:30]([CH2:11][CH:12]=[C:5]([CH3:6])[CH3:7])[C:31]([O:33][CH3:34])=[O:32])[CH3:35]Participants
Reactant 7


Product 1

Reagents, catalysts & solvents 1

Conditions
- Temperature
- -50 °C
Procedure
To a solution of diisopropylamine (19.93 milliliters (mL), 142 millimoles (mmol)) in anhydrous THF (99 mL) at −50° C. (deficient dry ice/acetone bath) was added n-butyllithium (n-BuLi; 54.3 mL, 130 mmol, 2.5 M in hexanes). This solution was removed from the cold bath for 15 minutes (min), then re-cooled to −50° C. To the lithium diisopropylamide (LDA) was added a solution of (S)-methyl 3-hydroxybutanoate (6.64 ml, 59.3 mmol) in THF (20.0 mL) dropwise over 15 min using a cannula. This solution was allowed to warm to −30° C. over 30 min, stirred at −30° C. for 1 h, and recooled to −78° C. To the enolate was added a solution of 1-bromo-3-methylbut-2-ene (13.7 mL, 119 mmol) in anhydrous 1,2-dimethoxyethane (20.0 mL, 193 mmol) dropwise over 15 min. The cold bath was at −60° C. after 1 h at which time the reaction flask was removed from the bath and stirred without cooling for 1.5 h. The reaction was quenched by the addition of sat. aq. ammonium chloride (NH4Cl; 50 mL), diluted with EtOAc (50 mL), and the phases were separated. The aqueous phase was further extracted with EtOAc (2×50 mL) and the combined organic extracts were washed with sat. aq. sodium chloride (NaCl, brine; 50 mL), dried over sodium sulfate Na2SO4, filtered, and concentrated to dryness. The crude residue was purified by flash column chromatography (120 grams (g) silica gel (SiO2), 0→40% EtOAc/hexanes) to afford the title compound (9.5 g, 51.0 mmol, 86%) as a slightly yellow oil: IR (thin film) 3452, 2971, 2929, 1730, 1437, 1198, 1160 cm−1; 1H NMR (400 MHz, CDCl3) δ 5.11-5.01 (m, 1H), 3.92 (p, J=6.3 Hz, 1H), 3.70 (s, 3H), 2.78 (s, 1H), 2.46-2.28 (m, 3H), 1.69 (d, J=1.4 Hz, 3H), 1.62 (s, 3H), 1.23 (d, J=6.4 Hz, 3H); 13C NMR (101 MHz, CDCl3) δ 175.54, 134.14, 120.30, 67.78, 52.72, 51.52, 27.90, 25.73, 21.46, 17.64.
Workup
- customThis solution was removed from the cold bath for 15 minutes (min)
- temperatureto warm to −30° C. over 30 min
- customrecooled to −78° C
- waitThe cold bath was at −60° C. after 1 h at which time
- customthe reaction flask was removed from the bath
- stirringstirred
- temperaturewithout cooling for 1.5 h
- customThe reaction was quenched by the addition of sat. aq. ammonium chloride (NH4Cl; 50 mL)
- additiondiluted with EtOAc (50 mL)
- customthe phases were separated
- extractionThe aqueous phase was further extracted with EtOAc (2×50 mL)
- washthe combined organic extracts were washed with sat. aq. sodium chloride (NaCl, brine; 50 mL)
- dry with materialdried over sodium sulfate Na2SO4
- filtrationfiltered
- concentrationconcentrated to dryness
- customThe crude residue was purified by flash column chromatography (120 grams (g) silica gel (SiO2), 0→40% EtOAc/hexanes)
Sources & evidence
- ORD record
- ord-bccc43aa834d4f4da8cc544b07d30c93
- Source dataset
- uspto-grants-2016_02
- Patent number
- US09247741B2
Reaction notes
Loading…